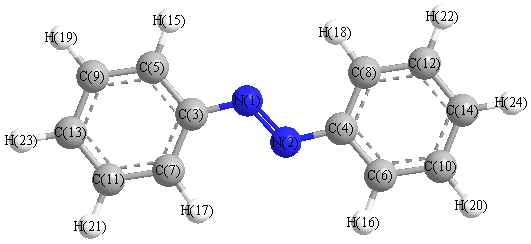
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3389 |
3086 |
0.00 |
|
|
|
| 2 |
Ag |
3366 |
3065 |
0.00 |
|
|
|
| 3 |
Ag |
3356 |
3056 |
0.00 |
|
|
|
| 4 |
Ag |
3343 |
3044 |
0.00 |
|
|
|
| 5 |
Ag |
3331 |
3034 |
0.00 |
|
|
|
| 6 |
Ag |
1869 |
1702 |
0.00 |
|
|
|
| 7 |
Ag |
1781 |
1621 |
0.00 |
|
|
|
| 8 |
Ag |
1768 |
1610 |
0.00 |
|
|
|
| 9 |
Ag |
1631 |
1485 |
0.00 |
|
|
|
| 10 |
Ag |
1593 |
1451 |
0.00 |
|
|
|
| 11 |
Ag |
1442 |
1313 |
0.00 |
|
|
|
| 12 |
Ag |
1345 |
1225 |
0.00 |
|
|
|
| 13 |
Ag |
1297 |
1181 |
0.00 |
|
|
|
| 14 |
Ag |
1261 |
1149 |
0.00 |
|
|
|
| 15 |
Ag |
1216 |
1107 |
0.00 |
|
|
|
| 16 |
Ag |
1162 |
1058 |
0.00 |
|
|
|
| 17 |
Ag |
1110 |
1010 |
0.00 |
|
|
|
| 18 |
Ag |
1079 |
983 |
0.00 |
|
|
|
| 19 |
Ag |
1000 |
911 |
0.00 |
|
|
|
| 20 |
Ag |
727 |
662 |
0.00 |
|
|
|
| 21 |
Ag |
667 |
607 |
0.00 |
|
|
|
| 22 |
Ag |
325 |
296 |
0.00 |
|
|
|
| 23 |
Ag |
238 |
217 |
0.00 |
|
|
|
| 24 |
Au |
1094 |
996 |
0.00 |
|
|
|
| 25 |
Au |
1086 |
989 |
0.01 |
|
|
|
| 26 |
Au |
1039 |
946 |
9.58 |
|
|
|
| 27 |
Au |
936 |
853 |
0.09 |
|
|
|
| 28 |
Au |
857 |
780 |
97.25 |
|
|
|
| 29 |
Au |
753 |
686 |
89.23 |
|
|
|
| 30 |
Au |
608 |
554 |
19.70 |
|
|
|
| 31 |
Au |
454 |
413 |
0.03 |
|
|
|
| 32 |
Au |
338 |
308 |
0.55 |
|
|
|
| 33 |
Au |
70 |
64 |
1.33 |
|
|
|
| 34 |
Au |
22 |
20 |
0.01 |
|
|
|
| 35 |
Bg |
1094 |
996 |
0.00 |
|
|
|
| 36 |
Bg |
1086 |
989 |
0.00 |
|
|
|
| 37 |
Bg |
1033 |
941 |
0.00 |
|
|
|
| 38 |
Bg |
937 |
853 |
0.00 |
|
|
|
| 39 |
Bg |
835 |
760 |
0.00 |
|
|
|
| 40 |
Bg |
742 |
676 |
0.00 |
|
|
|
| 41 |
Bg |
529 |
482 |
0.00 |
|
|
|
| 42 |
Bg |
456 |
416 |
0.00 |
|
|
|
| 43 |
Bg |
276 |
252 |
0.00 |
|
|
|
| 44 |
Bg |
100 |
91 |
0.00 |
|
|
|
| 45 |
Bu |
3389 |
3086 |
5.56 |
|
|
|
| 46 |
Bu |
3366 |
3065 |
28.67 |
|
|
|
| 47 |
Bu |
3355 |
3055 |
48.37 |
|
|
|
| 48 |
Bu |
3343 |
3044 |
21.52 |
|
|
|
| 49 |
Bu |
3331 |
3033 |
1.21 |
|
|
|
| 50 |
Bu |
1788 |
1628 |
5.34 |
|
|
|
| 51 |
Bu |
1769 |
1611 |
3.64 |
|
|
|
| 52 |
Bu |
1638 |
1492 |
31.27 |
|
|
|
| 53 |
Bu |
1588 |
1446 |
13.68 |
|
|
|
| 54 |
Bu |
1435 |
1306 |
2.43 |
|
|
|
| 55 |
Bu |
1356 |
1235 |
1.95 |
|
|
|
| 56 |
Bu |
1327 |
1209 |
12.76 |
|
|
|
| 57 |
Bu |
1260 |
1148 |
9.80 |
|
|
|
| 58 |
Bu |
1210 |
1102 |
20.86 |
|
|
|
| 59 |
Bu |
1165 |
1061 |
4.34 |
|
|
|
| 60 |
Bu |
1110 |
1011 |
8.58 |
|
|
|
| 61 |
Bu |
1078 |
982 |
0.76 |
|
|
|
| 62 |
Bu |
888 |
808 |
1.41 |
|
|
|
| 63 |
Bu |
673 |
612 |
0.65 |
|
|
|
| 64 |
Bu |
581 |
529 |
3.33 |
|
|
|
| 65 |
Bu |
562 |
512 |
25.90 |
|
|
|
| 66 |
Bu |
90 |
82 |
1.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 44470.6 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 40494.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.645 |
|
|
|
| 2 |
N |
-0.645 |
|
|
|
| 3 |
C |
0.790 |
|
|
|
| 4 |
C |
0.790 |
|
|
|
| 5 |
C |
0.088 |
|
|
|
| 6 |
C |
0.088 |
|
|
|
| 7 |
C |
0.287 |
|
|
|
| 8 |
C |
0.287 |
|
|
|
| 9 |
C |
0.684 |
|
|
|
| 10 |
C |
0.684 |
|
|
|
| 11 |
C |
0.698 |
|
|
|
| 12 |
C |
0.698 |
|
|
|
| 13 |
C |
0.559 |
|
|
|
| 14 |
C |
0.559 |
|
|
|
| 15 |
H |
-0.550 |
|
|
|
| 16 |
H |
-0.550 |
|
|
|
| 17 |
H |
-0.557 |
|
|
|
| 18 |
H |
-0.557 |
|
|
|
| 19 |
H |
-0.437 |
|
|
|
| 20 |
H |
-0.437 |
|
|
|
| 21 |
H |
-0.457 |
|
|
|
| 22 |
H |
-0.457 |
|
|
|
| 23 |
H |
-0.460 |
|
|
|
| 24 |
H |
-0.460 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-68.504 |
3.928 |
0.000 |
| y |
3.928 |
-70.337 |
0.000 |
| z |
0.000 |
0.000 |
-89.724 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
11.526 |
3.928 |
0.000 |
| y |
3.928 |
8.777 |
0.000 |
| z |
0.000 |
0.000 |
-20.303 |
|
| Polar |
|---|
| 3z2-r2 | -40.605 |
|---|
| x2-y2 | 1.833 |
|---|
| xy | 3.928 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
31.772 |
7.947 |
0.000 |
| y |
7.947 |
29.334 |
0.000 |
| z |
0.000 |
0.000 |
12.952 |
<r2> (average value of r
2) Å
2
| <r2> |
1093.007 |
| (<r2>)1/2 |
33.061 |