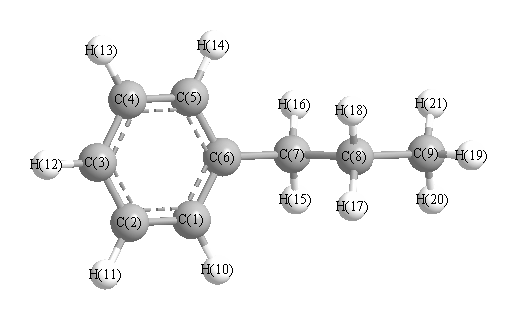
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3226 |
3091 |
14.53 |
|
|
|
| 2 |
A' |
3204 |
3070 |
4.66 |
|
|
|
| 3 |
A' |
3188 |
3055 |
9.54 |
|
|
|
| 4 |
A' |
3137 |
3006 |
33.83 |
|
|
|
| 5 |
A' |
3060 |
2932 |
52.89 |
|
|
|
| 6 |
A' |
3050 |
2923 |
33.43 |
|
|
|
| 7 |
A' |
3047 |
2920 |
6.13 |
|
|
|
| 8 |
A' |
1676 |
1606 |
9.36 |
|
|
|
| 9 |
A' |
1530 |
1466 |
13.26 |
|
|
|
| 10 |
A' |
1488 |
1426 |
7.88 |
|
|
|
| 11 |
A' |
1473 |
1411 |
0.11 |
|
|
|
| 12 |
A' |
1467 |
1406 |
0.48 |
|
|
|
| 13 |
A' |
1394 |
1336 |
2.52 |
|
|
|
| 14 |
A' |
1373 |
1316 |
1.05 |
|
|
|
| 15 |
A' |
1300 |
1246 |
0.63 |
|
|
|
| 16 |
A' |
1233 |
1181 |
0.64 |
|
|
|
| 17 |
A' |
1198 |
1148 |
0.17 |
|
|
|
| 18 |
A' |
1116 |
1069 |
6.53 |
|
|
|
| 19 |
A' |
1071 |
1026 |
0.56 |
|
|
|
| 20 |
A' |
1060 |
1015 |
3.85 |
|
|
|
| 21 |
A' |
1014 |
972 |
0.20 |
|
|
|
| 22 |
A' |
999 |
957 |
0.01 |
|
|
|
| 23 |
A' |
927 |
888 |
0.17 |
|
|
|
| 24 |
A' |
907 |
869 |
2.18 |
|
|
|
| 25 |
A' |
830 |
796 |
0.69 |
|
|
|
| 26 |
A' |
764 |
732 |
27.76 |
|
|
|
| 27 |
A' |
713 |
683 |
45.69 |
|
|
|
| 28 |
A' |
595 |
570 |
5.18 |
|
|
|
| 29 |
A' |
509 |
488 |
9.40 |
|
|
|
| 30 |
A' |
311 |
298 |
0.03 |
|
|
|
| 31 |
A' |
277 |
266 |
0.56 |
|
|
|
| 32 |
A' |
104 |
99 |
0.32 |
|
|
|
| 33 |
A" |
3213 |
3079 |
29.90 |
|
|
|
| 34 |
A" |
3190 |
3057 |
5.23 |
|
|
|
| 35 |
A" |
3131 |
3000 |
56.42 |
|
|
|
| 36 |
A" |
3108 |
2978 |
9.79 |
|
|
|
| 37 |
A" |
3086 |
2957 |
0.47 |
|
|
|
| 38 |
A" |
1653 |
1584 |
0.72 |
|
|
|
| 39 |
A" |
1483 |
1422 |
1.73 |
|
|
|
| 40 |
A" |
1476 |
1414 |
10.73 |
|
|
|
| 41 |
A" |
1382 |
1324 |
0.14 |
|
|
|
| 42 |
A" |
1345 |
1289 |
0.29 |
|
|
|
| 43 |
A" |
1311 |
1256 |
0.37 |
|
|
|
| 44 |
A" |
1239 |
1187 |
0.25 |
|
|
|
| 45 |
A" |
1174 |
1125 |
0.01 |
|
|
|
| 46 |
A" |
1122 |
1075 |
5.50 |
|
|
|
| 47 |
A" |
1070 |
1025 |
1.29 |
|
|
|
| 48 |
A" |
983 |
942 |
0.01 |
|
|
|
| 49 |
A" |
869 |
833 |
0.61 |
|
|
|
| 50 |
A" |
853 |
817 |
0.05 |
|
|
|
| 51 |
A" |
737 |
706 |
1.45 |
|
|
|
| 52 |
A" |
630 |
604 |
0.02 |
|
|
|
| 53 |
A" |
412 |
395 |
0.00 |
|
|
|
| 54 |
A" |
345 |
331 |
0.11 |
|
|
|
| 55 |
A" |
239 |
229 |
0.01 |
|
|
|
| 56 |
A" |
87 |
83 |
0.02 |
|
|
|
| 57 |
A" |
39 |
37 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40707.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 39009.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.492 |
|
|
|
| 2 |
C |
0.169 |
|
|
|
| 3 |
C |
0.504 |
|
|
|
| 4 |
C |
0.169 |
|
|
|
| 5 |
C |
0.492 |
|
|
|
| 6 |
C |
1.972 |
|
|
|
| 7 |
C |
0.322 |
|
|
|
| 8 |
C |
0.615 |
|
|
|
| 9 |
C |
0.387 |
|
|
|
| 10 |
H |
-0.826 |
|
|
|
| 11 |
H |
-0.592 |
|
|
|
| 12 |
H |
-0.576 |
|
|
|
| 13 |
H |
-0.592 |
|
|
|
| 14 |
H |
-0.826 |
|
|
|
| 15 |
H |
-0.375 |
|
|
|
| 16 |
H |
-0.375 |
|
|
|
| 17 |
H |
-0.307 |
|
|
|
| 18 |
H |
-0.307 |
|
|
|
| 19 |
H |
-0.130 |
|
|
|
| 20 |
H |
-0.107 |
|
|
|
| 21 |
H |
-0.107 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.163 |
0.418 |
0.000 |
0.448 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-59.434 |
-0.122 |
0.000 |
| y |
-0.122 |
-52.773 |
0.000 |
| z |
0.000 |
0.000 |
-51.673 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.211 |
-0.122 |
0.000 |
| y |
-0.122 |
2.780 |
0.000 |
| z |
0.000 |
0.000 |
4.431 |
|
| Polar |
|---|
| 3z2-r2 | 8.862 |
|---|
| x2-y2 | -6.661 |
|---|
| xy | -0.122 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.543 |
-1.155 |
0.000 |
| y |
-1.155 |
20.651 |
0.000 |
| z |
0.000 |
0.000 |
16.004 |
<r2> (average value of r
2) Å
2
| <r2> |
438.660 |
| (<r2>)1/2 |
20.944 |