Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -158.007998 |
| Energy at 298.15K | -158.018568 |
| HF Energy | -157.313557 |
| Nuclear repulsion energy | 129.553374 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
120 |
117 |
|
|
|
|
|
Au |
231 |
226 |
|
|
|
|
|
Bu |
264 |
258 |
|
|
|
|
|
Bg |
268 |
262 |
|
|
|
|
|
Ag |
433 |
423 |
|
|
|
|
|
Au |
746 |
729 |
|
|
|
|
|
Bg |
828 |
809 |
|
|
|
|
|
Ag |
865 |
845 |
|
|
|
|
|
Au |
981 |
959 |
|
|
|
|
|
Bu |
999 |
976 |
|
|
|
|
|
Bu |
1052 |
1028 |
|
|
|
|
|
Ag |
1103 |
1077 |
|
|
|
|
|
Ag |
1198 |
1170 |
|
|
|
|
|
Bg |
1248 |
1219 |
|
|
|
|
|
Au |
1321 |
1290 |
|
|
|
|
|
Bu |
1350 |
1318 |
|
|
|
|
|
Bg |
1358 |
1326 |
|
|
|
|
|
Ag |
1437 |
1404 |
|
|
|
|
|
Ag |
1458 |
1424 |
|
|
|
|
|
Bu |
1458 |
1425 |
|
|
|
|
|
Ag |
1537 |
1501 |
|
|
|
|
|
Bu |
1538 |
1503 |
|
|
|
|
|
Bg |
1546 |
1510 |
|
|
|
|
|
Au |
1546 |
1511 |
|
|
|
|
|
Ag |
1550 |
1514 |
|
|
|
|
|
Bu |
1559 |
1523 |
|
|
|
|
|
Ag |
3033 |
2963 |
|
|
|
|
|
Bu |
3039 |
2969 |
|
|
|
|
|
Bu |
3050 |
2980 |
|
|
|
|
|
Ag |
3051 |
2981 |
|
|
|
|
|
Bg |
3065 |
2994 |
|
|
|
|
|
Au |
3086 |
3015 |
|
|
|
|
|
Bg |
3128 |
3056 |
|
|
|
|
|
Au |
3131 |
3059 |
|
|
|
|
|
Ag |
3132 |
3060 |
|
|
|
|
|
Bu |
3133 |
3061 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29419.6 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 28740.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
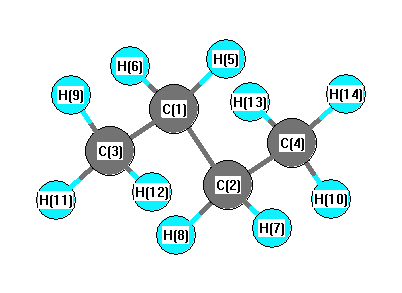
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.426 |
0.641 |
0.000 |
| C2 |
0.426 |
-0.641 |
0.000 |
| C3 |
0.426 |
1.922 |
0.000 |
| C4 |
-0.426 |
-1.922 |
0.000 |
| H5 |
-1.090 |
0.635 |
0.888 |
| H6 |
-1.090 |
0.635 |
-0.888 |
| H7 |
1.090 |
-0.635 |
0.888 |
| H8 |
1.090 |
-0.635 |
-0.888 |
| H9 |
-0.204 |
2.829 |
0.000 |
| H10 |
1.077 |
1.961 |
0.894 |
| H11 |
1.077 |
1.961 |
-0.894 |
| H12 |
0.204 |
-2.829 |
0.000 |
| H13 |
-1.077 |
-1.961 |
0.894 |
| H14 |
-1.077 |
-1.961 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5391 | 1.5393 | 2.5624 | 1.1087 | 1.1087 | 2.1715 | 2.1715 | 2.2002 | 2.1921 | 2.1921 | 3.5268 | 2.8272 | 2.8272 |
C2 | 1.5391 | | 2.5624 | 1.5393 | 2.1715 | 2.1715 | 1.1087 | 1.1087 | 3.5268 | 2.8272 | 2.8272 | 2.2002 | 2.1921 | 2.1921 | C3 | 1.5393 | 2.5624 | | 3.9372 | 2.1786 | 2.1786 | 2.7865 | 2.7865 | 1.1051 | 1.1063 | 1.1063 | 4.7565 | 4.2592 | 4.2592 | C4 | 2.5624 | 1.5393 | 3.9372 | | 2.7865 | 2.7865 | 2.1786 | 2.1786 | 4.7565 | 4.2592 | 4.2592 | 1.1051 | 1.1063 | 1.1063 | H5 | 1.1087 | 2.1715 | 2.1786 | 2.7865 | | 1.7757 | 2.5233 | 3.0855 | 2.5281 | 2.5418 | 3.1039 | 3.8031 | 2.5961 | 3.1486 | H6 | 1.1087 | 2.1715 | 2.1786 | 2.7865 | 1.7757 | | 3.0855 | 2.5233 | 2.5281 | 3.1039 | 2.5418 | 3.8031 | 3.1486 | 2.5961 | H7 | 2.1715 | 1.1087 | 2.7865 | 2.1786 | 2.5233 | 3.0855 | | 1.7757 | 3.8031 | 2.5961 | 3.1486 | 2.5281 | 2.5418 | 3.1039 | H8 | 2.1715 | 1.1087 | 2.7865 | 2.1786 | 3.0855 | 2.5233 | 1.7757 | | 3.8031 | 3.1486 | 2.5961 | 2.5281 | 3.1039 | 2.5418 | H9 | 2.2002 | 3.5268 | 1.1051 | 4.7565 | 2.5281 | 2.5281 | 3.8031 | 3.8031 | | 1.7873 | 1.7873 | 5.6736 | 4.9512 | 4.9512 | H10 | 2.1921 | 2.8272 | 1.1063 | 4.2592 | 2.5418 | 3.1039 | 2.5961 | 3.1486 | 1.7873 | | 1.7873 | 4.9512 | 4.4759 | 4.8195 | H11 | 2.1921 | 2.8272 | 1.1063 | 4.2592 | 3.1039 | 2.5418 | 3.1486 | 2.5961 | 1.7873 | 1.7873 | | 4.9512 | 4.8195 | 4.4759 | H12 | 3.5268 | 2.2002 | 4.7565 | 1.1051 | 3.8031 | 3.8031 | 2.5281 | 2.5281 | 5.6736 | 4.9512 | 4.9512 | | 1.7873 | 1.7873 | H13 | 2.8272 | 2.1921 | 4.2592 | 1.1063 | 2.5961 | 3.1486 | 2.5418 | 3.1039 | 4.9512 | 4.4759 | 4.8195 | 1.7873 | | 1.7873 | H14 | 2.8272 | 2.1921 | 4.2592 | 1.1063 | 3.1486 | 2.5961 | 3.1039 | 2.5418 | 4.9512 | 4.8195 | 4.4759 | 1.7873 | 1.7873 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.690 |
|
C1 |
C2 |
H7 |
109.112 |
| C1 |
C2 |
H8 |
109.112 |
|
C1 |
C3 |
H9 |
111.560 |
| C1 |
C3 |
H11 |
110.848 |
|
C1 |
C3 |
H12 |
30.972 |
| C2 |
C1 |
C3 |
112.690 |
|
C2 |
C1 |
H5 |
109.112 |
| C2 |
C1 |
H6 |
109.112 |
|
C2 |
C4 |
H10 |
17.325 |
| C2 |
C4 |
H13 |
110.848 |
|
C2 |
C4 |
H14 |
110.848 |
| C3 |
C1 |
H5 |
109.654 |
|
C3 |
C1 |
H6 |
109.654 |
| C4 |
C2 |
H7 |
109.654 |
|
C4 |
C2 |
H8 |
109.654 |
| H5 |
C1 |
H6 |
106.420 |
|
H7 |
C2 |
H8 |
106.420 |
| H9 |
C3 |
H11 |
107.836 |
|
H9 |
C3 |
H12 |
142.532 |
| H10 |
C4 |
H13 |
94.069 |
|
H10 |
C4 |
H14 |
114.200 |
| H11 |
C3 |
H12 |
93.629 |
|
H13 |
C4 |
H14 |
107.755 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -158.007102 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3144 |
3071 |
|
|
|
|
| 2 |
A |
3129 |
3057 |
|
|
|
|
| 3 |
A |
3077 |
3006 |
|
|
|
|
| 4 |
A |
3055 |
2985 |
|
|
|
|
| 5 |
A |
3039 |
2968 |
|
|
|
|
| 6 |
A |
1558 |
1522 |
|
|
|
|
| 7 |
A |
1553 |
1517 |
|
|
|
|
| 8 |
A |
1535 |
1500 |
|
|
|
|
| 9 |
A |
1459 |
1425 |
|
|
|
|
| 10 |
A |
1419 |
1386 |
|
|
|
|
| 11 |
A |
1341 |
1310 |
|
|
|
|
| 12 |
A |
1232 |
1203 |
|
|
|
|
| 13 |
A |
1122 |
1096 |
|
|
|
|
| 14 |
A |
1016 |
993 |
|
|
|
|
| 15 |
A |
855 |
835 |
|
|
|
|
| 16 |
A |
814 |
795 |
|
|
|
|
| 17 |
A |
327 |
320 |
|
|
|
|
| 18 |
A |
277 |
270 |
|
|
|
|
| 19 |
A |
111 |
108 |
|
|
|
|
| 20 |
B |
3136 |
3063 |
|
|
|
|
| 21 |
B |
3130 |
3058 |
|
|
|
|
| 22 |
B |
3083 |
3012 |
|
|
|
|
| 23 |
B |
3052 |
2982 |
|
|
|
|
| 24 |
B |
3036 |
2966 |
|
|
|
|
| 25 |
B |
1553 |
1517 |
|
|
|
|
| 26 |
B |
1545 |
1509 |
|
|
|
|
| 27 |
B |
1533 |
1498 |
|
|
|
|
| 28 |
B |
1460 |
1426 |
|
|
|
|
| 29 |
B |
1403 |
1371 |
|
|
|
|
| 30 |
B |
1319 |
1289 |
|
|
|
|
| 31 |
B |
1184 |
1156 |
|
|
|
|
| 32 |
B |
996 |
973 |
|
|
|
|
| 33 |
B |
987 |
964 |
|
|
|
|
| 34 |
B |
762 |
744 |
|
|
|
|
| 35 |
B |
444 |
434 |
|
|
|
|
| 36 |
B |
223 |
218 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29453.6 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 28773.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
