return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-156.782548
Energy at 298.15K-156.790446
HF Energy-156.122841
Nuclear repulsion energy115.978760
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3247 3172 0.00      
2 A 3162 3089 0.00      
3 A 3142 3070 0.00      
4 A 3141 3068 0.00      
5 A 3134 3061 0.00      
6 A 3083 3012 0.00      
7 A 3055 2985 0.00      
8 A 3037 2967 0.00      
9 A 1713 1673 0.00      
10 A 1551 1516 0.00      
11 A 1544 1508 0.00      
12 A 1532 1496 0.00      
13 A 1484 1450 0.00      
14 A 1454 1421 0.00      
15 A 1377 1345 0.00      
16 A 1336 1305 0.00      
17 A 1316 1286 0.00      
18 A 1225 1197 0.00      
19 A 1118 1092 0.00      
20 A 1060 1035 0.00      
21 A 1017 994 0.00      
22 A 1004 980 0.00      
23 A 908 887 0.00      
24 A 878 858 0.00      
25 A 807 788 0.00      
26 A 648 633 0.00      
27 A 435 425 0.00      
28 A 321 313 0.00      
29 A 240 234 0.00      
30 A 108 105 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 24037.8 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 23482.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.73479 0.13640 0.13323

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.730 -0.249 -0.302
C2 0.547 0.529 0.314
C3 -0.717 -0.304 0.348
C4 -1.872 0.021 -0.283
H5 -0.669 -1.245 0.918
H6 1.506 -0.534 -1.346
H7 2.652 0.359 -0.297
H8 1.930 -1.175 0.269
H9 0.812 0.835 1.346
H10 0.361 1.455 -0.263
H11 -1.961 0.948 -0.864
H12 -2.754 -0.627 -0.232

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.54442.53263.61272.86931.10511.10501.10602.17572.18663.92084.5010
C21.54441.51382.54282.23322.19212.19922.19561.10841.10702.80203.5402
C32.53261.51381.35541.10172.80433.49432.78852.15202.15122.14052.1427
C43.61272.54281.35542.11993.58524.53754.02453.24342.65401.09801.0959
H52.86932.23321.10172.11993.21873.88322.67972.58843.12143.10742.4609
H61.10512.19212.80433.58523.21871.79201.78823.09872.53793.80124.4047
H71.10502.19923.49434.53753.88321.79201.78822.51282.54034.68505.4965
H81.10602.19562.78854.02452.67971.78821.78822.54053.10894.57534.7432
H92.17571.10842.15203.24342.58843.09872.51282.54051.78203.54724.1650
H102.18661.10702.15122.65403.12142.53792.54033.10891.78202.45103.7473
H113.92082.80202.14051.09803.10743.80124.68504.57533.54722.45101.8740
H124.50103.54022.14271.09592.46094.40475.49654.74324.16503.74731.8740

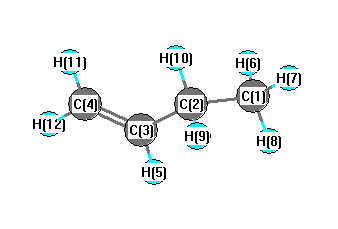
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.815 C1 C2 H9 109.097
C1 C2 H10 110.022 C2 C1 H6 110.567
C2 C1 H7 111.128 C2 C1 H8 110.788
C2 C3 C4 124.721 C2 C3 H5 116.378
C3 C2 H9 109.331 C3 C2 H10 109.355
C3 C4 H11 121.136 C3 C4 H12 121.520
C4 C3 H5 118.894 H6 C1 H7 108.352
H6 C1 H8 107.942 H7 C1 H8 107.948
H9 C2 H10 107.102 H11 C4 H12 117.344
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability