Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -155.967121 |
| Energy at 298.15K | -155.972530 |
| Nuclear repulsion energy | 103.571402 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3269 |
3133 |
0.00 |
|
|
|
| 2 |
Ag |
3175 |
3043 |
0.00 |
|
|
|
| 3 |
Ag |
3165 |
3033 |
0.00 |
|
|
|
| 4 |
Ag |
1729 |
1657 |
0.00 |
|
|
|
| 5 |
Ag |
1465 |
1404 |
0.00 |
|
|
|
| 6 |
Ag |
1311 |
1257 |
0.00 |
|
|
|
| 7 |
Ag |
1231 |
1179 |
0.00 |
|
|
|
| 8 |
Ag |
907 |
869 |
0.00 |
|
|
|
| 9 |
Ag |
516 |
495 |
0.00 |
|
|
|
| 10 |
Au |
1055 |
1011 |
36.08 |
|
|
|
| 11 |
Au |
953 |
913 |
84.66 |
|
|
|
| 12 |
Au |
541 |
519 |
13.10 |
|
|
|
| 13 |
Au |
181 |
174 |
0.72 |
|
|
|
| 14 |
Bg |
1004 |
962 |
0.00 |
|
|
|
| 15 |
Bg |
951 |
912 |
0.00 |
|
|
|
| 16 |
Bg |
785 |
752 |
0.00 |
|
|
|
| 17 |
Bu |
3269 |
3133 |
20.34 |
|
|
|
| 18 |
Bu |
3182 |
3049 |
12.38 |
|
|
|
| 19 |
Bu |
3168 |
3036 |
14.75 |
|
|
|
| 20 |
Bu |
1667 |
1597 |
23.19 |
|
|
|
| 21 |
Bu |
1407 |
1349 |
5.87 |
|
|
|
| 22 |
Bu |
1311 |
1256 |
2.18 |
|
|
|
| 23 |
Bu |
993 |
952 |
2.01 |
|
|
|
| 24 |
Bu |
296 |
283 |
2.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18764.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 17982.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.327 |
0.650 |
0.000 |
| C2 |
0.327 |
-0.650 |
0.000 |
| C3 |
0.327 |
1.820 |
0.000 |
| C4 |
-0.327 |
-1.820 |
0.000 |
| H5 |
-1.420 |
0.646 |
0.000 |
| H6 |
1.420 |
-0.646 |
0.000 |
| H7 |
-0.205 |
2.770 |
0.000 |
| H8 |
1.417 |
1.861 |
0.000 |
| H9 |
0.205 |
-2.770 |
0.000 |
| H10 |
-1.417 |
-1.861 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4546 | 1.3406 | 2.4700 | 1.0933 | 2.1749 | 2.1235 | 2.1229 | 3.4607 | 2.7372 |
C2 | 1.4546 | | 2.4700 | 1.3406 | 2.1749 | 1.0933 | 3.4607 | 2.7372 | 2.1235 | 2.1229 | C3 | 1.3406 | 2.4700 | | 3.6988 | 2.1050 | 2.6976 | 1.0885 | 1.0906 | 4.5917 | 4.0733 | C4 | 2.4700 | 1.3406 | 3.6988 | | 2.6976 | 2.1050 | 4.5917 | 4.0733 | 1.0885 | 1.0906 | H5 | 1.0933 | 2.1749 | 2.1050 | 2.6976 | | 3.1200 | 2.4468 | 3.0860 | 3.7827 | 2.5069 | H6 | 2.1749 | 1.0933 | 2.6976 | 2.1050 | 3.1200 | | 3.7827 | 2.5069 | 2.4468 | 3.0860 | H7 | 2.1235 | 3.4607 | 1.0885 | 4.5917 | 2.4468 | 3.7827 | | 1.8592 | 5.5548 | 4.7867 | H8 | 2.1229 | 2.7372 | 1.0906 | 4.0733 | 3.0860 | 2.5069 | 1.8592 | | 4.7867 | 4.6778 | H9 | 3.4607 | 2.1235 | 4.5917 | 1.0885 | 3.7827 | 2.4468 | 5.5548 | 4.7867 | | 1.8592 | H10 | 2.7372 | 2.1229 | 4.0733 | 1.0906 | 2.5069 | 3.0860 | 4.7867 | 4.6778 | 1.8592 | |
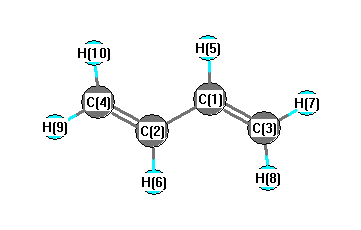 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
124.120 |
|
C1 |
C2 |
H6 |
116.495 |
| C1 |
C3 |
H7 |
121.551 |
|
C1 |
C3 |
H8 |
121.324 |
| C2 |
C1 |
C3 |
124.120 |
|
C2 |
C1 |
H5 |
116.495 |
| C2 |
C4 |
H9 |
121.551 |
|
C2 |
C4 |
H10 |
121.324 |
| C3 |
C1 |
H5 |
119.385 |
|
C4 |
C2 |
H6 |
119.385 |
| H7 |
C3 |
H8 |
117.125 |
|
H9 |
C4 |
H10 |
117.125 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.216 |
|
|
|
| 2 |
C |
1.216 |
|
|
|
| 3 |
C |
0.285 |
|
|
|
| 4 |
C |
0.285 |
|
|
|
| 5 |
H |
-0.646 |
|
|
|
| 6 |
H |
-0.646 |
|
|
|
| 7 |
H |
-0.411 |
|
|
|
| 8 |
H |
-0.445 |
|
|
|
| 9 |
H |
-0.411 |
|
|
|
| 10 |
H |
-0.445 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.849 |
0.027 |
0.000 |
| y |
0.027 |
-23.213 |
0.000 |
| z |
0.000 |
0.000 |
-29.065 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.291 |
0.027 |
0.000 |
| y |
0.027 |
2.744 |
0.000 |
| z |
0.000 |
0.000 |
-6.034 |
|
| Polar |
|---|
| 3z2-r2 | -12.069 |
|---|
| x2-y2 | 0.365 |
|---|
| xy | 0.027 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.117 |
1.518 |
0.000 |
| y |
1.518 |
12.759 |
0.000 |
| z |
0.000 |
0.000 |
5.469 |
<r2> (average value of r
2) Å
2
| <r2> |
94.268 |
| (<r2>)1/2 |
9.709 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -155.961482 |
| Energy at 298.15K | -155.966830 |
| HF Energy | -155.961482 |
| Nuclear repulsion energy | 104.666929 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3270 |
3133 |
4.71 |
|
|
|
| 2 |
A |
3184 |
3051 |
7.64 |
|
|
|
| 3 |
A |
3169 |
3037 |
16.75 |
|
|
|
| 4 |
A |
1702 |
1631 |
1.95 |
|
|
|
| 5 |
A |
1458 |
1397 |
11.86 |
|
|
|
| 6 |
A |
1338 |
1283 |
0.04 |
|
|
|
| 7 |
A |
1069 |
1024 |
0.06 |
|
|
|
| 8 |
A |
1011 |
969 |
2.35 |
|
|
|
| 9 |
A |
955 |
915 |
7.64 |
|
|
|
| 10 |
A |
896 |
858 |
0.79 |
|
|
|
| 11 |
A |
758 |
726 |
3.62 |
|
|
|
| 12 |
A |
270 |
259 |
0.00 |
|
|
|
| 13 |
A |
165 |
158 |
0.26 |
|
|
|
| 14 |
B |
3268 |
3132 |
14.36 |
|
|
|
| 15 |
B |
3173 |
3040 |
8.56 |
|
|
|
| 16 |
B |
3165 |
3033 |
4.02 |
|
|
|
| 17 |
B |
1709 |
1638 |
7.56 |
|
|
|
| 18 |
B |
1426 |
1366 |
2.60 |
|
|
|
| 19 |
B |
1307 |
1253 |
0.37 |
|
|
|
| 20 |
B |
1099 |
1053 |
4.70 |
|
|
|
| 21 |
B |
1026 |
984 |
29.65 |
|
|
|
| 22 |
B |
956 |
916 |
77.11 |
|
|
|
| 23 |
B |
618 |
592 |
9.79 |
|
|
|
| 24 |
B |
468 |
449 |
15.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18729.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 17948.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.103 |
0.726 |
0.551 |
| C2 |
-0.103 |
-0.726 |
0.551 |
| C3 |
-0.103 |
1.540 |
-0.491 |
| C4 |
0.103 |
-1.540 |
-0.491 |
| H5 |
0.431 |
1.162 |
1.498 |
| H6 |
-0.431 |
-1.162 |
1.498 |
| H7 |
0.089 |
2.609 |
-0.418 |
| H8 |
-0.485 |
1.167 |
-1.441 |
| H9 |
-0.089 |
-2.609 |
-0.418 |
| H10 |
0.485 |
-1.167 |
-1.441 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4662 | 1.3388 | 2.4943 | 1.0933 | 2.1789 | 2.1182 | 2.1240 | 3.4784 | 2.7748 |
C2 | 1.4662 | | 2.4943 | 1.3388 | 2.1789 | 1.0933 | 3.4784 | 2.7748 | 2.1182 | 2.1240 | C3 | 1.3388 | 2.4943 | | 3.0872 | 2.0944 | 3.3720 | 1.0887 | 1.0900 | 4.1502 | 2.9287 | C4 | 2.4943 | 1.3388 | 3.0872 | | 3.3720 | 2.0944 | 4.1502 | 2.9287 | 1.0887 | 1.0900 | H5 | 1.0933 | 2.1789 | 2.0944 | 3.3720 | | 2.4794 | 2.4254 | 3.0793 | 4.2624 | 3.7512 | H6 | 2.1789 | 1.0933 | 3.3720 | 2.0944 | 2.4794 | | 4.2624 | 3.7512 | 2.4254 | 3.0793 | H7 | 2.1182 | 3.4784 | 1.0887 | 4.1502 | 2.4254 | 4.2624 | | 1.8595 | 5.2217 | 3.9326 | H8 | 2.1240 | 2.7748 | 1.0900 | 2.9287 | 3.0793 | 3.7512 | 1.8595 | | 3.9326 | 2.5277 | H9 | 3.4784 | 2.1182 | 4.1502 | 1.0887 | 4.2624 | 2.4254 | 5.2217 | 3.9326 | | 1.8595 | H10 | 2.7748 | 2.1240 | 2.9287 | 1.0900 | 3.7512 | 3.0793 | 3.9326 | 2.5277 | 1.8595 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
125.493 |
|
C1 |
C2 |
H6 |
115.945 |
| C1 |
C3 |
H7 |
121.179 |
|
C1 |
C3 |
H8 |
121.631 |
| C2 |
C1 |
C3 |
125.493 |
|
C2 |
C1 |
H5 |
115.945 |
| C2 |
C4 |
H9 |
121.179 |
|
C2 |
C4 |
H10 |
121.631 |
| C3 |
C1 |
H5 |
118.547 |
|
C4 |
C2 |
H6 |
118.547 |
| H7 |
C3 |
H8 |
117.181 |
|
H9 |
C4 |
H10 |
117.181 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.925 |
|
|
|
| 2 |
C |
0.925 |
|
|
|
| 3 |
C |
0.396 |
|
|
|
| 4 |
C |
0.396 |
|
|
|
| 5 |
H |
-0.501 |
|
|
|
| 6 |
H |
-0.501 |
|
|
|
| 7 |
H |
-0.385 |
|
|
|
| 8 |
H |
-0.434 |
|
|
|
| 9 |
H |
-0.385 |
|
|
|
| 10 |
H |
-0.434 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.100 |
0.100 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.322 |
0.731 |
0.000 |
| y |
0.731 |
-23.455 |
0.000 |
| z |
0.000 |
0.000 |
-23.215 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.987 |
0.731 |
0.000 |
| y |
0.731 |
2.314 |
0.000 |
| z |
0.000 |
0.000 |
2.673 |
|
| Polar |
|---|
| 3z2-r2 | 5.346 |
|---|
| x2-y2 | -4.867 |
|---|
| xy | 0.731 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.594 |
-0.145 |
0.000 |
| y |
-0.145 |
10.321 |
0.000 |
| z |
0.000 |
0.000 |
7.796 |
<r2> (average value of r
2) Å
2
| <r2> |
86.093 |
| (<r2>)1/2 |
9.279 |
