return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHCCH2CH3 (1-Butyne)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-155.540744
Energy at 298.15K-155.545416
HF Energy-154.912629
Nuclear repulsion energy101.503415
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3481 3400        
2 A' 3149 3076        
3 A' 3067 2996        
4 A' 3052 2982        
5 A' 2153 2104        
6 A' 1549 1513        
7 A' 1528 1493        
8 A' 1455 1421        
9 A' 1382 1350        
10 A' 1114 1089        
11 A' 1049 1025        
12 A' 859 839        
13 A' 539 526        
14 A' 439 429        
15 A' 165 161        
16 A" 3158 3085        
17 A" 3092 3020        
18 A" 1541 1505        
19 A" 1313 1283        
20 A" 1131 1105        
21 A" 801 782        
22 A" 499 487        
23 A" 309 302        
24 A" 198 194        

Unscaled Zero Point Vibrational Energy (zpe) 18510.8 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 18083.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.88174 0.14808 0.13320

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.672 -1.887 0.000
C2 0.000 -0.850 0.000
C3 0.765 0.420 0.000
C4 -0.163 1.655 0.000
H5 -1.242 -2.801 0.000
H6 1.423 0.440 0.889
H7 1.423 0.440 -0.889
H8 0.437 2.583 0.000
H9 -0.810 1.654 -0.894
H10 -0.810 1.654 0.894

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.23592.71783.57911.07673.25463.25464.60593.65543.6554
C21.23591.48222.51062.31252.11612.11613.46072.77972.7797
C32.71781.48221.54493.79451.10611.10612.18802.19162.1916
C43.57912.51061.54494.58492.18672.18671.10461.10391.1039
H51.07672.31253.79454.58494.28854.28855.63954.56434.5643
H63.25462.11611.10612.18674.28851.77752.52113.10472.5417
H73.25462.11611.10612.18674.28851.77752.52112.54173.1047
H84.60593.46072.18801.10465.63952.52112.52111.79361.7936
H93.65542.77972.19161.10394.56433.10472.54171.79361.7885
H103.65542.77972.19161.10394.56432.54173.10471.79361.7885

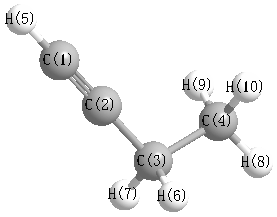
picture of 1-Butyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 178.135 C2 C1 H5 179.050
C2 C3 C4 112.050 C2 C3 H6 108.811
C2 C3 H7 108.811 C3 C4 H8 110.234
C3 C4 H9 110.556 C3 C4 H10 110.556
C4 C3 H6 110.044 C4 C3 H7 110.044
H6 C3 H7 106.933 H8 C4 H9 108.608
H8 C4 H10 108.608 H9 C4 H10 108.212
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability