Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.886503 |
| Energy at 298.15K | |
| HF Energy | -191.886503 |
| Nuclear repulsion energy | 103.002421 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3270 |
3133 |
3.95 |
65.14 |
0.53 |
0.70 |
| 2 |
A' |
3215 |
3081 |
1.17 |
101.93 |
0.28 |
0.43 |
| 3 |
A' |
3166 |
3034 |
4.72 |
86.34 |
0.08 |
0.15 |
| 4 |
A' |
2925 |
2803 |
84.28 |
109.40 |
0.27 |
0.42 |
| 5 |
A' |
1800 |
1725 |
275.56 |
89.53 |
0.38 |
0.55 |
| 6 |
A' |
1697 |
1626 |
2.13 |
50.60 |
0.12 |
0.21 |
| 7 |
A' |
1446 |
1385 |
13.56 |
6.13 |
0.50 |
0.67 |
| 8 |
A' |
1377 |
1320 |
6.56 |
20.60 |
0.29 |
0.45 |
| 9 |
A' |
1292 |
1238 |
3.87 |
9.20 |
0.15 |
0.25 |
| 10 |
A' |
1178 |
1129 |
37.42 |
11.57 |
0.64 |
0.78 |
| 11 |
A' |
924 |
885 |
17.57 |
1.68 |
0.00 |
0.00 |
| 12 |
A' |
573 |
549 |
5.37 |
6.23 |
0.17 |
0.30 |
| 13 |
A' |
318 |
305 |
10.49 |
1.27 |
0.33 |
0.49 |
| 14 |
A" |
1035 |
991 |
4.91 |
1.33 |
0.75 |
0.86 |
| 15 |
A" |
1027 |
984 |
6.34 |
1.85 |
0.75 |
0.86 |
| 16 |
A" |
1010 |
968 |
47.43 |
2.07 |
0.75 |
0.86 |
| 17 |
A" |
614 |
589 |
10.50 |
0.67 |
0.75 |
0.86 |
| 18 |
A" |
169 |
162 |
4.52 |
0.51 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13517.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12953.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.744 |
0.000 |
| C2 |
0.000 |
0.720 |
0.000 |
| C3 |
1.215 |
1.280 |
0.000 |
| O4 |
-1.218 |
-1.319 |
0.000 |
| H5 |
0.806 |
-1.316 |
0.000 |
| H6 |
-0.917 |
1.311 |
0.000 |
| H7 |
1.356 |
2.360 |
0.000 |
| H8 |
2.115 |
0.662 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4710 | 2.4417 | 1.2114 | 1.1157 | 2.1922 | 3.4498 | 2.6663 |
C2 | 1.4710 | | 1.3380 | 2.3742 | 2.1892 | 1.0907 | 2.1277 | 2.1153 | C3 | 2.4417 | 1.3380 | | 3.5595 | 2.6275 | 2.1319 | 1.0889 | 1.0914 | O4 | 1.2114 | 2.3742 | 3.5595 | | 2.0241 | 2.6466 | 4.4890 | 3.8762 | H5 | 1.1157 | 2.1892 | 2.6275 | 2.0241 | | 3.1411 | 3.7161 | 2.3708 | H6 | 2.1922 | 1.0907 | 2.1319 | 2.6466 | 3.1411 | | 2.5025 | 3.0998 | H7 | 3.4498 | 2.1277 | 1.0889 | 4.4890 | 3.7161 | 2.5025 | | 1.8599 | H8 | 2.6663 | 2.1153 | 1.0914 | 3.8762 | 2.3708 | 3.0998 | 1.8599 | |
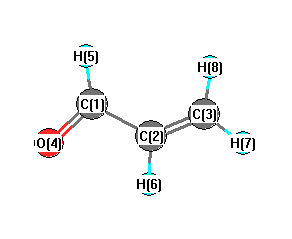 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.665 |
|
C1 |
C2 |
H6 |
116.911 |
| C2 |
C1 |
O4 |
124.248 |
|
C2 |
C1 |
H5 |
114.936 |
| C2 |
C3 |
H7 |
122.168 |
|
C2 |
C3 |
H8 |
120.745 |
| C3 |
C2 |
H6 |
122.425 |
|
O4 |
C1 |
H5 |
120.816 |
| H7 |
C3 |
H8 |
117.088 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.599 |
|
|
|
| 2 |
C |
1.032 |
|
|
|
| 3 |
C |
0.362 |
|
|
|
| 4 |
O |
-0.528 |
|
|
|
| 5 |
H |
-0.231 |
|
|
|
| 6 |
H |
-0.492 |
|
|
|
| 7 |
H |
-0.347 |
|
|
|
| 8 |
H |
-0.395 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.619 |
2.214 |
0.000 |
3.429 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.720 |
-2.786 |
0.000 |
| y |
-2.786 |
-24.019 |
0.000 |
| z |
0.000 |
0.000 |
-25.052 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.184 |
-2.786 |
0.000 |
| y |
-2.786 |
0.867 |
0.000 |
| z |
0.000 |
0.000 |
-0.683 |
|
| Polar |
|---|
| 3z2-r2 | -1.366 |
|---|
| x2-y2 | -0.701 |
|---|
| xy | -2.786 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.748 |
1.783 |
0.000 |
| y |
1.783 |
7.274 |
0.000 |
| z |
0.000 |
0.000 |
4.160 |
<r2> (average value of r
2) Å
2
| <r2> |
83.637 |
| (<r2>)1/2 |
9.145 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.882863 |
| Energy at 298.15K | |
| HF Energy | -191.882863 |
| Nuclear repulsion energy | 104.510679 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3281 |
3145 |
1.25 |
54.81 |
0.62 |
0.77 |
| 2 |
A' |
3203 |
3069 |
9.95 |
126.99 |
0.22 |
0.36 |
| 3 |
A' |
3174 |
3042 |
4.94 |
68.61 |
0.10 |
0.18 |
| 4 |
A' |
2948 |
2825 |
129.15 |
175.77 |
0.28 |
0.44 |
| 5 |
A' |
1802 |
1726 |
129.11 |
20.78 |
0.61 |
0.76 |
| 6 |
A' |
1690 |
1620 |
72.03 |
73.40 |
0.13 |
0.23 |
| 7 |
A' |
1430 |
1370 |
41.49 |
5.33 |
0.75 |
0.86 |
| 8 |
A' |
1415 |
1356 |
4.75 |
9.79 |
0.10 |
0.18 |
| 9 |
A' |
1309 |
1254 |
2.53 |
20.52 |
0.27 |
0.43 |
| 10 |
A' |
1065 |
1021 |
3.64 |
3.00 |
0.72 |
0.84 |
| 11 |
A' |
949 |
909 |
60.91 |
7.13 |
0.08 |
0.15 |
| 12 |
A' |
682 |
653 |
13.39 |
0.70 |
0.75 |
0.86 |
| 13 |
A' |
281 |
269 |
7.07 |
3.50 |
0.33 |
0.50 |
| 14 |
A" |
1040 |
997 |
0.01 |
4.82 |
0.75 |
0.86 |
| 15 |
A" |
1028 |
985 |
48.30 |
0.46 |
0.75 |
0.86 |
| 16 |
A" |
1016 |
973 |
2.54 |
0.40 |
0.75 |
0.86 |
| 17 |
A" |
558 |
535 |
10.53 |
0.83 |
0.75 |
0.86 |
| 18 |
A" |
152 |
146 |
6.49 |
0.21 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13509.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12946.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.888 |
-0.291 |
0.000 |
| C2 |
0.000 |
0.895 |
0.000 |
| C3 |
1.332 |
0.776 |
0.000 |
| O4 |
-0.498 |
-1.439 |
0.000 |
| H5 |
-1.979 |
-0.072 |
0.000 |
| H6 |
-0.482 |
1.874 |
0.000 |
| H7 |
1.985 |
1.647 |
0.000 |
| H8 |
1.792 |
-0.213 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4813 | 2.4629 | 1.2120 | 1.1135 | 2.2025 | 3.4652 | 2.6806 |
C2 | 1.4813 | | 1.3375 | 2.3860 | 2.2027 | 1.0913 | 2.1226 | 2.1064 | C3 | 2.4629 | 1.3375 | | 2.8726 | 3.4182 | 2.1208 | 1.0888 | 1.0900 | O4 | 1.2120 | 2.3860 | 2.8726 | | 2.0158 | 3.3124 | 3.9605 | 2.5971 | H5 | 1.1135 | 2.2027 | 3.4182 | 2.0158 | | 2.4548 | 4.3207 | 3.7738 | H6 | 2.2025 | 1.0913 | 2.1208 | 3.3124 | 2.4548 | | 2.4774 | 3.0862 | H7 | 3.4652 | 2.1226 | 1.0888 | 3.9605 | 4.3207 | 2.4774 | | 1.8699 | H8 | 2.6806 | 2.1064 | 1.0900 | 2.5971 | 3.7738 | 3.0862 | 1.8699 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.708 |
|
C1 |
C2 |
H6 |
116.955 |
| C2 |
C1 |
O4 |
124.416 |
|
C2 |
C1 |
H5 |
115.455 |
| C2 |
C3 |
H7 |
121.718 |
|
C2 |
C3 |
H8 |
120.050 |
| C3 |
C2 |
H6 |
121.337 |
|
O4 |
C1 |
H5 |
120.129 |
| H7 |
C3 |
H8 |
118.233 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.589 |
|
|
|
| 2 |
C |
0.932 |
|
|
|
| 3 |
C |
0.394 |
|
|
|
| 4 |
O |
-0.526 |
|
|
|
| 5 |
H |
-0.181 |
|
|
|
| 6 |
H |
-0.517 |
|
|
|
| 7 |
H |
-0.345 |
|
|
|
| 8 |
H |
-0.346 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.253 |
2.803 |
0.000 |
2.815 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.185 |
-0.672 |
0.000 |
| y |
-0.672 |
-26.500 |
0.000 |
| z |
0.000 |
0.000 |
-25.020 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.575 |
-0.672 |
0.000 |
| y |
-0.672 |
-3.897 |
0.000 |
| z |
0.000 |
0.000 |
-1.677 |
|
| Polar |
|---|
| 3z2-r2 | -3.355 |
|---|
| x2-y2 | 6.315 |
|---|
| xy | -0.672 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.859 |
0.627 |
0.000 |
| y |
0.627 |
6.632 |
0.000 |
| z |
0.000 |
0.000 |
4.149 |
<r2> (average value of r
2) Å
2
| <r2> |
75.342 |
| (<r2>)1/2 |
8.680 |
