Jump to
S1C2
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.824079 |
| Energy at 298.15K | |
| HF Energy | -191.824079 |
| Nuclear repulsion energy | 102.240605 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3188 |
3188 |
8.77 |
74.08 |
0.53 |
0.70 |
| 2 |
A' |
3131 |
3131 |
3.57 |
112.28 |
0.28 |
0.44 |
| 3 |
A' |
3087 |
3087 |
8.92 |
96.06 |
0.08 |
0.15 |
| 4 |
A' |
2806 |
2806 |
113.14 |
128.58 |
0.29 |
0.44 |
| 5 |
A' |
1704 |
1704 |
251.49 |
85.03 |
0.40 |
0.57 |
| 6 |
A' |
1630 |
1630 |
2.03 |
41.81 |
0.11 |
0.20 |
| 7 |
A' |
1415 |
1415 |
9.92 |
6.17 |
0.46 |
0.63 |
| 8 |
A' |
1346 |
1346 |
5.55 |
22.23 |
0.30 |
0.46 |
| 9 |
A' |
1265 |
1265 |
2.82 |
10.84 |
0.14 |
0.25 |
| 10 |
A' |
1144 |
1144 |
37.03 |
10.76 |
0.65 |
0.79 |
| 11 |
A' |
901 |
901 |
18.78 |
1.97 |
0.01 |
0.02 |
| 12 |
A' |
558 |
558 |
4.15 |
7.55 |
0.18 |
0.30 |
| 13 |
A' |
318 |
318 |
9.21 |
1.47 |
0.33 |
0.50 |
| 14 |
A" |
1001 |
1001 |
13.57 |
0.45 |
0.75 |
0.86 |
| 15 |
A" |
988 |
988 |
4.90 |
2.10 |
0.75 |
0.86 |
| 16 |
A" |
972 |
972 |
35.81 |
2.56 |
0.75 |
0.86 |
| 17 |
A" |
594 |
594 |
9.21 |
0.60 |
0.75 |
0.86 |
| 18 |
A" |
164 |
164 |
3.73 |
0.60 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13106.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13106.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.748 |
0.000 |
| C2 |
0.000 |
0.722 |
0.000 |
| C3 |
1.222 |
1.293 |
0.000 |
| O4 |
-1.226 |
-1.332 |
0.000 |
| H5 |
0.817 |
-1.319 |
0.000 |
| H6 |
-0.922 |
1.315 |
0.000 |
| H7 |
1.358 |
2.379 |
0.000 |
| H8 |
2.129 |
0.677 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4779 | 2.4603 | 1.2235 | 1.1240 | 2.2021 | 3.4719 | 2.6887 |
C2 | 1.4779 | | 1.3490 | 2.3923 | 2.1984 | 1.0959 | 2.1420 | 2.1294 | C3 | 2.4603 | 1.3490 | | 3.5898 | 2.6433 | 2.1441 | 1.0937 | 1.0964 | O4 | 1.2235 | 2.3923 | 3.5898 | | 2.0434 | 2.6642 | 4.5219 | 3.9106 | H5 | 1.1240 | 2.1984 | 2.6433 | 2.0434 | | 3.1559 | 3.7367 | 2.3883 | H6 | 2.2021 | 1.0959 | 2.1441 | 2.6642 | 3.1559 | | 2.5160 | 3.1167 | H7 | 3.4719 | 2.1420 | 1.0937 | 4.5219 | 3.7367 | 2.5160 | | 1.8680 | H8 | 2.6887 | 2.1294 | 1.0964 | 3.9106 | 2.3883 | 3.1167 | 1.8680 | |
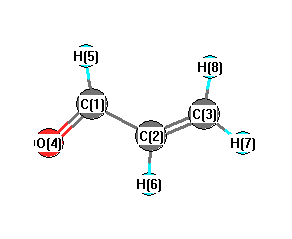 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.919 |
|
C1 |
C2 |
H6 |
116.865 |
| C2 |
C1 |
O4 |
124.376 |
|
C2 |
C1 |
H5 |
114.648 |
| C2 |
C3 |
H7 |
122.196 |
|
C2 |
C3 |
H8 |
120.747 |
| C3 |
C2 |
H6 |
122.215 |
|
O4 |
C1 |
H5 |
120.977 |
| H7 |
C3 |
H8 |
117.057 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.641 |
|
|
|
| 2 |
C |
1.195 |
|
|
|
| 3 |
C |
0.513 |
|
|
|
| 4 |
O |
-0.502 |
|
|
|
| 5 |
H |
-0.314 |
|
|
|
| 6 |
H |
-0.591 |
|
|
|
| 7 |
H |
-0.441 |
|
|
|
| 8 |
H |
-0.501 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.530 |
2.243 |
0.000 |
3.382 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.004 |
-2.667 |
0.000 |
| y |
-2.667 |
-24.286 |
0.000 |
| z |
0.000 |
0.000 |
-25.052 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.335 |
-2.667 |
0.000 |
| y |
-2.667 |
0.742 |
0.000 |
| z |
0.000 |
0.000 |
-0.406 |
|
| Polar |
|---|
| 3z2-r2 | -0.813 |
|---|
| x2-y2 | -0.718 |
|---|
| xy | -2.667 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.119 |
1.845 |
0.000 |
| y |
1.845 |
7.764 |
0.000 |
| z |
0.000 |
0.000 |
4.308 |
<r2> (average value of r
2) Å
2
| <r2> |
84.800 |
| (<r2>)1/2 |
9.209 |
Jump to
S1C1
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.820481 |
| Energy at 298.15K | |
| HF Energy | -191.820481 |
| Nuclear repulsion energy | 103.618489 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3201 |
3201 |
4.24 |
60.30 |
0.64 |
0.78 |
| 2 |
A' |
3115 |
3115 |
15.53 |
159.22 |
0.19 |
0.32 |
| 3 |
A' |
3098 |
3098 |
10.02 |
60.93 |
0.14 |
0.25 |
| 4 |
A' |
2826 |
2826 |
167.65 |
207.74 |
0.30 |
0.46 |
| 5 |
A' |
1714 |
1714 |
103.05 |
17.47 |
0.71 |
0.83 |
| 6 |
A' |
1617 |
1617 |
85.31 |
72.41 |
0.16 |
0.27 |
| 7 |
A' |
1399 |
1399 |
37.78 |
6.68 |
0.59 |
0.74 |
| 8 |
A' |
1383 |
1383 |
1.03 |
10.34 |
0.10 |
0.18 |
| 9 |
A' |
1281 |
1281 |
2.96 |
22.01 |
0.27 |
0.42 |
| 10 |
A' |
1044 |
1044 |
3.49 |
3.16 |
0.71 |
0.83 |
| 11 |
A' |
914 |
914 |
59.28 |
8.67 |
0.09 |
0.16 |
| 12 |
A' |
665 |
665 |
10.14 |
0.66 |
0.74 |
0.85 |
| 13 |
A' |
279 |
279 |
6.52 |
3.81 |
0.32 |
0.49 |
| 14 |
A" |
1000 |
1000 |
4.96 |
3.46 |
0.75 |
0.86 |
| 15 |
A" |
992 |
992 |
39.87 |
1.11 |
0.75 |
0.86 |
| 16 |
A" |
980 |
980 |
1.62 |
0.93 |
0.75 |
0.86 |
| 17 |
A" |
533 |
533 |
8.94 |
1.27 |
0.75 |
0.86 |
| 18 |
A" |
155 |
155 |
5.55 |
0.47 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13097.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13097.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.891 |
-0.296 |
0.000 |
| C2 |
0.000 |
0.896 |
0.000 |
| C3 |
1.345 |
0.794 |
0.000 |
| O4 |
-0.506 |
-1.458 |
0.000 |
| H5 |
-1.989 |
-0.064 |
0.000 |
| H6 |
-0.493 |
1.875 |
0.000 |
| H7 |
1.987 |
1.679 |
0.000 |
| H8 |
1.825 |
-0.190 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4884 | 2.4879 | 1.2240 | 1.1219 | 2.2078 | 3.4913 | 2.7182 |
C2 | 1.4884 | | 1.3486 | 2.4074 | 2.2084 | 1.0969 | 2.1361 | 2.1230 | C3 | 2.4879 | 1.3486 | | 2.9152 | 3.4424 | 2.1324 | 1.0937 | 1.0946 | O4 | 1.2240 | 2.4074 | 2.9152 | | 2.0352 | 3.3333 | 4.0074 | 2.6536 | H5 | 1.1219 | 2.2084 | 3.4424 | 2.0352 | | 2.4491 | 4.3416 | 3.8157 | H6 | 2.2078 | 1.0969 | 2.1324 | 3.3333 | 2.4491 | | 2.4882 | 3.1043 | H7 | 3.4913 | 2.1361 | 1.0937 | 4.0074 | 4.3416 | 2.4882 | | 1.8759 | H8 | 2.7182 | 2.1230 | 1.0946 | 2.6536 | 3.8157 | 3.1043 | 1.8759 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
122.481 |
|
C1 |
C2 |
H6 |
116.481 |
| C2 |
C1 |
O4 |
124.857 |
|
C2 |
C1 |
H5 |
114.844 |
| C2 |
C3 |
H7 |
121.663 |
|
C2 |
C3 |
H8 |
120.317 |
| C3 |
C2 |
H6 |
121.038 |
|
O4 |
C1 |
H5 |
120.299 |
| H7 |
C3 |
H8 |
118.020 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.642 |
|
|
|
| 2 |
C |
1.091 |
|
|
|
| 3 |
C |
0.547 |
|
|
|
| 4 |
O |
-0.505 |
|
|
|
| 5 |
H |
-0.266 |
|
|
|
| 6 |
H |
-0.623 |
|
|
|
| 7 |
H |
-0.441 |
|
|
|
| 8 |
H |
-0.445 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.357 |
2.771 |
0.000 |
2.794 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.687 |
-0.618 |
0.000 |
| y |
-0.618 |
-26.690 |
0.000 |
| z |
0.000 |
0.000 |
-25.020 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.168 |
-0.618 |
0.000 |
| y |
-0.618 |
-3.837 |
0.000 |
| z |
0.000 |
0.000 |
-1.332 |
|
| Polar |
|---|
| 3z2-r2 | -2.664 |
|---|
| x2-y2 | 6.003 |
|---|
| xy | -0.618 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.324 |
0.718 |
0.000 |
| y |
0.718 |
7.084 |
0.000 |
| z |
0.000 |
0.000 |
4.300 |
<r2> (average value of r
2) Å
2
| <r2> |
76.725 |
| (<r2>)1/2 |
8.759 |
