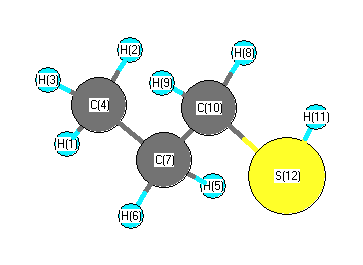
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3142 |
3011 |
29.62 |
|
|
|
| 2 |
A' |
3079 |
2950 |
37.29 |
|
|
|
| 3 |
A' |
3062 |
2934 |
7.42 |
|
|
|
| 4 |
A' |
3054 |
2926 |
24.80 |
|
|
|
| 5 |
A' |
2714 |
2601 |
3.69 |
|
|
|
| 6 |
A' |
1486 |
1424 |
6.10 |
|
|
|
| 7 |
A' |
1472 |
1410 |
1.15 |
|
|
|
| 8 |
A' |
1465 |
1404 |
3.23 |
|
|
|
| 9 |
A' |
1396 |
1338 |
1.78 |
|
|
|
| 10 |
A' |
1364 |
1307 |
3.68 |
|
|
|
| 11 |
A' |
1251 |
1199 |
22.53 |
|
|
|
| 12 |
A' |
1138 |
1090 |
3.20 |
|
|
|
| 13 |
A' |
1072 |
1027 |
0.03 |
|
|
|
| 14 |
A' |
935 |
896 |
2.49 |
|
|
|
| 15 |
A' |
856 |
821 |
2.72 |
|
|
|
| 16 |
A' |
745 |
714 |
2.31 |
|
|
|
| 17 |
A' |
355 |
340 |
0.43 |
|
|
|
| 18 |
A' |
222 |
213 |
1.97 |
|
|
|
| 19 |
A" |
3145 |
3013 |
37.59 |
|
|
|
| 20 |
A" |
3131 |
3000 |
13.40 |
|
|
|
| 21 |
A" |
3100 |
2971 |
1.32 |
|
|
|
| 22 |
A" |
1475 |
1414 |
8.32 |
|
|
|
| 23 |
A" |
1314 |
1259 |
0.07 |
|
|
|
| 24 |
A" |
1233 |
1181 |
0.42 |
|
|
|
| 25 |
A" |
1056 |
1012 |
2.22 |
|
|
|
| 26 |
A" |
862 |
826 |
0.00 |
|
|
|
| 27 |
A" |
743 |
712 |
3.29 |
|
|
|
| 28 |
A" |
230 |
221 |
0.26 |
|
|
|
| 29 |
A" |
181 |
174 |
13.18 |
|
|
|
| 30 |
A" |
101 |
97 |
3.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22688.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 21741.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.137 |
|
|
|
| 2 |
H |
-0.116 |
|
|
|
| 3 |
H |
-0.116 |
|
|
|
| 4 |
C |
0.348 |
|
|
|
| 5 |
H |
-0.226 |
|
|
|
| 6 |
H |
-0.226 |
|
|
|
| 7 |
C |
0.524 |
|
|
|
| 8 |
H |
-0.241 |
|
|
|
| 9 |
H |
-0.241 |
|
|
|
| 10 |
C |
0.179 |
|
|
|
| 11 |
H |
0.100 |
|
|
|
| 12 |
S |
0.152 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.403 |
1.006 |
0.000 |
1.726 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.398 |
-2.601 |
0.000 |
| y |
-2.601 |
-32.763 |
0.000 |
| z |
0.000 |
0.000 |
-35.563 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.236 |
-2.601 |
0.000 |
| y |
-2.601 |
2.718 |
0.000 |
| z |
0.000 |
0.000 |
-1.482 |
|
| Polar |
|---|
| 3z2-r2 | -2.964 |
|---|
| x2-y2 | -2.635 |
|---|
| xy | -2.601 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.293 |
-0.224 |
0.000 |
| y |
-0.224 |
8.292 |
0.000 |
| z |
0.000 |
0.000 |
7.865 |
<r2> (average value of r
2) Å
2
| <r2> |
157.959 |
| (<r2>)1/2 |
12.568 |