Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3113.164596 |
| Energy at 298.15K | |
| HF Energy | -3113.164596 |
| Nuclear repulsion energy | 288.607070 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3137 |
3006 |
6.57 |
27.96 |
0.03 |
0.06 |
| 2 |
A' |
3126 |
2995 |
2.00 |
142.38 |
0.03 |
0.05 |
| 3 |
A' |
1468 |
1407 |
0.68 |
5.60 |
0.74 |
0.85 |
| 4 |
A' |
1459 |
1399 |
7.85 |
0.45 |
0.74 |
0.85 |
| 5 |
A' |
1302 |
1247 |
0.88 |
21.31 |
0.35 |
0.52 |
| 6 |
A' |
1210 |
1159 |
36.61 |
2.50 |
0.26 |
0.42 |
| 7 |
A' |
1092 |
1046 |
1.14 |
4.87 |
0.70 |
0.82 |
| 8 |
A' |
752 |
721 |
30.17 |
52.81 |
0.24 |
0.39 |
| 9 |
A' |
655 |
627 |
59.13 |
16.30 |
0.22 |
0.36 |
| 10 |
A' |
251 |
241 |
0.78 |
4.75 |
0.22 |
0.36 |
| 11 |
A' |
194 |
186 |
7.25 |
0.21 |
0.20 |
0.33 |
| 12 |
A" |
3219 |
3085 |
0.24 |
5.71 |
0.75 |
0.86 |
| 13 |
A" |
3194 |
3061 |
0.23 |
71.68 |
0.75 |
0.86 |
| 14 |
A" |
1280 |
1226 |
0.02 |
2.39 |
0.75 |
0.86 |
| 15 |
A" |
1116 |
1070 |
1.55 |
0.03 |
0.75 |
0.86 |
| 16 |
A" |
970 |
930 |
0.13 |
1.03 |
0.75 |
0.86 |
| 17 |
A" |
762 |
730 |
2.95 |
0.00 |
0.75 |
0.86 |
| 18 |
A" |
110 |
106 |
5.65 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12648.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12121.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.644 |
0.000 |
| C2 |
1.230 |
-0.228 |
0.000 |
| Br3 |
-1.589 |
-0.495 |
0.000 |
| Cl4 |
2.693 |
0.824 |
0.000 |
| H5 |
-0.060 |
1.265 |
0.896 |
| H6 |
-0.060 |
1.265 |
-0.896 |
| H7 |
1.284 |
-0.853 |
0.895 |
| H8 |
1.284 |
-0.853 |
-0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5083 | 1.9549 | 2.6987 | 1.0923 | 1.0923 | 2.1660 | 2.1660 |
C2 | 1.5083 | | 2.8317 | 1.8015 | 2.1678 | 2.1678 | 1.0930 | 1.0930 | Br3 | 1.9549 | 2.8317 | | 4.4800 | 2.4979 | 2.4979 | 3.0300 | 3.0300 | Cl4 | 2.6987 | 1.8015 | 4.4800 | | 2.9284 | 2.9284 | 2.3662 | 2.3662 | H5 | 1.0923 | 2.1678 | 2.4979 | 2.9284 | | 1.7928 | 2.5089 | 3.0828 | H6 | 1.0923 | 2.1678 | 2.4979 | 2.9284 | 1.7928 | | 3.0828 | 2.5089 | H7 | 2.1660 | 1.0930 | 3.0300 | 2.3662 | 2.5089 | 3.0828 | | 1.7901 | H8 | 2.1660 | 1.0930 | 3.0300 | 2.3662 | 3.0828 | 2.5089 | 1.7901 | |
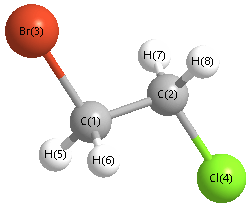 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.926 |
|
C1 |
C2 |
H7 |
111.754 |
| C1 |
C2 |
H8 |
111.754 |
|
C2 |
C1 |
Br3 |
109.022 |
| C2 |
C1 |
Cl4 |
39.157 |
|
C2 |
C1 |
H6 |
111.943 |
| Br3 |
C1 |
H5 |
106.670 |
|
Br3 |
C1 |
H6 |
106.670 |
| Cl4 |
C2 |
H7 |
107.117 |
|
Cl4 |
C2 |
H8 |
107.117 |
| H5 |
C1 |
H6 |
110.304 |
|
H7 |
C2 |
H8 |
109.948 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.012 |
|
|
|
| 2 |
C |
0.097 |
|
|
|
| 3 |
Br |
0.213 |
|
|
|
| 4 |
Cl |
-0.077 |
|
|
|
| 5 |
H |
-0.077 |
|
|
|
| 6 |
H |
-0.077 |
|
|
|
| 7 |
H |
-0.045 |
|
|
|
| 8 |
H |
-0.045 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.061 |
-0.018 |
0.000 |
0.064 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.283 |
-2.310 |
0.000 |
| y |
-2.310 |
-43.512 |
0.000 |
| z |
0.000 |
0.000 |
-43.655 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.700 |
-2.310 |
0.000 |
| y |
-2.310 |
3.457 |
0.000 |
| z |
0.000 |
0.000 |
3.243 |
|
| Polar |
|---|
| 3z2-r2 | 6.485 |
|---|
| x2-y2 | -6.771 |
|---|
| xy | -2.310 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.310 |
1.765 |
0.000 |
| y |
1.765 |
8.436 |
0.000 |
| z |
0.000 |
0.000 |
7.320 |
<r2> (average value of r
2) Å
2
| <r2> |
283.390 |
| (<r2>)1/2 |
16.834 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3113.161675 |
| Energy at 298.15K | |
| HF Energy | -3113.161675 |
| Nuclear repulsion energy | 304.667467 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3195 |
3062 |
0.67 |
37.45 |
0.75 |
0.86 |
| 2 |
A |
3174 |
3042 |
0.75 |
74.15 |
0.75 |
0.86 |
| 3 |
A |
3116 |
2986 |
11.42 |
175.80 |
0.03 |
0.06 |
| 4 |
A |
3099 |
2970 |
9.02 |
110.28 |
0.15 |
0.26 |
| 5 |
A |
1445 |
1385 |
3.34 |
3.70 |
0.75 |
0.86 |
| 6 |
A |
1435 |
1375 |
12.18 |
7.27 |
0.74 |
0.85 |
| 7 |
A |
1319 |
1264 |
17.06 |
1.82 |
0.31 |
0.47 |
| 8 |
A |
1277 |
1224 |
46.87 |
3.65 |
0.38 |
0.55 |
| 9 |
A |
1208 |
1158 |
1.38 |
5.29 |
0.70 |
0.82 |
| 10 |
A |
1136 |
1088 |
0.88 |
1.26 |
0.69 |
0.81 |
| 11 |
A |
1058 |
1014 |
2.39 |
1.13 |
0.69 |
0.82 |
| 12 |
A |
933 |
894 |
8.14 |
3.85 |
0.17 |
0.29 |
| 13 |
A |
867 |
831 |
20.85 |
1.34 |
0.74 |
0.85 |
| 14 |
A |
688 |
660 |
19.40 |
9.19 |
0.26 |
0.41 |
| 15 |
A |
587 |
563 |
11.50 |
17.54 |
0.13 |
0.23 |
| 16 |
A |
389 |
373 |
7.54 |
1.66 |
0.69 |
0.82 |
| 17 |
A |
243 |
233 |
1.28 |
1.19 |
0.23 |
0.37 |
| 18 |
A |
96 |
92 |
0.55 |
0.95 |
0.65 |
0.79 |
Unscaled Zero Point Vibrational Energy (zpe) 12633.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12106.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.010 |
1.100 |
-0.384 |
| C2 |
1.280 |
0.900 |
0.403 |
| Br3 |
-1.355 |
-0.225 |
0.036 |
| Cl4 |
2.169 |
-0.576 |
-0.086 |
| H5 |
-0.426 |
2.073 |
-0.133 |
| H6 |
0.185 |
1.038 |
-1.461 |
| H7 |
1.953 |
1.745 |
0.216 |
| H8 |
1.084 |
0.819 |
1.476 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5069 | 1.9483 | 2.7498 | 1.0950 | 1.0929 | 2.1333 | 2.1653 |
C2 | 1.5069 | | 2.8884 | 1.7915 | 2.1386 | 2.1653 | 1.0965 | 1.0937 | Br3 | 1.9483 | 2.8884 | | 3.5435 | 2.4839 | 2.4912 | 3.8546 | 3.0185 | Cl4 | 2.7498 | 1.7915 | 3.5435 | | 3.7091 | 2.9034 | 2.3512 | 2.3590 | H5 | 1.0950 | 2.1386 | 2.4839 | 3.7091 | | 1.7914 | 2.4273 | 2.5373 | H6 | 1.0929 | 2.1653 | 2.4912 | 2.9034 | 1.7914 | | 2.5372 | 3.0783 | H7 | 2.1333 | 1.0965 | 3.8546 | 2.3512 | 2.4273 | 2.5372 | | 1.7886 | H8 | 2.1653 | 1.0937 | 3.0185 | 2.3590 | 2.5373 | 3.0783 | 1.7886 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.668 |
|
C1 |
C2 |
H7 |
109.041 |
| C1 |
C2 |
H8 |
111.752 |
|
C2 |
C1 |
Br3 |
112.808 |
| C2 |
C1 |
Cl4 |
36.955 |
|
C2 |
C1 |
H6 |
111.804 |
| Br3 |
C1 |
H5 |
105.982 |
|
Br3 |
C1 |
H6 |
106.585 |
| Cl4 |
C2 |
H7 |
106.507 |
|
Cl4 |
C2 |
H8 |
107.204 |
| H5 |
C1 |
H6 |
109.924 |
|
H7 |
C2 |
H8 |
109.502 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.042 |
|
|
|
| 2 |
C |
0.031 |
|
|
|
| 3 |
Br |
0.237 |
|
|
|
| 4 |
Cl |
-0.052 |
|
|
|
| 5 |
H |
-0.071 |
|
|
|
| 6 |
H |
-0.069 |
|
|
|
| 7 |
H |
-0.019 |
|
|
|
| 8 |
H |
-0.016 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.382 |
2.710 |
0.102 |
2.739 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.943 |
2.733 |
0.485 |
| y |
2.733 |
-40.522 |
-0.106 |
| z |
0.485 |
-0.106 |
-43.271 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.046 |
2.733 |
0.485 |
| y |
2.733 |
5.084 |
-0.106 |
| z |
0.485 |
-0.106 |
0.962 |
|
| Polar |
|---|
| 3z2-r2 | 1.923 |
|---|
| x2-y2 | -7.420 |
|---|
| xy | 2.733 |
|---|
| xz | 0.485 |
|---|
| yz | -0.106 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.540 |
0.181 |
-0.165 |
| y |
0.181 |
9.004 |
0.033 |
| z |
-0.165 |
0.033 |
7.517 |
<r2> (average value of r
2) Å
2
| <r2> |
221.736 |
| (<r2>)1/2 |
14.891 |
