Jump to
S1C2
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3114.266671 |
| Energy at 298.15K | |
| HF Energy | -3114.266671 |
| Nuclear repulsion energy | 285.235647 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3066 |
3066 |
8.56 |
32.30 |
0.03 |
0.05 |
| 2 |
A' |
3053 |
3053 |
2.70 |
147.02 |
0.03 |
0.05 |
| 3 |
A' |
1437 |
1437 |
0.55 |
5.21 |
0.75 |
0.86 |
| 4 |
A' |
1429 |
1429 |
4.94 |
0.48 |
0.75 |
0.85 |
| 5 |
A' |
1259 |
1259 |
1.36 |
25.09 |
0.34 |
0.51 |
| 6 |
A' |
1175 |
1175 |
40.24 |
3.62 |
0.26 |
0.41 |
| 7 |
A' |
1049 |
1049 |
0.88 |
6.52 |
0.74 |
0.85 |
| 8 |
A' |
679 |
679 |
30.52 |
80.02 |
0.26 |
0.41 |
| 9 |
A' |
591 |
591 |
75.64 |
18.54 |
0.23 |
0.37 |
| 10 |
A' |
238 |
238 |
0.90 |
6.48 |
0.22 |
0.37 |
| 11 |
A' |
190 |
190 |
7.56 |
0.43 |
0.21 |
0.34 |
| 12 |
A" |
3149 |
3149 |
1.88 |
7.21 |
0.75 |
0.86 |
| 13 |
A" |
3122 |
3122 |
0.55 |
75.20 |
0.75 |
0.86 |
| 14 |
A" |
1249 |
1249 |
0.01 |
2.09 |
0.75 |
0.86 |
| 15 |
A" |
1083 |
1083 |
1.57 |
0.04 |
0.75 |
0.86 |
| 16 |
A" |
943 |
943 |
0.14 |
1.06 |
0.75 |
0.86 |
| 17 |
A" |
751 |
751 |
2.60 |
0.00 |
0.75 |
0.86 |
| 18 |
A" |
108 |
108 |
5.45 |
0.01 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12284.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12284.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.657 |
0.000 |
| C2 |
1.241 |
-0.212 |
0.000 |
| Br3 |
-1.599 |
-0.528 |
0.000 |
| Cl4 |
2.710 |
0.878 |
0.000 |
| H5 |
-0.076 |
1.275 |
0.903 |
| H6 |
-0.076 |
1.275 |
-0.903 |
| H7 |
1.308 |
-0.834 |
0.901 |
| H8 |
1.308 |
-0.834 |
-0.901 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5149 | 1.9907 | 2.7190 | 1.0964 | 1.0964 | 2.1789 | 2.1789 |
C2 | 1.5149 | | 2.8577 | 1.8295 | 2.1816 | 2.1816 | 1.0972 | 1.0972 | Br3 | 1.9907 | 2.8577 | | 4.5331 | 2.5268 | 2.5268 | 3.0591 | 3.0591 | Cl4 | 2.7190 | 1.8295 | 4.5331 | | 2.9556 | 2.9556 | 2.3898 | 2.3898 | H5 | 1.0964 | 2.1816 | 2.5268 | 2.9556 | | 1.8055 | 2.5227 | 3.1012 | H6 | 1.0964 | 2.1816 | 2.5268 | 2.9556 | 1.8055 | | 3.1012 | 2.5227 | H7 | 2.1789 | 1.0972 | 3.0591 | 2.3898 | 2.5227 | 3.1012 | | 1.8020 | H8 | 2.1789 | 1.0972 | 3.0591 | 2.3898 | 3.1012 | 2.5227 | 1.8020 | |
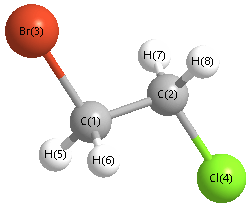 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.417 |
|
C1 |
C2 |
H7 |
112.061 |
| C1 |
C2 |
H8 |
112.061 |
|
C2 |
C1 |
Br3 |
108.451 |
| C2 |
C1 |
Cl4 |
39.671 |
|
C2 |
C1 |
H6 |
112.336 |
| Br3 |
C1 |
H5 |
106.225 |
|
Br3 |
C1 |
H6 |
106.225 |
| Cl4 |
C2 |
H7 |
106.799 |
|
Cl4 |
C2 |
H8 |
106.799 |
| H5 |
C1 |
H6 |
110.858 |
|
H7 |
C2 |
H8 |
110.402 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.245 |
|
|
|
| 2 |
C |
0.307 |
|
|
|
| 3 |
Br |
0.166 |
|
|
|
| 4 |
Cl |
-0.085 |
|
|
|
| 5 |
H |
-0.173 |
|
|
|
| 6 |
H |
-0.173 |
|
|
|
| 7 |
H |
-0.143 |
|
|
|
| 8 |
H |
-0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.049 |
-0.014 |
0.000 |
0.051 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.428 |
-2.484 |
0.000 |
| y |
-2.484 |
-43.690 |
0.000 |
| z |
0.000 |
0.000 |
-43.727 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.719 |
-2.484 |
0.000 |
| y |
-2.484 |
3.387 |
0.000 |
| z |
0.000 |
0.000 |
3.332 |
|
| Polar |
|---|
| 3z2-r2 | 6.663 |
|---|
| x2-y2 | -6.738 |
|---|
| xy | -2.484 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.214 |
1.993 |
0.000 |
| y |
1.993 |
8.863 |
0.000 |
| z |
0.000 |
0.000 |
7.612 |
<r2> (average value of r
2) Å
2
| <r2> |
289.381 |
| (<r2>)1/2 |
17.011 |
Jump to
S1C1
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -3114.263677 |
| Energy at 298.15K | |
| HF Energy | -3114.263677 |
| Nuclear repulsion energy | 300.871027 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3120 |
3120 |
2.40 |
41.40 |
0.75 |
0.86 |
| 2 |
A |
3096 |
3096 |
1.78 |
78.16 |
0.75 |
0.86 |
| 3 |
A |
3039 |
3039 |
14.53 |
188.78 |
0.03 |
0.07 |
| 4 |
A |
3018 |
3018 |
12.21 |
139.67 |
0.15 |
0.27 |
| 5 |
A |
1413 |
1413 |
2.14 |
3.70 |
0.74 |
0.85 |
| 6 |
A |
1403 |
1403 |
9.37 |
8.14 |
0.74 |
0.85 |
| 7 |
A |
1280 |
1280 |
18.38 |
2.49 |
0.35 |
0.52 |
| 8 |
A |
1238 |
1238 |
53.52 |
4.37 |
0.36 |
0.53 |
| 9 |
A |
1168 |
1168 |
2.65 |
6.12 |
0.72 |
0.84 |
| 10 |
A |
1102 |
1102 |
0.91 |
1.31 |
0.69 |
0.81 |
| 11 |
A |
1018 |
1018 |
2.49 |
1.17 |
0.64 |
0.78 |
| 12 |
A |
901 |
901 |
8.18 |
4.66 |
0.20 |
0.34 |
| 13 |
A |
840 |
840 |
19.93 |
1.03 |
0.75 |
0.86 |
| 14 |
A |
633 |
633 |
26.06 |
10.14 |
0.26 |
0.41 |
| 15 |
A |
536 |
536 |
13.61 |
18.81 |
0.12 |
0.22 |
| 16 |
A |
378 |
378 |
9.30 |
2.18 |
0.67 |
0.80 |
| 17 |
A |
239 |
239 |
1.46 |
1.64 |
0.20 |
0.34 |
| 18 |
A |
94 |
94 |
0.52 |
1.06 |
0.64 |
0.78 |
Unscaled Zero Point Vibrational Energy (zpe) 12257.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12257.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.022 |
1.112 |
-0.390 |
| C2 |
1.291 |
0.908 |
0.410 |
| Br3 |
-1.375 |
-0.227 |
0.036 |
| Cl4 |
2.202 |
-0.581 |
-0.087 |
| H5 |
-0.420 |
2.087 |
-0.141 |
| H6 |
0.197 |
1.037 |
-1.470 |
| H7 |
1.968 |
1.758 |
0.229 |
| H8 |
1.088 |
0.817 |
1.485 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5138 | 1.9810 | 2.7771 | 1.0998 | 1.0970 | 2.1424 | 2.1775 |
C2 | 1.5138 | | 2.9216 | 1.8154 | 2.1493 | 2.1789 | 1.1016 | 1.0981 | Br3 | 1.9810 | 2.9216 | | 3.5971 | 2.5099 | 2.5178 | 3.8930 | 3.0431 | Cl4 | 2.7771 | 1.8154 | 3.5971 | | 3.7416 | 2.9247 | 2.3717 | 2.3803 | H5 | 1.0998 | 2.1493 | 2.5099 | 3.7416 | | 1.8034 | 2.4389 | 2.5558 | H6 | 1.0970 | 2.1789 | 2.5178 | 2.9247 | 1.8034 | | 2.5580 | 3.0950 | H7 | 2.1424 | 1.1016 | 3.8930 | 2.3717 | 2.4389 | 2.5580 | | 1.7994 | H8 | 2.1775 | 1.0981 | 3.0431 | 2.3803 | 2.5558 | 3.0950 | 1.7994 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.745 |
|
C1 |
C2 |
H7 |
108.978 |
| C1 |
C2 |
H8 |
111.982 |
|
C2 |
C1 |
Br3 |
112.755 |
| C2 |
C1 |
Cl4 |
37.075 |
|
C2 |
C1 |
H6 |
112.157 |
| Br3 |
C1 |
H5 |
105.519 |
|
Br3 |
C1 |
H6 |
106.187 |
| Cl4 |
C2 |
H7 |
106.196 |
|
Cl4 |
C2 |
H8 |
106.982 |
| H5 |
C1 |
H6 |
110.356 |
|
H7 |
C2 |
H8 |
109.772 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.185 |
|
|
|
| 2 |
C |
0.239 |
|
|
|
| 3 |
Br |
0.188 |
|
|
|
| 4 |
Cl |
-0.062 |
|
|
|
| 5 |
H |
-0.166 |
|
|
|
| 6 |
H |
-0.162 |
|
|
|
| 7 |
H |
-0.114 |
|
|
|
| 8 |
H |
-0.108 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.398 |
2.703 |
0.105 |
2.734 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.175 |
2.762 |
0.490 |
| y |
2.762 |
-40.619 |
-0.102 |
| z |
0.490 |
-0.102 |
-43.340 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.196 |
2.762 |
0.490 |
| y |
2.762 |
5.138 |
-0.102 |
| z |
0.490 |
-0.102 |
1.058 |
|
| Polar |
|---|
| 3z2-r2 | 2.115 |
|---|
| x2-y2 | -7.556 |
|---|
| xy | 2.762 |
|---|
| xz | 0.490 |
|---|
| yz | -0.102 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.155 |
0.184 |
-0.151 |
| y |
0.184 |
9.389 |
0.036 |
| z |
-0.151 |
0.036 |
7.831 |
<r2> (average value of r
2) Å
2
| <r2> |
227.055 |
| (<r2>)1/2 |
15.068 |
