Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -999.071478 |
| Energy at 298.15K | -999.076162 |
| Nuclear repulsion energy | 194.273313 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3124 |
2993 |
0.00 |
|
|
|
| 2 |
Ag |
1470 |
1409 |
0.00 |
|
|
|
| 3 |
Ag |
1321 |
1266 |
0.00 |
|
|
|
| 4 |
Ag |
1092 |
1046 |
0.00 |
|
|
|
| 5 |
Ag |
777 |
744 |
0.00 |
|
|
|
| 6 |
Ag |
304 |
291 |
0.00 |
|
|
|
| 7 |
Au |
3212 |
3078 |
1.47 |
|
|
|
| 8 |
Au |
1131 |
1084 |
1.04 |
|
|
|
| 9 |
Au |
773 |
741 |
2.37 |
|
|
|
| 10 |
Au |
115 |
111 |
6.86 |
|
|
|
| 11 |
Bg |
3190 |
3057 |
0.00 |
|
|
|
| 12 |
Bg |
1283 |
1230 |
0.00 |
|
|
|
| 13 |
Bg |
1002 |
960 |
0.00 |
|
|
|
| 14 |
Bu |
3132 |
3001 |
13.39 |
|
|
|
| 15 |
Bu |
1466 |
1405 |
8.45 |
|
|
|
| 16 |
Bu |
1234 |
1182 |
28.51 |
|
|
|
| 17 |
Bu |
739 |
708 |
100.49 |
|
|
|
| 18 |
Bu |
215 |
206 |
9.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12789.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12255.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.476 |
0.587 |
0.000 |
| C2 |
-0.476 |
-0.587 |
0.000 |
| Cl3 |
-0.476 |
2.113 |
0.000 |
| Cl4 |
0.476 |
-2.113 |
0.000 |
| H5 |
1.103 |
0.595 |
0.895 |
| H6 |
1.103 |
0.595 |
-0.895 |
| H7 |
-1.103 |
-0.595 |
0.895 |
| H8 |
-1.103 |
-0.595 |
-0.895 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5107 | 1.7987 | 2.7002 | 1.0931 | 1.0931 | 2.1658 | 2.1658 |
C2 | 1.5107 | | 2.7002 | 1.7987 | 2.1658 | 2.1658 | 1.0931 | 1.0931 | Cl3 | 1.7987 | 2.7002 | | 4.3325 | 2.3660 | 2.3660 | 2.9210 | 2.9210 | Cl4 | 2.7002 | 1.7987 | 4.3325 | | 2.9210 | 2.9210 | 2.3660 | 2.3660 | H5 | 1.0931 | 2.1658 | 2.3660 | 2.9210 | | 1.7900 | 2.5069 | 3.0804 | H6 | 1.0931 | 2.1658 | 2.3660 | 2.9210 | 1.7900 | | 3.0804 | 2.5069 | H7 | 2.1658 | 1.0931 | 2.9210 | 2.3660 | 2.5069 | 3.0804 | | 1.7900 | H8 | 2.1658 | 1.0931 | 2.9210 | 2.3660 | 3.0804 | 2.5069 | 1.7900 | |
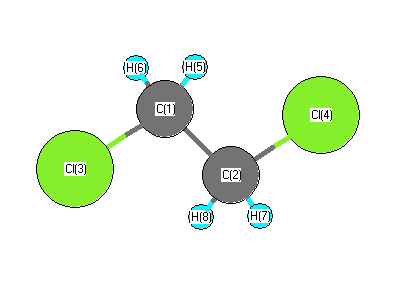 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.052 |
|
C1 |
C2 |
H7 |
111.559 |
| C1 |
C2 |
H8 |
111.559 |
|
C2 |
C1 |
Cl3 |
109.052 |
| C2 |
C1 |
H5 |
111.559 |
|
C2 |
C1 |
H6 |
111.559 |
| Cl3 |
C1 |
H5 |
107.276 |
|
Cl3 |
C1 |
H6 |
107.276 |
| Cl4 |
C2 |
H7 |
107.276 |
|
Cl4 |
C2 |
H8 |
107.276 |
| H5 |
C1 |
H6 |
109.927 |
|
H7 |
C2 |
H8 |
109.927 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.121 |
|
|
|
| 2 |
C |
0.121 |
|
|
|
| 3 |
Cl |
-0.070 |
|
|
|
| 4 |
Cl |
-0.070 |
|
|
|
| 5 |
H |
-0.026 |
|
|
|
| 6 |
H |
-0.026 |
|
|
|
| 7 |
H |
-0.026 |
|
|
|
| 8 |
H |
-0.026 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.422 |
2.050 |
0.000 |
| y |
2.050 |
-45.068 |
0.000 |
| z |
0.000 |
0.000 |
-37.714 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.969 |
2.050 |
0.000 |
| y |
2.050 |
-7.500 |
0.000 |
| z |
0.000 |
0.000 |
3.531 |
|
| Polar |
|---|
| 3z2-r2 | 7.061 |
|---|
| x2-y2 | 7.646 |
|---|
| xy | 2.050 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.333 |
-1.194 |
0.000 |
| y |
-1.194 |
10.742 |
0.000 |
| z |
0.000 |
0.000 |
6.519 |
<r2> (average value of r
2) Å
2
| <r2> |
200.913 |
| (<r2>)1/2 |
14.174 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -999.069049 |
| Energy at 298.15K | -999.073872 |
| HF Energy | -999.069049 |
| Nuclear repulsion energy | 201.858151 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3175 |
3042 |
0.41 |
|
|
|
| 2 |
A |
3109 |
2980 |
21.64 |
|
|
|
| 3 |
A |
1446 |
1386 |
1.30 |
|
|
|
| 4 |
A |
1329 |
1273 |
15.78 |
|
|
|
| 5 |
A |
1224 |
1173 |
0.23 |
|
|
|
| 6 |
A |
1065 |
1021 |
2.26 |
|
|
|
| 7 |
A |
954 |
914 |
10.60 |
|
|
|
| 8 |
A |
679 |
650 |
17.10 |
|
|
|
| 9 |
A |
261 |
250 |
1.01 |
|
|
|
| 10 |
A |
111 |
107 |
0.87 |
|
|
|
| 11 |
B |
3187 |
3054 |
2.53 |
|
|
|
| 12 |
B |
3102 |
2973 |
3.46 |
|
|
|
| 13 |
B |
1440 |
1380 |
14.17 |
|
|
|
| 14 |
B |
1297 |
1243 |
35.27 |
|
|
|
| 15 |
B |
1152 |
1104 |
0.50 |
|
|
|
| 16 |
B |
892 |
854 |
20.10 |
|
|
|
| 17 |
B |
699 |
670 |
25.68 |
|
|
|
| 18 |
B |
413 |
396 |
8.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12766.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 12234.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.296 |
0.693 |
0.894 |
| C2 |
-0.296 |
-0.693 |
0.894 |
| Cl3 |
-0.296 |
1.693 |
-0.470 |
| Cl4 |
0.296 |
-1.693 |
-0.470 |
| H5 |
0.003 |
1.211 |
1.814 |
| H6 |
1.387 |
0.662 |
0.821 |
| H7 |
-0.003 |
-1.211 |
1.814 |
| H8 |
-1.387 |
-0.662 |
0.821 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5075 | 1.7924 | 2.7487 | 1.0959 | 1.0936 | 2.1355 | 2.1626 |
C2 | 1.5075 | | 2.7487 | 1.7924 | 2.1355 | 2.1626 | 1.0959 | 1.0936 | Cl3 | 1.7924 | 2.7487 | | 3.4384 | 2.3537 | 2.3591 | 3.7064 | 2.8996 | Cl4 | 2.7487 | 1.7924 | 3.4384 | | 3.7064 | 2.8996 | 2.3537 | 2.3591 | H5 | 1.0959 | 2.1355 | 2.3537 | 3.7064 | | 1.7897 | 2.4213 | 2.5349 | H6 | 1.0936 | 2.1626 | 2.3591 | 2.8996 | 1.7897 | | 2.5349 | 3.0744 | H7 | 2.1355 | 1.0959 | 3.7064 | 2.3537 | 2.4213 | 2.5349 | | 1.7897 | H8 | 2.1626 | 1.0936 | 2.8996 | 2.3591 | 2.5349 | 3.0744 | 1.7897 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.528 |
|
C1 |
C2 |
H7 |
109.202 |
| C1 |
C2 |
H8 |
111.499 |
|
C2 |
C1 |
Cl3 |
112.528 |
| C2 |
C1 |
H5 |
109.202 |
|
C2 |
C1 |
H6 |
111.499 |
| Cl3 |
C1 |
H5 |
106.660 |
|
Cl3 |
C1 |
H6 |
107.160 |
| Cl4 |
C2 |
H7 |
106.660 |
|
Cl4 |
C2 |
H8 |
107.160 |
| H5 |
C1 |
H6 |
109.645 |
|
H7 |
C2 |
H8 |
109.645 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.065 |
|
|
|
| 2 |
C |
0.065 |
|
|
|
| 3 |
Cl |
-0.050 |
|
|
|
| 4 |
Cl |
-0.050 |
|
|
|
| 5 |
H |
-0.020 |
|
|
|
| 6 |
H |
0.005 |
|
|
|
| 7 |
H |
-0.020 |
|
|
|
| 8 |
H |
0.005 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.798 |
2.798 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.605 |
1.259 |
0.000 |
| y |
1.259 |
-42.538 |
0.000 |
| z |
0.000 |
0.000 |
-35.327 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.327 |
1.259 |
0.000 |
| y |
1.259 |
-6.072 |
0.000 |
| z |
0.000 |
0.000 |
4.744 |
|
| Polar |
|---|
| 3z2-r2 | 9.489 |
|---|
| x2-y2 | 4.933 |
|---|
| xy | 1.259 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.817 |
-0.465 |
0.000 |
| y |
-0.465 |
9.048 |
0.000 |
| z |
0.000 |
0.000 |
8.081 |
<r2> (average value of r
2) Å
2
| <r2> |
164.037 |
| (<r2>)1/2 |
12.808 |
