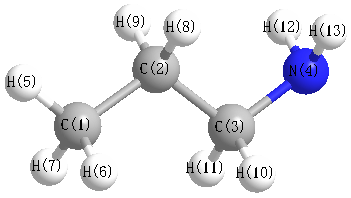
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3521 |
3374 |
1.79 |
|
|
|
| 2 |
A' |
3137 |
3007 |
33.47 |
|
|
|
| 3 |
A' |
3052 |
2925 |
75.38 |
|
|
|
| 4 |
A' |
3048 |
2921 |
17.99 |
|
|
|
| 5 |
A' |
3038 |
2911 |
13.80 |
|
|
|
| 6 |
A' |
1650 |
1581 |
23.15 |
|
|
|
| 7 |
A' |
1488 |
1426 |
6.69 |
|
|
|
| 8 |
A' |
1475 |
1413 |
1.14 |
|
|
|
| 9 |
A' |
1466 |
1405 |
0.07 |
|
|
|
| 10 |
A' |
1394 |
1336 |
0.79 |
|
|
|
| 11 |
A' |
1390 |
1332 |
8.94 |
|
|
|
| 12 |
A' |
1309 |
1255 |
4.25 |
|
|
|
| 13 |
A' |
1143 |
1096 |
3.86 |
|
|
|
| 14 |
A' |
1116 |
1069 |
18.31 |
|
|
|
| 15 |
A' |
1059 |
1015 |
0.31 |
|
|
|
| 16 |
A' |
909 |
871 |
56.85 |
|
|
|
| 17 |
A' |
828 |
793 |
117.46 |
|
|
|
| 18 |
A' |
448 |
429 |
3.60 |
|
|
|
| 19 |
A' |
265 |
254 |
3.90 |
|
|
|
| 20 |
A" |
3606 |
3456 |
0.67 |
|
|
|
| 21 |
A" |
3128 |
2997 |
58.23 |
|
|
|
| 22 |
A" |
3099 |
2970 |
35.72 |
|
|
|
| 23 |
A" |
3071 |
2943 |
1.44 |
|
|
|
| 24 |
A" |
1476 |
1415 |
7.59 |
|
|
|
| 25 |
A" |
1380 |
1323 |
0.13 |
|
|
|
| 26 |
A" |
1313 |
1259 |
0.53 |
|
|
|
| 27 |
A" |
1231 |
1180 |
0.02 |
|
|
|
| 28 |
A" |
1028 |
985 |
0.01 |
|
|
|
| 29 |
A" |
854 |
818 |
0.90 |
|
|
|
| 30 |
A" |
739 |
708 |
2.50 |
|
|
|
| 31 |
A" |
271 |
260 |
35.32 |
|
|
|
| 32 |
A" |
219 |
210 |
8.47 |
|
|
|
| 33 |
A" |
127 |
122 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26638.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 25527.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.378 |
|
|
|
| 2 |
C |
0.554 |
|
|
|
| 3 |
C |
0.698 |
|
|
|
| 4 |
N |
-0.285 |
|
|
|
| 5 |
H |
-0.144 |
|
|
|
| 6 |
H |
-0.113 |
|
|
|
| 7 |
H |
-0.113 |
|
|
|
| 8 |
H |
-0.216 |
|
|
|
| 9 |
H |
-0.216 |
|
|
|
| 10 |
H |
-0.250 |
|
|
|
| 11 |
H |
-0.250 |
|
|
|
| 12 |
H |
-0.021 |
|
|
|
| 13 |
H |
-0.021 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.104 |
1.385 |
0.000 |
1.389 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.547 |
-1.937 |
0.000 |
| y |
-1.937 |
-32.833 |
0.000 |
| z |
0.000 |
0.000 |
-25.738 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.739 |
-1.937 |
0.000 |
| y |
-1.937 |
-6.690 |
0.000 |
| z |
0.000 |
0.000 |
3.951 |
|
| Polar |
|---|
| 3z2-r2 | 7.903 |
|---|
| x2-y2 | 6.286 |
|---|
| xy | -1.937 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.985 |
0.722 |
0.000 |
| y |
0.722 |
7.776 |
0.000 |
| z |
0.000 |
0.000 |
6.662 |
<r2> (average value of r
2) Å
2
| <r2> |
112.839 |
| (<r2>)1/2 |
10.623 |