return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-174.027080
Energy at 298.15K-174.037425
HF Energy-173.309200
Nuclear repulsion energy130.022583
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3444 3364        
2 A' 3137 3065        
3 A' 3054 2983        
4 A' 3048 2977        
5 A' 3030 2960        
6 A' 1718 1679        
7 A' 1555 1519        
8 A' 1542 1506        
9 A' 1533 1497        
10 A' 1457 1423        
11 A' 1434 1401        
12 A' 1351 1320        
13 A' 1176 1149        
14 A' 1118 1092        
15 A' 1066 1041        
16 A' 967 945        
17 A' 882 861        
18 A' 453 443        
19 A' 270 264        
20 A" 3537 3455        
21 A" 3131 3058        
22 A" 3099 3027        
23 A" 3068 2997        
24 A" 1546 1511        
25 A" 1417 1384        
26 A" 1350 1319        
27 A" 1278 1248        
28 A" 1054 1030        
29 A" 878 858        
30 A" 755 737        
31 A" 311 304        
32 A" 235 230        
33 A" 136 133        

Unscaled Zero Point Vibrational Energy (zpe) 27014.2 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 26390.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.81034 0.12137 0.11434

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.446 1.288 0.000
C2 0.000 0.758 0.000
C3 -0.055 -0.783 0.000
N4 -1.406 -1.383 0.000
H5 1.472 2.392 0.000
H6 1.995 0.935 0.894
H7 1.995 0.935 -0.894
H8 -0.542 1.137 0.890
H9 -0.542 1.137 -0.890
H10 0.483 -1.170 -0.888
H11 0.483 -1.170 0.888
H12 -1.927 -1.046 -0.815
H13 -1.927 -1.046 0.815

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.54022.55703.90671.10471.10631.10632.18322.18322.78512.78514.18164.1816
C21.54021.54132.56052.19942.19282.19281.10871.10872.17652.17652.76212.7621
C32.55701.54131.47833.52262.81932.81932.17142.17141.10751.10752.05892.0589
N43.90672.56051.47834.74634.21094.21092.80842.80842.09732.09731.02451.0245
H51.10472.19943.52264.74631.78741.78742.53402.53403.80203.80204.90234.9023
H61.10632.19282.81934.21091.78741.78752.54443.10723.14492.59184.71394.3939
H71.10632.19282.81934.21091.78741.78753.10722.54442.59183.14494.39394.7139
H82.18321.10872.17142.80842.53402.54443.10721.77963.08742.52453.09682.5862
H92.18321.10872.17142.80842.53403.10722.54441.77962.52453.08742.58623.0968
H102.78512.17651.10752.09733.80203.14492.59183.08742.52451.77512.41382.9531
H112.78512.17651.10752.09733.80202.59183.14492.52453.08741.77512.95312.4138
H124.18162.76212.05891.02454.90234.71394.39393.09682.58622.41382.95311.6304
H134.18162.76212.05891.02454.90234.39394.71392.58623.09682.95312.41381.6304

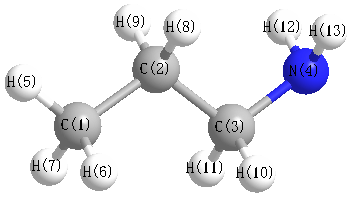
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.156 C1 C2 H8 109.940
C1 C2 H9 109.940 C2 C1 H5 111.460
C2 C1 H6 110.842 C2 C1 H7 110.842
C2 C3 N4 115.969 C2 C3 H10 109.418
C2 C3 H11 109.418 C3 C2 H8 108.951
C3 C2 H9 108.951 C3 N4 H12 109.358
C3 N4 H13 109.358 N4 C3 H10 107.544
N4 C3 H11 107.544 H5 C1 H6 107.884
H5 C1 H7 107.884 H6 C1 H7 107.781
H8 C2 H9 106.745 H10 C3 H11 106.528
H12 N4 H13 105.445
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability