return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: B2PLYP=FULLultrafine/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
 hartrees
Energy at 0K-174.311095
Energy at 298.15K 
HF Energy-174.097021
Nuclear repulsion energy130.697595
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3494 3362 2.04      
2 A' 3128 3010 34.92      
3 A' 3050 2935 75.92      
4 A' 3045 2931 10.35      
5 A' 3034 2920 18.41      
6 A' 1651 1589 21.57      
7 A' 1497 1441 5.01      
8 A' 1484 1428 0.99      
9 A' 1475 1419 0.06      
10 A' 1400 1348 2.38      
11 A' 1391 1339 6.04      
12 A' 1315 1266 5.27      
13 A' 1138 1095 2.64      
14 A' 1100 1058 18.51      
15 A' 1045 1005 0.55      
16 A' 910 876 63.50      
17 A' 831 800 106.69      
18 A' 447 430 3.06      
19 A' 267 257 3.86      
20 A" 3581 3446 0.53      
21 A" 3120 3002 63.50      
22 A" 3096 2979 31.95      
23 A" 3067 2952 2.54      
24 A" 1488 1432 6.71      
25 A" 1382 1330 0.18      
26 A" 1317 1267 0.61      
27 A" 1236 1190 0.00      
28 A" 1029 991 0.07      
29 A" 857 825 0.91      
30 A" 742 714 1.60      
31 A" 271 260 33.82      
32 A" 223 214 9.30      
33 A" 129 125 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 26619.7 cm-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 25616.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/aug-cc-pVDZ
ABC
0.82340 0.12227 0.11529

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.433 1.292 0.000
C2 0.000 0.753 0.000
C3 -0.053 -0.780 0.000
N4 -1.392 -1.381 0.000
H5 1.448 2.390 0.000
H6 1.984 0.949 0.888
H7 1.984 0.949 -0.888
H8 -0.541 1.129 0.883
H9 -0.541 1.129 -0.883
H10 0.484 -1.164 -0.881
H11 0.484 -1.164 0.881
H12 -1.919 -1.071 -0.814
H13 -1.919 -1.071 0.814

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.53102.55003.88861.09811.09961.09962.16862.16862.77662.77664.18094.1809
C21.53101.53362.54692.18632.18222.18221.10191.10192.16402.16402.76942.7694
C32.55001.53361.46703.50802.81582.81582.15912.15911.10081.10082.05612.0561
N43.88862.54691.46704.72074.19654.19652.79282.79282.08342.08341.01811.0181
H51.09812.18633.50804.72071.77491.77492.51582.51583.78663.78664.89694.8969
H61.09962.18222.81584.19651.77491.77542.53123.08913.13752.59164.71254.3950
H71.09962.18222.81584.19651.77491.77543.08912.53122.59163.13754.39504.7125
H82.16861.10192.15912.79282.51582.53123.08911.76623.06902.51143.10122.5963
H92.16861.10192.15912.79282.51583.08912.53121.76622.51143.06902.59633.1012
H102.77662.16401.10082.08343.78663.13752.59163.06902.51141.76182.40522.9417
H112.77662.16401.10082.08343.78662.59163.13752.51143.06901.76182.94172.4052
H124.18092.76942.05611.01814.89694.71254.39503.10122.59632.40522.94171.6285
H134.18092.76942.05611.01814.89694.39504.71252.59633.10122.94172.40521.6285

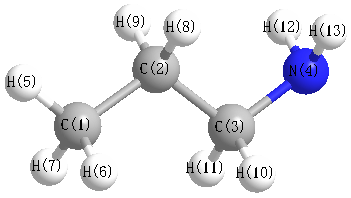
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.627 C1 C2 H8 109.829
C1 C2 H9 109.829 C2 C1 H5 111.455
C2 C1 H6 111.041 C2 C1 H7 111.041
C2 C3 N4 116.146 C2 C3 H10 109.365
C2 C3 H11 109.365 C3 C2 H8 108.915
C3 C2 H9 108.915 C3 N4 H12 110.352
C3 N4 H13 110.352 N4 C3 H10 107.601
N4 C3 H11 107.601 H5 C1 H6 107.732
H5 C1 H7 107.732 H6 C1 H7 107.666
H8 C2 H9 106.534 H10 C3 H11 106.310
H12 N4 H13 106.220
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability