return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5CN (ethyl cyanide)

using model chemistry: CCSD=FULL/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes Cs 1A'
Energy calculated at CCSD=FULL/aug-cc-pVDZ
 hartrees
Energy at 0K-171.613409
Energy at 298.15K 
HF Energy-170.980014
Nuclear repulsion energy101.979523
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3146 3146 20.51      
2 A' 3077 3077 7.69      
3 A' 3061 3061 17.12      
4 A' 2321 2321 1.57      
5 A' 1498 1498 3.83      
6 A' 1474 1474 3.56      
7 A' 1416 1416 0.84      
8 A' 1350 1350 4.79      
9 A' 1098 1098 2.57      
10 A' 1035 1035 0.85      
11 A' 843 843 0.62      
12 A' 535 535 0.48      
13 A' 207 207 4.47      
14 A" 3149 3149 20.36      
15 A" 3121 3121 0.38      
16 A" 1492 1492 5.84      
17 A" 1289 1289 0.03      
18 A" 1114 1114 0.47      
19 A" 786 786 1.79      
20 A" 380 380 0.49      
21 A" 222 222 1.13      

Unscaled Zero Point Vibrational Energy (zpe) 16306.5 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16306.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCSD=FULL/aug-cc-pVDZ
ABC
0.91327 0.15417 0.13885

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.519 0.572 0.000
C2 0.000 0.827 0.000
C3 -0.774 -0.437 0.000
N4 -1.367 -1.446 0.000
H5 2.057 1.532 0.000
H6 1.819 0.002 0.893
H7 1.819 0.002 -0.893
H8 -0.298 1.407 0.888
H9 -0.298 1.407 -0.888

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9
C11.54032.50483.52121.10121.10091.10092.18852.1885
C21.54031.48232.65252.17442.18812.18811.10181.1018
C32.50481.48231.17043.44842.77732.77732.10122.1012
N43.52122.65251.17044.53803.61173.61173.17333.1733
H51.10122.17443.44844.53801.78791.78792.52032.5203
H61.10092.18812.77733.61171.78791.78592.54133.1033
H71.10092.18812.77733.61171.78791.78593.10332.5413
H82.18851.10182.10123.17332.52032.54133.10331.7764
H92.18851.10182.10123.17332.52033.10332.54131.7764

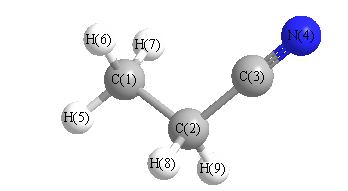
picture of ethyl cyanide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.918 C1 C2 H8 110.757
C1 C2 H9 110.757 C2 C1 H5 109.686
C2 C1 H6 110.777 C2 C1 H7 110.777
C2 C3 N4 178.975 C3 C2 H8 107.898
C3 C2 H9 107.898 H5 C1 H6 108.567
H5 C1 H7 108.567 H6 C1 H7 108.402
H8 C2 H9 107.437
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability