Jump to
S1C2
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.466103 |
| Energy at 298.15K | -207.469435 |
| HF Energy | -206.799813 |
| Nuclear repulsion energy | 101.246637 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3843 |
3843 |
43.71 |
|
|
|
| 2 |
A |
3144 |
3144 |
3.61 |
|
|
|
| 3 |
A |
3066 |
3066 |
21.01 |
|
|
|
| 4 |
A |
2320 |
2320 |
2.07 |
|
|
|
| 5 |
A |
1490 |
1490 |
2.00 |
|
|
|
| 6 |
A |
1426 |
1426 |
48.78 |
|
|
|
| 7 |
A |
1375 |
1375 |
2.89 |
|
|
|
| 8 |
A |
1229 |
1229 |
15.29 |
|
|
|
| 9 |
A |
1100 |
1100 |
94.13 |
|
|
|
| 10 |
A |
992 |
992 |
17.87 |
|
|
|
| 11 |
A |
903 |
903 |
17.57 |
|
|
|
| 12 |
A |
572 |
572 |
1.49 |
|
|
|
| 13 |
A |
372 |
372 |
44.98 |
|
|
|
| 14 |
A |
298 |
298 |
91.54 |
|
|
|
| 15 |
A |
211 |
211 |
11.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11170.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11170.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.579 |
0.594 |
0.038 |
| C2 |
0.831 |
0.111 |
-0.006 |
| O3 |
-1.522 |
-0.455 |
-0.111 |
| H4 |
-0.722 |
1.155 |
0.978 |
| H5 |
-0.741 |
1.283 |
-0.803 |
| H6 |
-1.414 |
-1.074 |
0.621 |
| N7 |
1.935 |
-0.278 |
-0.015 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4915 | 1.4187 | 1.1041 | 1.0995 | 1.9539 | 2.6613 |
C2 | 1.4915 | | 2.4232 | 2.1146 | 2.1169 | 2.6148 | 1.1700 | O3 | 1.4187 | 2.4232 | | 2.1028 | 2.0275 | 0.9649 | 3.4630 | H4 | 1.1041 | 2.1146 | 2.1028 | | 1.7856 | 2.3615 | 3.1779 | H5 | 1.0995 | 2.1169 | 2.0275 | 1.7856 | | 2.8348 | 3.1965 | H6 | 1.9539 | 2.6148 | 0.9649 | 2.3615 | 2.8348 | | 3.4999 | N7 | 2.6613 | 1.1700 | 3.4630 | 3.1779 | 3.1965 | 3.4999 | |
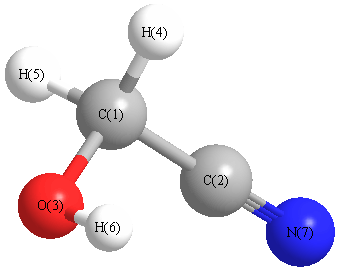 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.646 |
|
C1 |
O3 |
H6 |
108.619 |
| C2 |
C1 |
O3 |
112.723 |
|
C2 |
C1 |
H4 |
108.187 |
| C2 |
C1 |
H5 |
108.638 |
|
O3 |
C1 |
H4 |
112.329 |
| O3 |
C1 |
H5 |
106.560 |
|
H4 |
C1 |
H5 |
108.255 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
