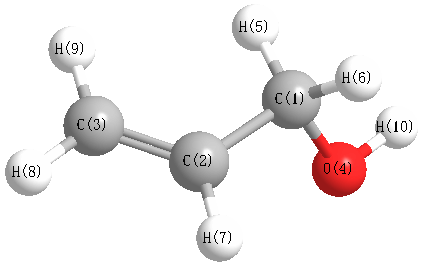
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3886 |
3724 |
32.03 |
|
|
|
| 2 |
A |
3262 |
3126 |
12.60 |
|
|
|
| 3 |
A |
3188 |
3055 |
6.11 |
|
|
|
| 4 |
A |
3164 |
3032 |
9.77 |
|
|
|
| 5 |
A |
3054 |
2927 |
36.97 |
|
|
|
| 6 |
A |
3003 |
2878 |
50.94 |
|
|
|
| 7 |
A |
1736 |
1663 |
2.76 |
|
|
|
| 8 |
A |
1487 |
1425 |
2.23 |
|
|
|
| 9 |
A |
1461 |
1400 |
15.59 |
|
|
|
| 10 |
A |
1407 |
1349 |
6.37 |
|
|
|
| 11 |
A |
1310 |
1255 |
1.13 |
|
|
|
| 12 |
A |
1271 |
1218 |
23.25 |
|
|
|
| 13 |
A |
1234 |
1182 |
45.00 |
|
|
|
| 14 |
A |
1158 |
1110 |
17.22 |
|
|
|
| 15 |
A |
1086 |
1041 |
95.59 |
|
|
|
| 16 |
A |
1024 |
981 |
15.36 |
|
|
|
| 17 |
A |
972 |
931 |
37.36 |
|
|
|
| 18 |
A |
961 |
921 |
14.43 |
|
|
|
| 19 |
A |
923 |
884 |
0.63 |
|
|
|
| 20 |
A |
654 |
627 |
6.18 |
|
|
|
| 21 |
A |
448 |
429 |
3.74 |
|
|
|
| 22 |
A |
330 |
316 |
9.75 |
|
|
|
| 23 |
A |
258 |
247 |
112.85 |
|
|
|
| 24 |
A |
114 |
109 |
1.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18694.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 17914.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.776 |
|
|
|
| 2 |
C |
0.902 |
|
|
|
| 3 |
C |
0.390 |
|
|
|
| 4 |
O |
-0.495 |
|
|
|
| 5 |
H |
-0.279 |
|
|
|
| 6 |
H |
-0.142 |
|
|
|
| 7 |
H |
-0.486 |
|
|
|
| 8 |
H |
-0.364 |
|
|
|
| 9 |
H |
-0.408 |
|
|
|
| 10 |
H |
0.106 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.330 |
1.331 |
0.840 |
1.608 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.222 |
2.853 |
1.726 |
| y |
2.853 |
-24.826 |
-1.342 |
| z |
1.726 |
-1.342 |
-27.043 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.713 |
2.853 |
1.726 |
| y |
2.853 |
-0.693 |
-1.342 |
| z |
1.726 |
-1.342 |
-4.020 |
|
| Polar |
|---|
| 3z2-r2 | -8.039 |
|---|
| x2-y2 | 3.604 |
|---|
| xy | 2.853 |
|---|
| xz | 1.726 |
|---|
| yz | -1.342 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.602 |
-0.224 |
0.553 |
| y |
-0.224 |
6.027 |
-0.175 |
| z |
0.553 |
-0.175 |
5.559 |
<r2> (average value of r
2) Å
2
| <r2> |
93.190 |
| (<r2>)1/2 |
9.653 |