Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.923061 |
| Energy at 298.15K | -207.929276 |
| Nuclear repulsion energy | 118.494497 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4188 |
3814 |
125.46 |
|
|
|
| 2 |
A |
3286 |
2992 |
29.36 |
|
|
|
| 3 |
A |
3271 |
2979 |
1.08 |
|
|
|
| 4 |
A |
3172 |
2888 |
24.94 |
|
|
|
| 5 |
A |
1940 |
1767 |
8.22 |
|
|
|
| 6 |
A |
1582 |
1441 |
19.81 |
|
|
|
| 7 |
A |
1567 |
1427 |
12.42 |
|
|
|
| 8 |
A |
1508 |
1373 |
21.22 |
|
|
|
| 9 |
A |
1408 |
1282 |
59.36 |
|
|
|
| 10 |
A |
1239 |
1128 |
6.07 |
|
|
|
| 11 |
A |
1142 |
1040 |
150.70 |
|
|
|
| 12 |
A |
981 |
893 |
3.13 |
|
|
|
| 13 |
A |
611 |
556 |
15.16 |
|
|
|
| 14 |
A |
351 |
320 |
2.82 |
|
|
|
| 15 |
A |
3228 |
2940 |
25.28 |
|
|
|
| 16 |
A |
1573 |
1432 |
8.08 |
|
|
|
| 17 |
A |
1188 |
1081 |
3.32 |
|
|
|
| 18 |
A |
996 |
907 |
8.17 |
|
|
|
| 19 |
A |
379 |
345 |
146.72 |
|
|
|
| 20 |
A |
318 |
290 |
4.60 |
|
|
|
| 21 |
A |
217 |
197 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17072.3 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 15546.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.297 |
1.296 |
0.000 |
| C2 |
0.000 |
0.549 |
0.000 |
| N3 |
0.018 |
-0.704 |
0.000 |
| O4 |
1.298 |
-1.199 |
0.000 |
| H5 |
1.192 |
-2.135 |
0.000 |
| H6 |
-2.134 |
0.603 |
0.000 |
| H7 |
-1.361 |
1.938 |
0.880 |
| H8 |
-1.361 |
1.938 |
-0.880 |
| H9 |
0.935 |
1.104 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4965 | 2.3930 | 3.6002 | 4.2388 | 1.0870 | 1.0914 | 1.0914 | 2.2399 |
C2 | 1.4965 | | 1.2530 | 2.1779 | 2.9372 | 2.1349 | 2.1346 | 2.1346 | 1.0872 | N3 | 2.3930 | 1.2530 | | 1.3734 | 1.8516 | 2.5174 | 3.1071 | 3.1071 | 2.0272 | O4 | 3.6002 | 2.1779 | 1.3734 | | 0.9420 | 3.8769 | 4.2060 | 4.2060 | 2.3319 | H5 | 4.2388 | 2.9372 | 1.8516 | 0.9420 | | 4.3082 | 4.8872 | 4.8872 | 3.2496 | H6 | 1.0870 | 2.1349 | 2.5174 | 3.8769 | 4.3082 | | 1.7762 | 1.7762 | 3.1098 | H7 | 1.0914 | 2.1346 | 3.1071 | 4.2060 | 4.8872 | 1.7762 | | 1.7605 | 2.5966 | H8 | 1.0914 | 2.1346 | 3.1071 | 4.2060 | 4.8872 | 1.7762 | 1.7605 | | 2.5966 | H9 | 2.2399 | 1.0872 | 2.0272 | 2.3319 | 3.2496 | 3.1098 | 2.5966 | 2.5966 | |
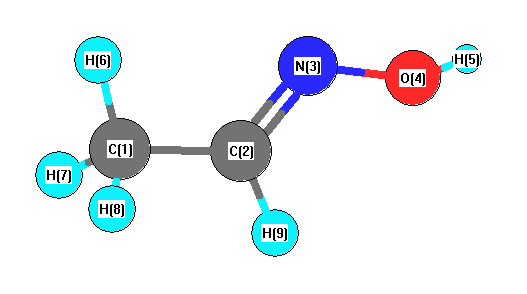 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
120.740 |
|
C1 |
C2 |
H9 |
119.371 |
| C2 |
C1 |
H6 |
110.448 |
|
C2 |
C1 |
H7 |
110.162 |
| C2 |
C1 |
H8 |
110.162 |
|
C2 |
N3 |
O4 |
111.954 |
| N3 |
C2 |
H9 |
119.889 |
|
N3 |
O4 |
H5 |
104.659 |
| H6 |
C1 |
H7 |
109.247 |
|
H6 |
C1 |
H8 |
109.247 |
| H7 |
C1 |
H8 |
107.517 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.149 |
|
|
|
| 2 |
C |
0.677 |
|
|
|
| 3 |
N |
-0.211 |
|
|
|
| 4 |
O |
-0.560 |
|
|
|
| 5 |
H |
0.144 |
|
|
|
| 6 |
H |
-0.053 |
|
|
|
| 7 |
H |
0.024 |
|
|
|
| 8 |
H |
0.024 |
|
|
|
| 9 |
H |
-0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.441 |
0.888 |
0.000 |
0.991 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.732 |
-1.946 |
0.000 |
| y |
-1.946 |
-19.000 |
0.000 |
| z |
0.000 |
0.000 |
-25.446 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.509 |
-1.946 |
0.000 |
| y |
-1.946 |
6.589 |
0.000 |
| z |
0.000 |
0.000 |
-3.080 |
|
| Polar |
|---|
| 3z2-r2 | -6.160 |
|---|
| x2-y2 | -6.732 |
|---|
| xy | -1.946 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.753 |
-1.245 |
0.000 |
| y |
-1.245 |
7.163 |
0.000 |
| z |
0.000 |
0.000 |
4.189 |
<r2> (average value of r
2) Å
2
| <r2> |
90.808 |
| (<r2>)1/2 |
9.529 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.921623 |
| Energy at 298.15K | -207.927812 |
| HF Energy | -207.921623 |
| Nuclear repulsion energy | 120.921571 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4192 |
3817 |
129.11 |
|
|
|
| 2 |
A' |
3326 |
3028 |
20.33 |
|
|
|
| 3 |
A' |
3311 |
3015 |
0.57 |
|
|
|
| 4 |
A' |
3176 |
2892 |
20.03 |
|
|
|
| 5 |
A' |
1948 |
1774 |
17.45 |
|
|
|
| 6 |
A' |
1576 |
1435 |
22.03 |
|
|
|
| 7 |
A' |
1519 |
1383 |
27.13 |
|
|
|
| 8 |
A' |
1508 |
1373 |
48.14 |
|
|
|
| 9 |
A' |
1462 |
1331 |
23.60 |
|
|
|
| 10 |
A' |
1232 |
1122 |
12.31 |
|
|
|
| 11 |
A' |
1070 |
974 |
142.07 |
|
|
|
| 12 |
A' |
972 |
885 |
0.70 |
|
|
|
| 13 |
A' |
726 |
661 |
15.10 |
|
|
|
| 14 |
A' |
336 |
306 |
1.61 |
|
|
|
| 15 |
A" |
3230 |
2941 |
22.54 |
|
|
|
| 16 |
A" |
1582 |
1441 |
8.68 |
|
|
|
| 17 |
A" |
1171 |
1066 |
1.88 |
|
|
|
| 18 |
A" |
958 |
872 |
9.67 |
|
|
|
| 19 |
A" |
529 |
482 |
20.39 |
|
|
|
| 20 |
A" |
400 |
364 |
120.73 |
|
|
|
| 21 |
A" |
60 |
55 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17140.9 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 15608.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.454 |
0.477 |
0.000 |
| C2 |
0.000 |
0.845 |
0.000 |
| N3 |
1.001 |
0.087 |
0.000 |
| O4 |
0.678 |
-1.247 |
0.000 |
| H5 |
1.511 |
-1.686 |
0.000 |
| H6 |
-1.602 |
-0.596 |
0.000 |
| H7 |
-1.935 |
0.909 |
0.879 |
| H8 |
-1.935 |
0.909 |
-0.879 |
| H9 |
0.257 |
1.897 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4999 | 2.4858 | 2.7418 | 3.6703 | 1.0835 | 1.0913 | 1.0913 | 2.2240 |
C2 | 1.4999 | | 1.2557 | 2.1993 | 2.9480 | 2.1552 | 2.1266 | 2.1266 | 1.0832 | N3 | 2.4858 | 1.2557 | | 1.3725 | 1.8450 | 2.6913 | 3.1733 | 3.1733 | 1.9572 | O4 | 2.7418 | 2.1993 | 1.3725 | | 0.9418 | 2.3711 | 3.4997 | 3.4997 | 3.1724 | H5 | 3.6703 | 2.9480 | 1.8450 | 0.9418 | | 3.2984 | 4.4026 | 4.4026 | 3.7965 | H6 | 1.0835 | 2.1552 | 2.6913 | 2.3711 | 3.2984 | | 1.7745 | 1.7745 | 3.1106 | H7 | 1.0913 | 2.1266 | 3.1733 | 3.4997 | 4.4026 | 1.7745 | | 1.7587 | 2.5610 | H8 | 1.0913 | 2.1266 | 3.1733 | 3.4997 | 4.4026 | 1.7745 | 1.7587 | | 2.5610 | H9 | 2.2240 | 1.0832 | 1.9572 | 3.1724 | 3.7965 | 3.1106 | 2.5610 | 2.5610 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.648 |
|
C1 |
C2 |
H9 |
117.948 |
| C2 |
C1 |
H6 |
112.062 |
|
C2 |
C1 |
H7 |
109.295 |
| C2 |
C1 |
H8 |
109.295 |
|
C2 |
N3 |
O4 |
113.531 |
| N3 |
C2 |
H9 |
113.404 |
|
N3 |
O4 |
H5 |
104.181 |
| H6 |
C1 |
H7 |
109.352 |
|
H6 |
C1 |
H8 |
109.352 |
| H7 |
C1 |
H8 |
107.369 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.255 |
|
|
|
| 2 |
C |
0.562 |
|
|
|
| 3 |
N |
-0.147 |
|
|
|
| 4 |
O |
-0.580 |
|
|
|
| 5 |
H |
0.146 |
|
|
|
| 6 |
H |
-0.044 |
|
|
|
| 7 |
H |
0.013 |
|
|
|
| 8 |
H |
0.013 |
|
|
|
| 9 |
H |
-0.218 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.915 |
0.406 |
0.000 |
1.001 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.338 |
-3.943 |
0.000 |
| y |
-3.943 |
-21.470 |
0.000 |
| z |
0.000 |
0.000 |
-25.473 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.133 |
-3.943 |
0.000 |
| y |
-3.943 |
2.936 |
0.000 |
| z |
0.000 |
0.000 |
-3.069 |
|
| Polar |
|---|
| 3z2-r2 | -6.138 |
|---|
| x2-y2 | -1.869 |
|---|
| xy | -3.943 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.299 |
-1.116 |
0.000 |
| y |
-1.116 |
6.318 |
0.000 |
| z |
0.000 |
0.000 |
4.194 |
<r2> (average value of r
2) Å
2
| <r2> |
78.555 |
| (<r2>)1/2 |
8.863 |
