Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.114364 |
| Energy at 298.15K | -209.120430 |
| Nuclear repulsion energy | 117.587428 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3876 |
3715 |
96.36 |
|
|
|
| 2 |
A |
3173 |
3040 |
10.92 |
|
|
|
| 3 |
A |
3126 |
2995 |
13.45 |
|
|
|
| 4 |
A |
3054 |
2926 |
18.41 |
|
|
|
| 5 |
A |
1764 |
1691 |
0.62 |
|
|
|
| 6 |
A |
1464 |
1403 |
14.53 |
|
|
|
| 7 |
A |
1444 |
1384 |
8.43 |
|
|
|
| 8 |
A |
1381 |
1323 |
24.45 |
|
|
|
| 9 |
A |
1290 |
1236 |
48.26 |
|
|
|
| 10 |
A |
1157 |
1109 |
3.66 |
|
|
|
| 11 |
A |
1044 |
1000 |
162.15 |
|
|
|
| 12 |
A |
918 |
880 |
7.38 |
|
|
|
| 13 |
A |
574 |
550 |
13.84 |
|
|
|
| 14 |
A |
325 |
312 |
3.42 |
|
|
|
| 15 |
A |
3120 |
2990 |
12.48 |
|
|
|
| 16 |
A |
1451 |
1391 |
9.20 |
|
|
|
| 17 |
A |
1068 |
1024 |
1.61 |
|
|
|
| 18 |
A |
908 |
870 |
9.13 |
|
|
|
| 19 |
A |
425 |
407 |
130.26 |
|
|
|
| 20 |
A |
292 |
280 |
0.16 |
|
|
|
| 21 |
A |
204 |
196 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16028.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15359.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.294 |
1.299 |
0.000 |
| C2 |
0.000 |
0.564 |
0.000 |
| N3 |
0.008 |
-0.709 |
0.000 |
| O4 |
1.307 |
-1.208 |
0.000 |
| H5 |
1.174 |
-2.160 |
0.000 |
| H6 |
-2.133 |
0.596 |
0.000 |
| H7 |
-1.369 |
1.948 |
0.884 |
| H8 |
-1.369 |
1.948 |
-0.884 |
| H9 |
0.946 |
1.118 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4884 | 2.3936 | 3.6125 | 4.2500 | 1.0947 | 1.0989 | 1.0989 | 2.2477 |
C2 | 1.4884 | | 1.2726 | 2.2014 | 2.9664 | 2.1328 | 2.1380 | 2.1380 | 1.0971 | N3 | 2.3936 | 1.2726 | | 1.3909 | 1.8617 | 2.5072 | 3.1205 | 3.1205 | 2.0541 | O4 | 3.6125 | 2.2014 | 1.3909 | | 0.9616 | 3.8835 | 4.2308 | 4.2308 | 2.3542 | H5 | 4.2500 | 2.9664 | 1.8617 | 0.9616 | | 4.3049 | 4.9120 | 4.9120 | 3.2868 | H6 | 1.0947 | 2.1328 | 2.5072 | 3.8835 | 4.3049 | | 1.7871 | 1.7871 | 3.1231 | H7 | 1.0989 | 2.1380 | 3.1205 | 4.2308 | 4.9120 | 1.7871 | | 1.7677 | 2.6133 | H8 | 1.0989 | 2.1380 | 3.1205 | 4.2308 | 4.9120 | 1.7871 | 1.7677 | | 2.6133 | H9 | 2.2477 | 1.0971 | 2.0541 | 2.3542 | 3.2868 | 3.1231 | 2.6133 | 2.6133 | |
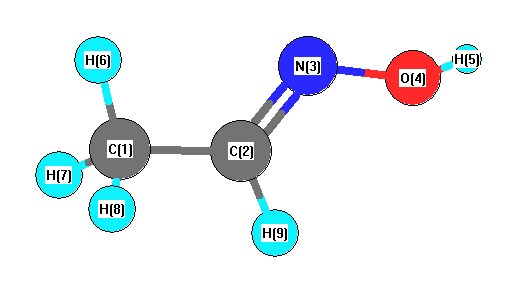 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
120.002 |
|
C1 |
C2 |
H9 |
119.998 |
| C2 |
C1 |
H6 |
110.382 |
|
C2 |
C1 |
H7 |
110.545 |
| C2 |
C1 |
H8 |
110.545 |
|
C2 |
N3 |
O4 |
111.406 |
| N3 |
C2 |
H9 |
120.000 |
|
N3 |
O4 |
H5 |
103.113 |
| H6 |
C1 |
H7 |
109.106 |
|
H6 |
C1 |
H8 |
109.106 |
| H7 |
C1 |
H8 |
107.081 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.166 |
|
|
|
| 2 |
C |
0.762 |
|
|
|
| 3 |
N |
-0.221 |
|
|
|
| 4 |
O |
-0.410 |
|
|
|
| 5 |
H |
0.137 |
|
|
|
| 6 |
H |
-0.097 |
|
|
|
| 7 |
H |
-0.028 |
|
|
|
| 8 |
H |
-0.028 |
|
|
|
| 9 |
H |
-0.281 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.377 |
0.711 |
0.000 |
0.805 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.454 |
-2.104 |
0.000 |
| y |
-2.104 |
-19.032 |
0.000 |
| z |
0.000 |
0.000 |
-25.386 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.245 |
-2.104 |
0.000 |
| y |
-2.104 |
6.388 |
0.000 |
| z |
0.000 |
0.000 |
-3.143 |
|
| Polar |
|---|
| 3z2-r2 | -6.286 |
|---|
| x2-y2 | -6.422 |
|---|
| xy | -2.104 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.388 |
-1.425 |
0.000 |
| y |
-1.425 |
7.692 |
0.000 |
| z |
0.000 |
0.000 |
4.460 |
<r2> (average value of r
2) Å
2
| <r2> |
91.476 |
| (<r2>)1/2 |
9.564 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.113700 |
| Energy at 298.15K | |
| HF Energy | -209.113700 |
| Nuclear repulsion energy | 120.067562 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3881 |
3719 |
97.07 |
|
|
|
| 2 |
A' |
3202 |
3068 |
7.50 |
|
|
|
| 3 |
A' |
3183 |
3051 |
3.82 |
|
|
|
| 4 |
A' |
3061 |
2933 |
12.65 |
|
|
|
| 5 |
A' |
1770 |
1696 |
9.37 |
|
|
|
| 6 |
A' |
1456 |
1396 |
22.01 |
|
|
|
| 7 |
A' |
1393 |
1335 |
35.49 |
|
|
|
| 8 |
A' |
1377 |
1320 |
26.31 |
|
|
|
| 9 |
A' |
1340 |
1284 |
27.10 |
|
|
|
| 10 |
A' |
1146 |
1099 |
8.20 |
|
|
|
| 11 |
A' |
968 |
927 |
149.81 |
|
|
|
| 12 |
A' |
930 |
891 |
2.88 |
|
|
|
| 13 |
A' |
681 |
653 |
12.13 |
|
|
|
| 14 |
A' |
310 |
297 |
1.50 |
|
|
|
| 15 |
A" |
3125 |
2994 |
10.16 |
|
|
|
| 16 |
A" |
1458 |
1398 |
9.59 |
|
|
|
| 17 |
A" |
1051 |
1008 |
0.77 |
|
|
|
| 18 |
A" |
864 |
828 |
15.40 |
|
|
|
| 19 |
A" |
513 |
492 |
44.05 |
|
|
|
| 20 |
A" |
410 |
393 |
72.96 |
|
|
|
| 21 |
A" |
24i |
23i |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16048.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15379.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.445 |
0.490 |
0.000 |
| C2 |
0.000 |
0.857 |
0.000 |
| N3 |
1.013 |
0.082 |
0.000 |
| O4 |
0.657 |
-1.263 |
0.000 |
| H5 |
1.511 |
-1.706 |
0.000 |
| H6 |
-1.586 |
-0.593 |
0.000 |
| H7 |
-1.939 |
0.918 |
0.883 |
| H8 |
-1.939 |
0.918 |
-0.883 |
| H9 |
0.277 |
1.914 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4913 | 2.4920 | 2.7373 | 3.6819 | 1.0920 | 1.0987 | 1.0987 | 2.2347 |
C2 | 1.4913 | | 1.2755 | 2.2198 | 2.9747 | 2.1492 | 2.1316 | 2.1316 | 1.0922 | N3 | 2.4920 | 1.2755 | | 1.3918 | 1.8557 | 2.6855 | 3.1930 | 3.1930 | 1.9738 | O4 | 2.7373 | 2.2198 | 1.3918 | | 0.9613 | 2.3411 | 3.5043 | 3.5043 | 3.1995 | H5 | 3.6819 | 2.9747 | 1.8557 | 0.9613 | | 3.2903 | 4.4233 | 4.4233 | 3.8236 | H6 | 1.0920 | 2.1492 | 2.6855 | 2.3411 | 3.2903 | | 1.7859 | 1.7859 | 3.1233 | H7 | 1.0987 | 2.1316 | 3.1930 | 3.5043 | 4.4233 | 1.7859 | | 1.7655 | 2.5848 | H8 | 1.0987 | 2.1316 | 3.1930 | 3.5043 | 4.4233 | 1.7859 | 1.7655 | | 2.5848 | H9 | 2.2347 | 1.0922 | 1.9738 | 3.1995 | 3.8236 | 3.1233 | 2.5848 | 2.5848 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.329 |
|
C1 |
C2 |
H9 |
118.954 |
| C2 |
C1 |
H6 |
111.668 |
|
C2 |
C1 |
H7 |
109.847 |
| C2 |
C1 |
H8 |
109.847 |
|
C2 |
N3 |
O4 |
112.586 |
| N3 |
C2 |
H9 |
112.717 |
|
N3 |
O4 |
H5 |
102.572 |
| H6 |
C1 |
H7 |
109.215 |
|
H6 |
C1 |
H8 |
109.215 |
| H7 |
C1 |
H8 |
106.928 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.310 |
|
|
|
| 2 |
C |
0.599 |
|
|
|
| 3 |
N |
-0.141 |
|
|
|
| 4 |
O |
-0.448 |
|
|
|
| 5 |
H |
0.144 |
|
|
|
| 6 |
H |
-0.098 |
|
|
|
| 7 |
H |
-0.038 |
|
|
|
| 8 |
H |
-0.038 |
|
|
|
| 9 |
H |
-0.289 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.851 |
0.238 |
0.000 |
0.883 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.323 |
-3.862 |
0.000 |
| y |
-3.862 |
-21.329 |
0.000 |
| z |
0.000 |
0.000 |
-25.412 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.048 |
-3.862 |
0.000 |
| y |
-3.862 |
3.039 |
0.000 |
| z |
0.000 |
0.000 |
-3.086 |
|
| Polar |
|---|
| 3z2-r2 | -6.172 |
|---|
| x2-y2 | -1.994 |
|---|
| xy | -3.862 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.828 |
-1.124 |
0.000 |
| y |
-1.124 |
6.794 |
0.000 |
| z |
0.000 |
0.000 |
4.458 |
<r2> (average value of r
2) Å
2
| <r2> |
78.983 |
| (<r2>)1/2 |
8.887 |
