Jump to
S1C2
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3738 |
3652 |
|
|
|
|
| 2 |
A |
3166 |
3093 |
|
|
|
|
| 3 |
A |
3133 |
3061 |
|
|
|
|
| 4 |
A |
3060 |
2989 |
|
|
|
|
| 5 |
A |
1702 |
1662 |
|
|
|
|
| 6 |
A |
1529 |
1494 |
|
|
|
|
| 7 |
A |
1472 |
1438 |
|
|
|
|
| 8 |
A |
1434 |
1401 |
|
|
|
|
| 9 |
A |
1300 |
1270 |
|
|
|
|
| 10 |
A |
1172 |
1145 |
|
|
|
|
| 11 |
A |
1011 |
988 |
|
|
|
|
| 12 |
A |
910 |
889 |
|
|
|
|
| 13 |
A |
561 |
548 |
|
|
|
|
| 14 |
A |
321 |
314 |
|
|
|
|
| 15 |
A |
3128 |
3055 |
|
|
|
|
| 16 |
A |
1523 |
1488 |
|
|
|
|
| 17 |
A |
1096 |
1071 |
|
|
|
|
| 18 |
A |
901 |
880 |
|
|
|
|
| 19 |
A |
392 |
383 |
|
|
|
|
| 20 |
A |
278 |
272 |
|
|
|
|
| 21 |
A |
184 |
180 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16005.9 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 15636.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.645945 |
| Energy at 298.15K | |
| HF Energy | -207.915318 |
| Nuclear repulsion energy | 118.341151 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3739 |
3652 |
|
|
|
|
| 2 |
A' |
3197 |
3123 |
|
|
|
|
| 3 |
A' |
3176 |
3102 |
|
|
|
|
| 4 |
A' |
3065 |
2995 |
|
|
|
|
| 5 |
A' |
1714 |
1674 |
|
|
|
|
| 6 |
A' |
1526 |
1491 |
|
|
|
|
| 7 |
A' |
1447 |
1414 |
|
|
|
|
| 8 |
A' |
1402 |
1370 |
|
|
|
|
| 9 |
A' |
1354 |
1322 |
|
|
|
|
| 10 |
A' |
1168 |
1141 |
|
|
|
|
| 11 |
A' |
943 |
921 |
|
|
|
|
| 12 |
A' |
904 |
883 |
|
|
|
|
| 13 |
A' |
669 |
654 |
|
|
|
|
| 14 |
A' |
317 |
309 |
|
|
|
|
| 15 |
A" |
3129 |
3056 |
|
|
|
|
| 16 |
A" |
1534 |
1499 |
|
|
|
|
| 17 |
A" |
1087 |
1062 |
|
|
|
|
| 18 |
A" |
858 |
838 |
|
|
|
|
| 19 |
A" |
486 |
475 |
|
|
|
|
| 20 |
A" |
378 |
370 |
|
|
|
|
| 21 |
A" |
82i |
80i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16005.4 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 15635.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.459 |
0.486 |
0.000 |
| C2 |
0.000 |
0.875 |
0.000 |
| N3 |
1.039 |
0.096 |
0.000 |
| O4 |
0.649 |
-1.283 |
0.000 |
| H5 |
1.512 |
-1.723 |
0.000 |
| H6 |
-1.584 |
-0.607 |
0.000 |
| H7 |
-1.957 |
0.910 |
0.891 |
| H8 |
-1.957 |
0.910 |
-0.891 |
| H9 |
0.276 |
1.938 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5099 | 2.5280 | 2.7513 | 3.7015 | 1.0994 | 1.1055 | 1.1055 | 2.2624 |
C2 | 1.5099 | | 1.2983 | 2.2532 | 3.0056 | 2.1686 | 2.1502 | 2.1502 | 1.0984 | N3 | 2.5280 | 1.2983 | | 1.4335 | 1.8797 | 2.7152 | 3.2296 | 3.2296 | 1.9937 | O4 | 2.7513 | 2.2532 | 1.4335 | | 0.9686 | 2.3325 | 3.5202 | 3.5202 | 3.2426 | H5 | 3.7015 | 3.0056 | 1.8797 | 0.9686 | | 3.2903 | 4.4448 | 4.4448 | 3.8641 | H6 | 1.0994 | 2.1686 | 2.7152 | 2.3325 | 3.2903 | | 1.7984 | 1.7984 | 3.1516 | H7 | 1.1055 | 2.1502 | 3.2296 | 3.5202 | 4.4448 | 1.7984 | | 1.7820 | 2.6141 | H8 | 1.1055 | 2.1502 | 3.2296 | 3.5202 | 4.4448 | 1.7984 | 1.7820 | | 2.6141 | H9 | 2.2624 | 1.0984 | 1.9937 | 3.2426 | 3.8641 | 3.1516 | 2.6141 | 2.6141 | |
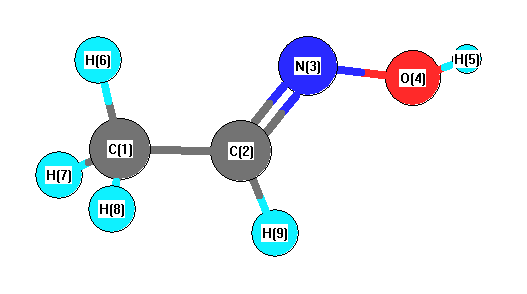 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.217 |
|
C1 |
C2 |
H9 |
119.476 |
| C2 |
C1 |
H6 |
111.456 |
|
C2 |
C1 |
H7 |
109.633 |
| C2 |
C1 |
H8 |
109.633 |
|
C2 |
N3 |
O4 |
111.041 |
| N3 |
C2 |
H9 |
112.308 |
|
N3 |
O4 |
H5 |
101.220 |
| H6 |
C1 |
H7 |
109.308 |
|
H6 |
C1 |
H8 |
109.308 |
| H7 |
C1 |
H8 |
107.414 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
