Jump to
S1C2
Vibrational Frequencies calculated at M06-2X/aug-cc-pVDZ
Geometric Data calculated at M06-2X/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at M06-2X/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.076150 |
| Energy at 298.15K | |
| HF Energy | -209.076150 |
| Nuclear repulsion energy | 120.179937 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3887 |
3887 |
110.45 |
|
|
|
| 2 |
A' |
3211 |
3211 |
9.02 |
|
|
|
| 3 |
A' |
3191 |
3191 |
0.01 |
|
|
|
| 4 |
A' |
3059 |
3059 |
9.69 |
|
|
|
| 5 |
A' |
1796 |
1796 |
9.57 |
|
|
|
| 6 |
A' |
1454 |
1454 |
29.73 |
|
|
|
| 7 |
A' |
1402 |
1402 |
44.51 |
|
|
|
| 8 |
A' |
1382 |
1382 |
21.26 |
|
|
|
| 9 |
A' |
1341 |
1341 |
17.69 |
|
|
|
| 10 |
A' |
1150 |
1150 |
8.35 |
|
|
|
| 11 |
A' |
991 |
991 |
131.09 |
|
|
|
| 12 |
A' |
929 |
929 |
6.32 |
|
|
|
| 13 |
A' |
684 |
684 |
11.74 |
|
|
|
| 14 |
A' |
333 |
333 |
1.52 |
|
|
|
| 15 |
A" |
3121 |
3121 |
7.69 |
|
|
|
| 16 |
A" |
1454 |
1454 |
10.13 |
|
|
|
| 17 |
A" |
1058 |
1058 |
1.10 |
|
|
|
| 18 |
A" |
876 |
876 |
14.54 |
|
|
|
| 19 |
A" |
517 |
517 |
58.20 |
|
|
|
| 20 |
A" |
427 |
427 |
63.38 |
|
|
|
| 21 |
A" |
114i |
114i |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16073.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16073.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.446 |
0.474 |
0.000 |
| C2 |
0.000 |
0.861 |
0.000 |
| N3 |
1.014 |
0.090 |
0.000 |
| O4 |
0.656 |
-1.255 |
0.000 |
| H5 |
1.507 |
-1.704 |
0.000 |
| H6 |
-1.567 |
-0.611 |
0.000 |
| H7 |
-1.940 |
0.897 |
0.884 |
| H8 |
-1.940 |
0.897 |
-0.884 |
| H9 |
0.268 |
1.919 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4966 | 2.4893 | 2.7211 | 3.6687 | 1.0915 | 1.0977 | 1.0977 | 2.2417 |
C2 | 1.4966 | | 1.2736 | 2.2150 | 2.9747 | 2.1495 | 2.1319 | 2.1319 | 1.0918 | N3 | 2.4893 | 1.2736 | | 1.3917 | 1.8604 | 2.6740 | 3.1869 | 3.1869 | 1.9757 | O4 | 2.7211 | 2.2150 | 1.3917 | | 0.9622 | 2.3138 | 3.4857 | 3.4857 | 3.1978 | H5 | 3.6687 | 2.9747 | 1.8604 | 0.9622 | | 3.2619 | 4.4075 | 4.4075 | 3.8293 | H6 | 1.0915 | 2.1495 | 2.6740 | 2.3138 | 3.2619 | | 1.7877 | 1.7877 | 3.1252 | H7 | 1.0977 | 2.1319 | 3.1869 | 3.4857 | 4.4075 | 1.7877 | | 1.7680 | 2.5880 | H8 | 1.0977 | 2.1319 | 3.1869 | 3.4857 | 4.4075 | 1.7877 | 1.7680 | | 2.5880 | H9 | 2.2417 | 1.0918 | 1.9757 | 3.1978 | 3.8293 | 3.1252 | 2.5880 | 2.5880 | |
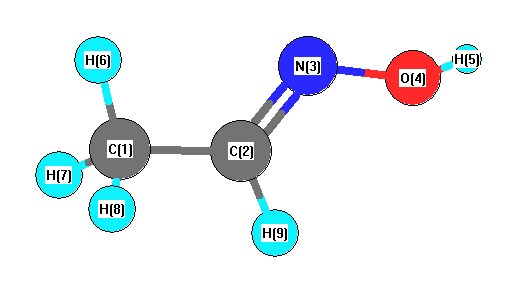 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.764 |
|
C1 |
C2 |
H9 |
119.182 |
| C2 |
C1 |
H6 |
111.352 |
|
C2 |
C1 |
H7 |
109.565 |
| C2 |
C1 |
H8 |
109.565 |
|
C2 |
N3 |
O4 |
112.337 |
| N3 |
C2 |
H9 |
113.054 |
|
N3 |
O4 |
H5 |
102.916 |
| H6 |
C1 |
H7 |
109.494 |
|
H6 |
C1 |
H8 |
109.494 |
| H7 |
C1 |
H8 |
107.281 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.602 |
|
|
|
| 2 |
C |
0.652 |
|
|
|
| 3 |
N |
-0.064 |
|
|
|
| 4 |
O |
-0.398 |
|
|
|
| 5 |
H |
0.130 |
|
|
|
| 6 |
H |
-0.231 |
|
|
|
| 7 |
H |
-0.139 |
|
|
|
| 8 |
H |
-0.139 |
|
|
|
| 9 |
H |
-0.411 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.825 |
0.256 |
0.000 |
0.864 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.391 |
-3.863 |
0.000 |
| y |
-3.863 |
-21.410 |
0.000 |
| z |
0.000 |
0.000 |
-25.563 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.095 |
-3.863 |
0.000 |
| y |
-3.863 |
3.067 |
0.000 |
| z |
0.000 |
0.000 |
-3.162 |
|
| Polar |
|---|
| 3z2-r2 | -6.324 |
|---|
| x2-y2 | -1.981 |
|---|
| xy | -3.863 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.691 |
-1.073 |
0.000 |
| y |
-1.073 |
6.679 |
0.000 |
| z |
0.000 |
0.000 |
4.427 |
<r2> (average value of r
2) Å
2
| <r2> |
78.726 |
| (<r2>)1/2 |
8.873 |
