Jump to
S1C2
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -208.628111 |
| Energy at 298.15K | |
| HF Energy | -207.917960 |
| Nuclear repulsion energy | 119.023503 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3853 |
3853 |
75.42 |
|
|
|
| 2 |
A' |
3198 |
3198 |
12.39 |
|
|
|
| 3 |
A' |
3182 |
3182 |
0.07 |
|
|
|
| 4 |
A' |
3054 |
3054 |
14.82 |
|
|
|
| 5 |
A' |
1755 |
1755 |
6.27 |
|
|
|
| 6 |
A' |
1477 |
1477 |
15.10 |
|
|
|
| 7 |
A' |
1407 |
1407 |
24.18 |
|
|
|
| 8 |
A' |
1390 |
1390 |
29.40 |
|
|
|
| 9 |
A' |
1347 |
1347 |
24.92 |
|
|
|
| 10 |
A' |
1151 |
1151 |
7.48 |
|
|
|
| 11 |
A' |
935 |
935 |
76.34 |
|
|
|
| 12 |
A' |
914 |
914 |
35.85 |
|
|
|
| 13 |
A' |
668 |
668 |
12.63 |
|
|
|
| 14 |
A' |
318 |
318 |
1.52 |
|
|
|
| 15 |
A" |
3117 |
3117 |
13.72 |
|
|
|
| 16 |
A" |
1481 |
1481 |
7.29 |
|
|
|
| 17 |
A" |
1063 |
1063 |
1.36 |
|
|
|
| 18 |
A" |
864 |
864 |
11.93 |
|
|
|
| 19 |
A" |
486 |
486 |
34.37 |
|
|
|
| 20 |
A" |
386 |
386 |
83.05 |
|
|
|
| 21 |
A" |
51i |
51i |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15996.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15996.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.457 |
0.480 |
0.000 |
| C2 |
0.000 |
0.868 |
0.000 |
| N3 |
1.029 |
0.095 |
0.000 |
| O4 |
0.655 |
-1.271 |
0.000 |
| H5 |
1.512 |
-1.714 |
0.000 |
| H6 |
-1.585 |
-0.609 |
0.000 |
| H7 |
-1.952 |
0.906 |
0.889 |
| H8 |
-1.952 |
0.906 |
-0.889 |
| H9 |
0.268 |
1.930 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5073 | 2.5158 | 2.7438 | 3.6914 | 1.0967 | 1.1031 | 1.1031 | 2.2528 |
C2 | 1.5073 | | 1.2872 | 2.2371 | 2.9916 | 2.1665 | 2.1451 | 2.1451 | 1.0954 | N3 | 2.5158 | 1.2872 | | 1.4163 | 1.8719 | 2.7078 | 3.2149 | 3.2149 | 1.9868 | O4 | 2.7438 | 2.2371 | 1.4163 | | 0.9641 | 2.3366 | 3.5110 | 3.5110 | 3.2245 | H5 | 3.6914 | 2.9916 | 1.8719 | 0.9641 | | 3.2885 | 4.4329 | 4.4329 | 3.8502 | H6 | 1.0967 | 2.1665 | 2.7078 | 2.3366 | 3.2885 | | 1.7942 | 1.7942 | 3.1431 | H7 | 1.1031 | 2.1451 | 3.2149 | 3.5110 | 4.4329 | 1.7942 | | 1.7783 | 2.6011 | H8 | 1.1031 | 2.1451 | 3.2149 | 3.5110 | 4.4329 | 1.7942 | 1.7783 | | 2.6011 | H9 | 2.2528 | 1.0954 | 1.9868 | 3.2245 | 3.8502 | 3.1431 | 2.6011 | 2.6011 | |
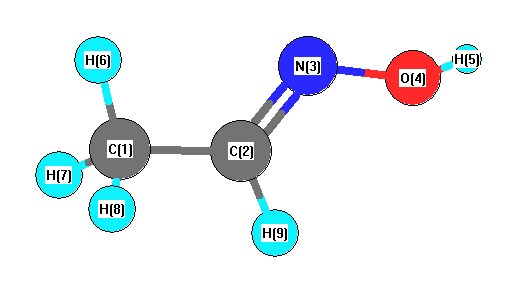 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.207 |
|
C1 |
C2 |
H9 |
119.040 |
| C2 |
C1 |
H6 |
111.632 |
|
C2 |
C1 |
H7 |
109.550 |
| C2 |
C1 |
H8 |
109.550 |
|
C2 |
N3 |
O4 |
111.588 |
| N3 |
C2 |
H9 |
112.752 |
|
N3 |
O4 |
H5 |
102.011 |
| H6 |
C1 |
H7 |
109.294 |
|
H6 |
C1 |
H8 |
109.294 |
| H7 |
C1 |
H8 |
107.426 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
