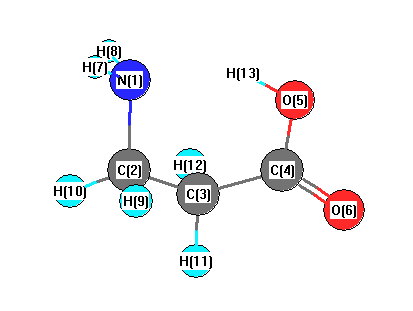
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3904 |
3555 |
457.00 |
|
|
|
| 2 |
A |
3810 |
3469 |
9.78 |
|
|
|
| 3 |
A |
3725 |
3392 |
6.20 |
|
|
|
| 4 |
A |
3275 |
2982 |
13.41 |
|
|
|
| 5 |
A |
3227 |
2938 |
34.62 |
|
|
|
| 6 |
A |
3177 |
2893 |
12.03 |
|
|
|
| 7 |
A |
3168 |
2885 |
57.78 |
|
|
|
| 8 |
A |
1995 |
1817 |
509.19 |
|
|
|
| 9 |
A |
1776 |
1617 |
36.47 |
|
|
|
| 10 |
A |
1622 |
1477 |
4.07 |
|
|
|
| 11 |
A |
1570 |
1429 |
106.89 |
|
|
|
| 12 |
A |
1563 |
1423 |
282.58 |
|
|
|
| 13 |
A |
1535 |
1398 |
7.64 |
|
|
|
| 14 |
A |
1484 |
1352 |
4.75 |
|
|
|
| 15 |
A |
1419 |
1292 |
4.16 |
|
|
|
| 16 |
A |
1399 |
1274 |
6.87 |
|
|
|
| 17 |
A |
1346 |
1225 |
53.89 |
|
|
|
| 18 |
A |
1238 |
1127 |
6.91 |
|
|
|
| 19 |
A |
1166 |
1061 |
14.06 |
|
|
|
| 20 |
A |
1100 |
1002 |
16.99 |
|
|
|
| 21 |
A |
1043 |
950 |
15.27 |
|
|
|
| 22 |
A |
984 |
896 |
55.39 |
|
|
|
| 23 |
A |
937 |
854 |
58.70 |
|
|
|
| 24 |
A |
893 |
813 |
31.41 |
|
|
|
| 25 |
A |
855 |
779 |
61.65 |
|
|
|
| 26 |
A |
760 |
692 |
18.50 |
|
|
|
| 27 |
A |
619 |
564 |
7.20 |
|
|
|
| 28 |
A |
520 |
474 |
10.77 |
|
|
|
| 29 |
A |
413 |
376 |
15.83 |
|
|
|
| 30 |
A |
339 |
308 |
9.78 |
|
|
|
| 31 |
A |
268 |
244 |
3.25 |
|
|
|
| 32 |
A |
200 |
183 |
5.32 |
|
|
|
| 33 |
A |
83 |
76 |
1.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25705.7 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 23407.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.564 |
|
|
|
| 2 |
C |
0.790 |
|
|
|
| 3 |
C |
0.195 |
|
|
|
| 4 |
C |
0.942 |
|
|
|
| 5 |
O |
-0.666 |
|
|
|
| 6 |
O |
-0.696 |
|
|
|
| 7 |
H |
0.012 |
|
|
|
| 8 |
H |
0.057 |
|
|
|
| 9 |
H |
-0.109 |
|
|
|
| 10 |
H |
-0.206 |
|
|
|
| 11 |
H |
-0.060 |
|
|
|
| 12 |
H |
0.009 |
|
|
|
| 13 |
H |
0.297 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
6.519 |
-1.211 |
0.629 |
6.660 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.097 |
0.994 |
-0.640 |
| y |
0.994 |
-37.700 |
-0.074 |
| z |
-0.640 |
-0.074 |
-33.836 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.329 |
0.994 |
-0.640 |
| y |
0.994 |
-0.234 |
-0.074 |
| z |
-0.640 |
-0.074 |
5.563 |
|
| Polar |
|---|
| 3z2-r2 | 11.125 |
|---|
| x2-y2 | -3.397 |
|---|
| xy | 0.994 |
|---|
| xz | -0.640 |
|---|
| yz | -0.074 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.628 |
0.378 |
0.110 |
| y |
0.378 |
7.263 |
0.023 |
| z |
0.110 |
0.023 |
6.100 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |