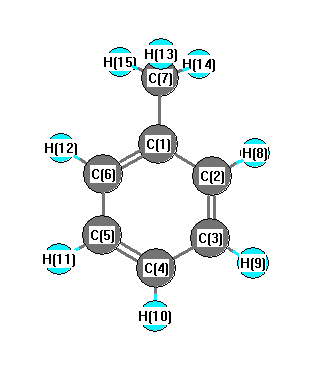
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3227 |
3093 |
12.51 |
|
|
|
| 2 |
A |
3215 |
3081 |
31.01 |
|
|
|
| 3 |
A |
3206 |
3072 |
5.58 |
|
|
|
| 4 |
A |
3193 |
3060 |
6.06 |
|
|
|
| 5 |
A |
3191 |
3058 |
7.93 |
|
|
|
| 6 |
A |
3152 |
3020 |
14.69 |
|
|
|
| 7 |
A |
3122 |
2991 |
13.71 |
|
|
|
| 8 |
A |
3053 |
2926 |
28.69 |
|
|
|
| 9 |
A |
1678 |
1608 |
9.90 |
|
|
|
| 10 |
A |
1654 |
1585 |
0.24 |
|
|
|
| 11 |
A |
1532 |
1468 |
14.39 |
|
|
|
| 12 |
A |
1494 |
1432 |
12.41 |
|
|
|
| 13 |
A |
1464 |
1403 |
6.48 |
|
|
|
| 14 |
A |
1460 |
1399 |
1.12 |
|
|
|
| 15 |
A |
1396 |
1338 |
1.25 |
|
|
|
| 16 |
A |
1377 |
1320 |
0.27 |
|
|
|
| 17 |
A |
1336 |
1280 |
0.25 |
|
|
|
| 18 |
A |
1245 |
1193 |
0.45 |
|
|
|
| 19 |
A |
1199 |
1149 |
0.36 |
|
|
|
| 20 |
A |
1174 |
1125 |
0.03 |
|
|
|
| 21 |
A |
1113 |
1067 |
6.78 |
|
|
|
| 22 |
A |
1060 |
1016 |
3.86 |
|
|
|
| 23 |
A |
1050 |
1006 |
3.23 |
|
|
|
| 24 |
A |
1015 |
973 |
0.64 |
|
|
|
| 25 |
A |
995 |
954 |
2.13 |
|
|
|
| 26 |
A |
994 |
952 |
0.02 |
|
|
|
| 27 |
A |
983 |
942 |
0.00 |
|
|
|
| 28 |
A |
908 |
870 |
0.36 |
|
|
|
| 29 |
A |
854 |
819 |
0.00 |
|
|
|
| 30 |
A |
806 |
772 |
0.39 |
|
|
|
| 31 |
A |
738 |
707 |
56.76 |
|
|
|
| 32 |
A |
708 |
679 |
21.02 |
|
|
|
| 33 |
A |
631 |
604 |
0.09 |
|
|
|
| 34 |
A |
526 |
504 |
0.40 |
|
|
|
| 35 |
A |
477 |
457 |
9.59 |
|
|
|
| 36 |
A |
413 |
396 |
0.01 |
|
|
|
| 37 |
A |
342 |
328 |
0.46 |
|
|
|
| 38 |
A |
212 |
203 |
2.08 |
|
|
|
| 39 |
A |
56 |
54 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28123.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 26950.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.665 |
|
|
|
| 2 |
C |
0.855 |
|
|
|
| 3 |
C |
0.266 |
|
|
|
| 4 |
C |
0.400 |
|
|
|
| 5 |
C |
0.298 |
|
|
|
| 6 |
C |
0.690 |
|
|
|
| 7 |
C |
0.261 |
|
|
|
| 8 |
H |
-0.728 |
|
|
|
| 9 |
H |
-0.586 |
|
|
|
| 10 |
H |
-0.565 |
|
|
|
| 11 |
H |
-0.594 |
|
|
|
| 12 |
H |
-0.743 |
|
|
|
| 13 |
H |
0.026 |
|
|
|
| 14 |
H |
-0.270 |
|
|
|
| 15 |
H |
0.025 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.416 |
-0.039 |
0.001 |
0.418 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.232 |
0.063 |
0.002 |
| y |
0.063 |
-38.616 |
-0.000 |
| z |
0.002 |
-0.000 |
-46.090 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.121 |
0.063 |
0.002 |
| y |
0.063 |
3.545 |
-0.000 |
| z |
0.002 |
-0.000 |
-7.666 |
|
| Polar |
|---|
| 3z2-r2 | -15.332 |
|---|
| x2-y2 | 0.384 |
|---|
| xy | 0.063 |
|---|
| xz | 0.002 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.411 |
0.067 |
0.000 |
| y |
0.067 |
13.318 |
0.000 |
| z |
0.000 |
0.000 |
7.959 |
<r2> (average value of r
2) Å
2
| <r2> |
201.183 |
| (<r2>)1/2 |
14.184 |