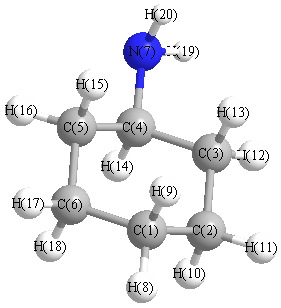
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3791 |
3452 |
0.83 |
|
|
|
| 2 |
A |
3709 |
3377 |
0.66 |
|
|
|
| 3 |
A |
3219 |
2931 |
71.40 |
|
|
|
| 4 |
A |
3205 |
2918 |
124.91 |
|
|
|
| 5 |
A |
3199 |
2913 |
77.67 |
|
|
|
| 6 |
A |
3197 |
2911 |
70.84 |
|
|
|
| 7 |
A |
3186 |
2901 |
86.67 |
|
|
|
| 8 |
A |
3156 |
2873 |
84.10 |
|
|
|
| 9 |
A |
3149 |
2868 |
12.14 |
|
|
|
| 10 |
A |
3148 |
2866 |
15.45 |
|
|
|
| 11 |
A |
3147 |
2866 |
14.05 |
|
|
|
| 12 |
A |
3137 |
2857 |
27.34 |
|
|
|
| 13 |
A |
3133 |
2853 |
12.54 |
|
|
|
| 14 |
A |
1777 |
1619 |
35.56 |
|
|
|
| 15 |
A |
1618 |
1474 |
1.58 |
|
|
|
| 16 |
A |
1600 |
1457 |
5.90 |
|
|
|
| 17 |
A |
1598 |
1455 |
5.35 |
|
|
|
| 18 |
A |
1594 |
1451 |
2.52 |
|
|
|
| 19 |
A |
1587 |
1445 |
0.10 |
|
|
|
| 20 |
A |
1545 |
1407 |
7.68 |
|
|
|
| 21 |
A |
1506 |
1371 |
4.42 |
|
|
|
| 22 |
A |
1501 |
1366 |
0.60 |
|
|
|
| 23 |
A |
1496 |
1362 |
1.14 |
|
|
|
| 24 |
A |
1479 |
1346 |
0.65 |
|
|
|
| 25 |
A |
1468 |
1337 |
1.80 |
|
|
|
| 26 |
A |
1436 |
1308 |
0.72 |
|
|
|
| 27 |
A |
1396 |
1271 |
0.73 |
|
|
|
| 28 |
A |
1379 |
1256 |
2.23 |
|
|
|
| 29 |
A |
1378 |
1255 |
1.08 |
|
|
|
| 30 |
A |
1318 |
1200 |
4.55 |
|
|
|
| 31 |
A |
1289 |
1174 |
1.58 |
|
|
|
| 32 |
A |
1216 |
1107 |
21.01 |
|
|
|
| 33 |
A |
1203 |
1096 |
4.76 |
|
|
|
| 34 |
A |
1160 |
1056 |
0.46 |
|
|
|
| 35 |
A |
1146 |
1044 |
1.24 |
|
|
|
| 36 |
A |
1119 |
1019 |
6.67 |
|
|
|
| 37 |
A |
1103 |
1005 |
0.47 |
|
|
|
| 38 |
A |
1056 |
962 |
3.03 |
|
|
|
| 39 |
A |
998 |
909 |
11.87 |
|
|
|
| 40 |
A |
971 |
884 |
51.79 |
|
|
|
| 41 |
A |
947 |
862 |
4.66 |
|
|
|
| 42 |
A |
921 |
838 |
88.16 |
|
|
|
| 43 |
A |
904 |
823 |
5.69 |
|
|
|
| 44 |
A |
847 |
771 |
0.40 |
|
|
|
| 45 |
A |
836 |
761 |
3.77 |
|
|
|
| 46 |
A |
589 |
536 |
0.97 |
|
|
|
| 47 |
A |
491 |
447 |
0.99 |
|
|
|
| 48 |
A |
482 |
439 |
1.70 |
|
|
|
| 49 |
A |
436 |
397 |
0.23 |
|
|
|
| 50 |
A |
368 |
335 |
10.99 |
|
|
|
| 51 |
A |
355 |
324 |
0.71 |
|
|
|
| 52 |
A |
267 |
243 |
30.77 |
|
|
|
| 53 |
A |
240 |
218 |
9.03 |
|
|
|
| 54 |
A |
169 |
154 |
0.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43581.1 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 39685.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.332 |
|
|
|
| 2 |
C |
0.327 |
|
|
|
| 3 |
C |
0.514 |
|
|
|
| 4 |
C |
-0.134 |
|
|
|
| 5 |
C |
0.547 |
|
|
|
| 6 |
C |
0.319 |
|
|
|
| 7 |
N |
-0.341 |
|
|
|
| 8 |
H |
-0.142 |
|
|
|
| 9 |
H |
-0.131 |
|
|
|
| 10 |
H |
-0.117 |
|
|
|
| 11 |
H |
-0.132 |
|
|
|
| 12 |
H |
-0.103 |
|
|
|
| 13 |
H |
-0.121 |
|
|
|
| 14 |
H |
-0.275 |
|
|
|
| 15 |
H |
-0.081 |
|
|
|
| 16 |
H |
-0.121 |
|
|
|
| 17 |
H |
-0.143 |
|
|
|
| 18 |
H |
-0.125 |
|
|
|
| 19 |
H |
-0.061 |
|
|
|
| 20 |
H |
-0.010 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.640 |
1.076 |
0.432 |
1.324 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.889 |
-3.913 |
-1.504 |
| y |
-3.913 |
-46.994 |
-0.731 |
| z |
-1.504 |
-0.731 |
-44.681 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.051 |
-3.913 |
-1.504 |
| y |
-3.913 |
-0.209 |
-0.731 |
| z |
-1.504 |
-0.731 |
3.260 |
|
| Polar |
|---|
| 3z2-r2 | 6.520 |
|---|
| x2-y2 | -1.894 |
|---|
| xy | -3.913 |
|---|
| xz | -1.504 |
|---|
| yz | -0.731 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.254 |
-0.034 |
-0.087 |
| y |
-0.034 |
11.300 |
-0.005 |
| z |
-0.087 |
-0.005 |
9.956 |
<r2> (average value of r
2) Å
2
| <r2> |
232.874 |
| (<r2>)1/2 |
15.260 |