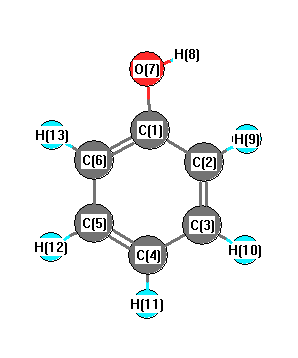
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3877 |
3716 |
63.85 |
|
|
|
| 2 |
A' |
3235 |
3100 |
4.70 |
|
|
|
| 3 |
A' |
3229 |
3094 |
13.76 |
|
|
|
| 4 |
A' |
3215 |
3081 |
14.36 |
|
|
|
| 5 |
A' |
3207 |
3073 |
0.18 |
|
|
|
| 6 |
A' |
3188 |
3055 |
12.67 |
|
|
|
| 7 |
A' |
1678 |
1608 |
40.59 |
|
|
|
| 8 |
A' |
1668 |
1598 |
52.38 |
|
|
|
| 9 |
A' |
1534 |
1470 |
66.18 |
|
|
|
| 10 |
A' |
1504 |
1441 |
27.70 |
|
|
|
| 11 |
A' |
1395 |
1337 |
16.18 |
|
|
|
| 12 |
A' |
1353 |
1296 |
17.54 |
|
|
|
| 13 |
A' |
1298 |
1244 |
86.88 |
|
|
|
| 14 |
A' |
1201 |
1151 |
122.11 |
|
|
|
| 15 |
A' |
1186 |
1137 |
8.24 |
|
|
|
| 16 |
A' |
1171 |
1122 |
5.59 |
|
|
|
| 17 |
A' |
1098 |
1052 |
12.21 |
|
|
|
| 18 |
A' |
1054 |
1010 |
5.77 |
|
|
|
| 19 |
A' |
1010 |
968 |
0.37 |
|
|
|
| 20 |
A' |
835 |
800 |
18.88 |
|
|
|
| 21 |
A' |
627 |
601 |
0.37 |
|
|
|
| 22 |
A' |
532 |
510 |
1.52 |
|
|
|
| 23 |
A' |
404 |
387 |
9.84 |
|
|
|
| 24 |
A" |
989 |
947 |
0.19 |
|
|
|
| 25 |
A" |
973 |
933 |
0.03 |
|
|
|
| 26 |
A" |
900 |
862 |
7.20 |
|
|
|
| 27 |
A" |
829 |
794 |
0.02 |
|
|
|
| 28 |
A" |
767 |
735 |
52.69 |
|
|
|
| 29 |
A" |
704 |
675 |
32.70 |
|
|
|
| 30 |
A" |
517 |
495 |
16.06 |
|
|
|
| 31 |
A" |
420 |
403 |
0.32 |
|
|
|
| 32 |
A" |
370 |
355 |
96.38 |
|
|
|
| 33 |
A" |
230 |
221 |
0.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23097.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 22134.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.852 |
|
|
|
| 2 |
C |
0.420 |
|
|
|
| 3 |
C |
0.670 |
|
|
|
| 4 |
C |
0.302 |
|
|
|
| 5 |
C |
0.867 |
|
|
|
| 6 |
C |
0.405 |
|
|
|
| 7 |
O |
-0.691 |
|
|
|
| 8 |
H |
0.180 |
|
|
|
| 9 |
H |
-0.746 |
|
|
|
| 10 |
H |
-0.568 |
|
|
|
| 11 |
H |
-0.558 |
|
|
|
| 12 |
H |
-0.572 |
|
|
|
| 13 |
H |
-0.561 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.273 |
-0.008 |
0.000 |
1.273 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.360 |
-4.186 |
0.000 |
| y |
-4.186 |
-37.759 |
0.000 |
| z |
0.000 |
0.000 |
-45.151 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.095 |
-4.186 |
0.000 |
| y |
-4.186 |
2.496 |
0.000 |
| z |
0.000 |
0.000 |
-8.591 |
|
| Polar |
|---|
| 3z2-r2 | -17.182 |
|---|
| x2-y2 | 2.399 |
|---|
| xy | -4.186 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.452 |
-0.209 |
0.000 |
| y |
-0.209 |
13.746 |
0.000 |
| z |
0.000 |
0.000 |
6.885 |
<r2> (average value of r
2) Å
2
| <r2> |
186.323 |
| (<r2>)1/2 |
13.650 |