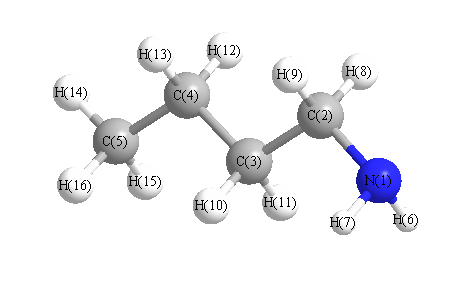
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3715 |
3383 |
1.11 |
|
|
|
| 2 |
A' |
3227 |
2938 |
64.82 |
|
|
|
| 3 |
A' |
3174 |
2890 |
72.02 |
|
|
|
| 4 |
A' |
3159 |
2877 |
28.81 |
|
|
|
| 5 |
A' |
3152 |
2870 |
45.26 |
|
|
|
| 6 |
A' |
3138 |
2857 |
26.40 |
|
|
|
| 7 |
A' |
1781 |
1622 |
28.32 |
|
|
|
| 8 |
A' |
1611 |
1467 |
4.04 |
|
|
|
| 9 |
A' |
1601 |
1458 |
0.84 |
|
|
|
| 10 |
A' |
1589 |
1447 |
0.61 |
|
|
|
| 11 |
A' |
1587 |
1445 |
0.37 |
|
|
|
| 12 |
A' |
1534 |
1396 |
9.80 |
|
|
|
| 13 |
A' |
1520 |
1384 |
1.96 |
|
|
|
| 14 |
A' |
1484 |
1352 |
3.37 |
|
|
|
| 15 |
A' |
1381 |
1258 |
6.85 |
|
|
|
| 16 |
A' |
1222 |
1113 |
6.48 |
|
|
|
| 17 |
A' |
1169 |
1064 |
28.39 |
|
|
|
| 18 |
A' |
1130 |
1029 |
2.19 |
|
|
|
| 19 |
A' |
1081 |
985 |
15.07 |
|
|
|
| 20 |
A' |
972 |
885 |
31.75 |
|
|
|
| 21 |
A' |
917 |
835 |
142.03 |
|
|
|
| 22 |
A' |
458 |
417 |
4.76 |
|
|
|
| 23 |
A' |
417 |
380 |
0.34 |
|
|
|
| 24 |
A' |
197 |
180 |
2.17 |
|
|
|
| 25 |
A" |
3796 |
3457 |
1.28 |
|
|
|
| 26 |
A" |
3225 |
2937 |
150.52 |
|
|
|
| 27 |
A" |
3213 |
2926 |
30.07 |
|
|
|
| 28 |
A" |
3184 |
2899 |
10.99 |
|
|
|
| 29 |
A" |
3161 |
2878 |
4.04 |
|
|
|
| 30 |
A" |
1593 |
1451 |
6.04 |
|
|
|
| 31 |
A" |
1490 |
1357 |
0.25 |
|
|
|
| 32 |
A" |
1424 |
1297 |
0.26 |
|
|
|
| 33 |
A" |
1401 |
1276 |
0.05 |
|
|
|
| 34 |
A" |
1320 |
1202 |
0.00 |
|
|
|
| 35 |
A" |
1113 |
1013 |
0.46 |
|
|
|
| 36 |
A" |
980 |
892 |
0.03 |
|
|
|
| 37 |
A" |
837 |
762 |
0.28 |
|
|
|
| 38 |
A" |
778 |
709 |
1.87 |
|
|
|
| 39 |
A" |
286 |
260 |
44.58 |
|
|
|
| 40 |
A" |
267 |
243 |
0.97 |
|
|
|
| 41 |
A" |
128 |
117 |
0.00 |
|
|
|
| 42 |
A" |
114 |
104 |
2.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34762.9 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 31655.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.416 |
|
|
|
| 2 |
C |
0.876 |
|
|
|
| 3 |
C |
0.079 |
|
|
|
| 4 |
C |
0.265 |
|
|
|
| 5 |
C |
0.424 |
|
|
|
| 6 |
H |
-0.015 |
|
|
|
| 7 |
H |
-0.015 |
|
|
|
| 8 |
H |
-0.163 |
|
|
|
| 9 |
H |
-0.163 |
|
|
|
| 10 |
H |
-0.158 |
|
|
|
| 11 |
H |
-0.158 |
|
|
|
| 12 |
H |
-0.162 |
|
|
|
| 13 |
H |
-0.162 |
|
|
|
| 14 |
H |
-0.072 |
|
|
|
| 15 |
H |
-0.080 |
|
|
|
| 16 |
H |
-0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.178 |
0.737 |
0.000 |
1.389 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.772 |
2.894 |
0.000 |
| y |
2.894 |
-33.429 |
0.000 |
| z |
0.000 |
0.000 |
-32.357 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.880 |
2.894 |
0.000 |
| y |
2.894 |
3.636 |
0.000 |
| z |
0.000 |
0.000 |
5.243 |
|
| Polar |
|---|
| 3z2-r2 | 10.486 |
|---|
| x2-y2 | -8.344 |
|---|
| xy | 2.894 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.004 |
0.182 |
0.000 |
| y |
0.182 |
8.220 |
0.000 |
| z |
0.000 |
0.000 |
7.691 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |