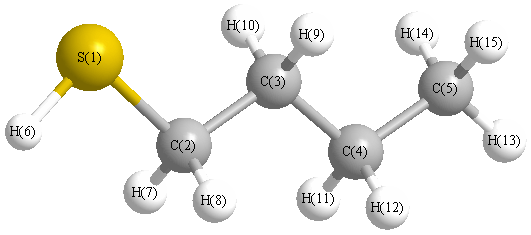
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3139 |
3008 |
34.53 |
|
|
|
| 2 |
A' |
3080 |
2951 |
34.97 |
|
|
|
| 3 |
A' |
3055 |
2927 |
49.21 |
|
|
|
| 4 |
A' |
3052 |
2925 |
12.27 |
|
|
|
| 5 |
A' |
3045 |
2918 |
7.60 |
|
|
|
| 6 |
A' |
2712 |
2599 |
4.24 |
|
|
|
| 7 |
A' |
1489 |
1427 |
7.40 |
|
|
|
| 8 |
A' |
1476 |
1415 |
0.88 |
|
|
|
| 9 |
A' |
1471 |
1409 |
1.64 |
|
|
|
| 10 |
A' |
1465 |
1404 |
1.59 |
|
|
|
| 11 |
A' |
1401 |
1342 |
4.10 |
|
|
|
| 12 |
A' |
1391 |
1333 |
3.18 |
|
|
|
| 13 |
A' |
1326 |
1270 |
6.62 |
|
|
|
| 14 |
A' |
1237 |
1185 |
18.13 |
|
|
|
| 15 |
A' |
1140 |
1092 |
1.49 |
|
|
|
| 16 |
A' |
1093 |
1048 |
0.29 |
|
|
|
| 17 |
A' |
1056 |
1012 |
0.21 |
|
|
|
| 18 |
A' |
930 |
891 |
1.91 |
|
|
|
| 19 |
A' |
854 |
818 |
1.21 |
|
|
|
| 20 |
A' |
758 |
727 |
4.27 |
|
|
|
| 21 |
A' |
392 |
375 |
0.68 |
|
|
|
| 22 |
A' |
323 |
309 |
0.80 |
|
|
|
| 23 |
A' |
154 |
148 |
1.27 |
|
|
|
| 24 |
A" |
3143 |
3012 |
26.92 |
|
|
|
| 25 |
A" |
3132 |
3002 |
38.39 |
|
|
|
| 26 |
A" |
3101 |
2972 |
16.27 |
|
|
|
| 27 |
A" |
3078 |
2950 |
1.56 |
|
|
|
| 28 |
A" |
1478 |
1416 |
7.70 |
|
|
|
| 29 |
A" |
1323 |
1268 |
0.38 |
|
|
|
| 30 |
A" |
1298 |
1244 |
0.52 |
|
|
|
| 31 |
A" |
1219 |
1168 |
0.52 |
|
|
|
| 32 |
A" |
1067 |
1023 |
0.91 |
|
|
|
| 33 |
A" |
921 |
882 |
1.43 |
|
|
|
| 34 |
A" |
786 |
754 |
0.00 |
|
|
|
| 35 |
A" |
733 |
702 |
3.77 |
|
|
|
| 36 |
A" |
246 |
235 |
0.01 |
|
|
|
| 37 |
A" |
179 |
171 |
12.89 |
|
|
|
| 38 |
A" |
109 |
104 |
0.68 |
|
|
|
| 39 |
A" |
92 |
88 |
3.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28971.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 27763.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.178 |
|
|
|
| 2 |
C |
0.257 |
|
|
|
| 3 |
C |
0.532 |
|
|
|
| 4 |
C |
0.534 |
|
|
|
| 5 |
C |
0.407 |
|
|
|
| 6 |
H |
0.096 |
|
|
|
| 7 |
H |
-0.243 |
|
|
|
| 8 |
H |
-0.243 |
|
|
|
| 9 |
H |
-0.294 |
|
|
|
| 10 |
H |
-0.294 |
|
|
|
| 11 |
H |
-0.283 |
|
|
|
| 12 |
H |
-0.283 |
|
|
|
| 13 |
H |
-0.128 |
|
|
|
| 14 |
H |
-0.117 |
|
|
|
| 15 |
H |
-0.117 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.772 |
0.360 |
0.000 |
1.809 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.699 |
2.214 |
0.000 |
| y |
2.214 |
-38.931 |
0.000 |
| z |
0.000 |
0.000 |
-42.059 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.204 |
2.214 |
0.000 |
| y |
2.214 |
3.948 |
0.000 |
| z |
0.000 |
0.000 |
-0.744 |
|
| Polar |
|---|
| 3z2-r2 | -1.488 |
|---|
| x2-y2 | -4.768 |
|---|
| xy | 2.214 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.976 |
-1.650 |
0.000 |
| y |
-1.650 |
12.788 |
0.000 |
| z |
0.000 |
0.000 |
9.330 |
<r2> (average value of r
2) Å
2
| <r2> |
262.679 |
| (<r2>)1/2 |
16.207 |