return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9SH (1-Butanethiol)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-555.696777
Energy at 298.15K 
HF Energy-554.846084
Nuclear repulsion energy221.343779
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3136 3064        
2 A' 3074 3003        
3 A' 3054 2983        
4 A' 3046 2976        
5 A' 3038 2968        
6 A' 2717 2654        
7 A' 1558 1522        
8 A' 1545 1509        
9 A' 1538 1502        
10 A' 1535 1499        
11 A' 1458 1425        
12 A' 1439 1406        
13 A' 1377 1345        
14 A' 1290 1260        
15 A' 1161 1134        
16 A' 1104 1078        
17 A' 1065 1041        
18 A' 950 928        
19 A' 872 852        
20 A' 773 755        
21 A' 397 388        
22 A' 328 321        
23 A' 155 151        
24 A" 3134 3062        
25 A" 3131 3059        
26 A" 3095 3024        
27 A" 3074 3003        
28 A" 1546 1510        
29 A" 1358 1326        
30 A" 1339 1308        
31 A" 1268 1239        
32 A" 1114 1088        
33 A" 953 931        
34 A" 813 794        
35 A" 752 734        
36 A" 255 249        
37 A" 177 173        
38 A" 113 111        
39 A" 98 96        

Unscaled Zero Point Vibrational Energy (zpe) 29414.4 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 28734.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.52065 0.04388 0.04175

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.395 -1.862 0.000
C2 -0.241 -0.991 0.000
C3 0.000 0.526 0.000
C4 -1.321 1.317 0.000
C5 -1.087 2.839 0.000
H6 0.902 -3.125 0.000
H7 -0.807 -1.290 0.899
H8 -0.807 -1.290 -0.899
H9 0.599 0.802 -0.890
H10 0.599 0.802 0.890
H11 -1.918 1.032 0.889
H12 -1.918 1.032 -0.889
H13 -2.043 3.393 0.000
H14 -0.515 3.148 0.894
H15 -0.515 3.148 -0.894

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.85382.76574.18185.31631.35542.44712.44712.92002.92004.48834.48836.27995.43635.4363
C21.85381.53592.54843.92232.42061.10401.10402.17132.17132.77402.77404.73974.24374.2437
C32.76571.53591.54012.55593.76032.18102.18101.10821.10822.17402.17403.52052.81822.8182
C44.18182.54841.54011.53954.96722.80512.80512.17832.17831.10811.10812.19752.19162.1916
C55.31633.92232.55591.53956.28674.23464.23462.79012.79012.17842.17841.10471.10601.1060
H61.35542.42063.76034.96726.28672.66432.66434.03814.03815.10115.10117.15206.49306.4930
H72.44711.10402.18102.80514.23462.66431.79903.09162.52072.57373.13394.92554.44774.7957
H82.44711.10402.18102.80514.23462.66431.79902.52073.09163.13392.57374.92554.79574.4477
H92.92002.17131.10822.17832.79014.03813.09162.52071.78083.09122.52783.80593.15092.5970
H102.92002.17131.10822.17832.79014.03812.52073.09161.78082.52783.09123.80592.59703.1509
H114.48832.77402.17401.10812.17845.10112.57373.13393.09122.52781.77762.52572.53983.1030
H124.48832.77402.17401.10812.17845.10113.13392.57372.52783.09121.77762.52573.10302.5398
H136.27994.73973.52052.19751.10477.15204.92554.92553.80593.80592.52572.52571.78771.7877
H145.43634.24372.81822.19161.10606.49304.44774.79573.15092.59702.53983.10301.78771.7879
H155.43634.24372.81822.19161.10606.49304.79574.44772.59703.15093.10302.53981.78771.7879

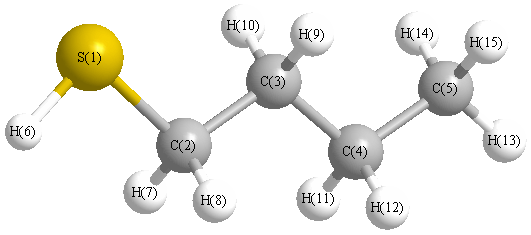
picture of 1-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C3 108.999 S1 C2 H7 109.002
S1 C2 H8 109.002 C2 S1 H6 96.693
C2 C3 C4 111.888 C2 C3 H9 109.346
C2 C3 H10 109.346 C3 C2 H7 110.343
C3 C2 H8 110.343 C3 C4 C5 112.187
C3 C4 H11 109.267 C3 C4 H12 109.267
C4 C3 H9 109.603 C4 C3 H10 109.603
C4 C5 H13 111.354 C4 C5 H14 110.814
C4 C5 H15 110.814 C5 C4 H11 109.654
C5 C4 H12 109.654 H7 C2 H8 109.124
H9 C3 H10 106.926 H11 C4 H12 106.655
H13 C5 H14 107.928 H13 C5 H15 107.928
H14 C5 H15 107.862
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability