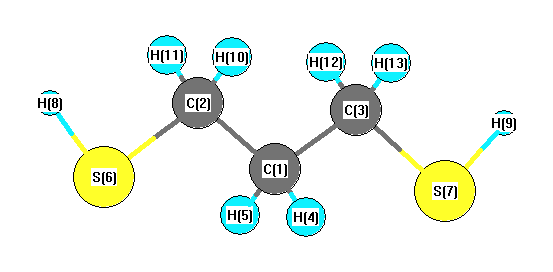
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3086 |
2957 |
46.10 |
|
|
|
| 2 |
A1 |
3067 |
2939 |
0.33 |
|
|
|
| 3 |
A1 |
2713 |
2600 |
0.95 |
|
|
|
| 4 |
A1 |
1483 |
1421 |
4.83 |
|
|
|
| 5 |
A1 |
1470 |
1409 |
0.85 |
|
|
|
| 6 |
A1 |
1283 |
1229 |
20.28 |
|
|
|
| 7 |
A1 |
1109 |
1063 |
0.69 |
|
|
|
| 8 |
A1 |
858 |
822 |
0.16 |
|
|
|
| 9 |
A1 |
796 |
763 |
0.28 |
|
|
|
| 10 |
A1 |
300 |
287 |
0.26 |
|
|
|
| 11 |
A1 |
127 |
122 |
0.29 |
|
|
|
| 12 |
A2 |
3139 |
3009 |
0.00 |
|
|
|
| 13 |
A2 |
1317 |
1263 |
0.00 |
|
|
|
| 14 |
A2 |
1101 |
1055 |
0.00 |
|
|
|
| 15 |
A2 |
830 |
795 |
0.00 |
|
|
|
| 16 |
A2 |
188 |
180 |
0.00 |
|
|
|
| 17 |
A2 |
89 |
85 |
0.00 |
|
|
|
| 18 |
B1 |
3152 |
3020 |
20.61 |
|
|
|
| 19 |
B1 |
3110 |
2981 |
0.98 |
|
|
|
| 20 |
B1 |
1267 |
1214 |
2.13 |
|
|
|
| 21 |
B1 |
1011 |
969 |
3.16 |
|
|
|
| 22 |
B1 |
746 |
715 |
3.14 |
|
|
|
| 23 |
B1 |
181 |
173 |
31.45 |
|
|
|
| 24 |
B1 |
97 |
93 |
0.11 |
|
|
|
| 25 |
B2 |
3078 |
2950 |
4.83 |
|
|
|
| 26 |
B2 |
2713 |
2600 |
4.82 |
|
|
|
| 27 |
B2 |
1465 |
1404 |
2.69 |
|
|
|
| 28 |
B2 |
1372 |
1315 |
16.83 |
|
|
|
| 29 |
B2 |
1217 |
1167 |
12.29 |
|
|
|
| 30 |
B2 |
1082 |
1037 |
0.02 |
|
|
|
| 31 |
B2 |
891 |
854 |
0.23 |
|
|
|
| 32 |
B2 |
720 |
690 |
5.37 |
|
|
|
| 33 |
B2 |
330 |
316 |
6.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22693.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 21746.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.574 |
|
|
|
| 2 |
C |
0.178 |
|
|
|
| 3 |
C |
0.178 |
|
|
|
| 4 |
H |
-0.237 |
|
|
|
| 5 |
H |
-0.237 |
|
|
|
| 6 |
S |
0.163 |
|
|
|
| 7 |
S |
0.163 |
|
|
|
| 8 |
H |
0.105 |
|
|
|
| 9 |
H |
0.105 |
|
|
|
| 10 |
H |
-0.248 |
|
|
|
| 11 |
H |
-0.248 |
|
|
|
| 12 |
H |
-0.248 |
|
|
|
| 13 |
H |
-0.248 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.768 |
2.768 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.694 |
0.000 |
0.000 |
| y |
0.000 |
-48.530 |
0.000 |
| z |
0.000 |
0.000 |
-46.266 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.296 |
0.000 |
0.000 |
| y |
0.000 |
-0.550 |
0.000 |
| z |
0.000 |
0.000 |
2.846 |
|
| Polar |
|---|
| 3z2-r2 | 5.692 |
|---|
| x2-y2 | -1.164 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.073 |
0.000 |
0.000 |
| y |
0.000 |
16.012 |
0.000 |
| z |
0.000 |
0.000 |
10.851 |
<r2> (average value of r
2) Å
2
| <r2> |
345.911 |
| (<r2>)1/2 |
18.599 |