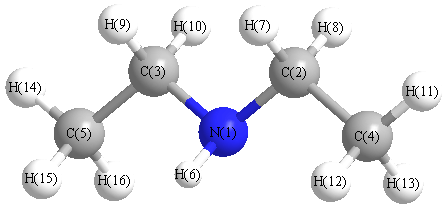
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3735 |
3401 |
0.13 |
|
|
|
| 2 |
A' |
3246 |
2955 |
82.52 |
|
|
|
| 3 |
A' |
3226 |
2937 |
120.72 |
|
|
|
| 4 |
A' |
3191 |
2906 |
37.64 |
|
|
|
| 5 |
A' |
3162 |
2879 |
23.64 |
|
|
|
| 6 |
A' |
3090 |
2814 |
185.94 |
|
|
|
| 7 |
A' |
1630 |
1484 |
1.33 |
|
|
|
| 8 |
A' |
1594 |
1452 |
3.13 |
|
|
|
| 9 |
A' |
1585 |
1443 |
11.07 |
|
|
|
| 10 |
A' |
1550 |
1411 |
1.47 |
|
|
|
| 11 |
A' |
1499 |
1365 |
0.56 |
|
|
|
| 12 |
A' |
1420 |
1293 |
5.70 |
|
|
|
| 13 |
A' |
1321 |
1203 |
2.28 |
|
|
|
| 14 |
A' |
1260 |
1147 |
18.05 |
|
|
|
| 15 |
A' |
1133 |
1031 |
6.61 |
|
|
|
| 16 |
A' |
958 |
873 |
3.22 |
|
|
|
| 17 |
A' |
884 |
805 |
5.46 |
|
|
|
| 18 |
A' |
824 |
750 |
70.86 |
|
|
|
| 19 |
A' |
450 |
410 |
0.36 |
|
|
|
| 20 |
A' |
276 |
252 |
1.41 |
|
|
|
| 21 |
A' |
200 |
183 |
0.89 |
|
|
|
| 22 |
A' |
120 |
110 |
1.55 |
|
|
|
| 23 |
A" |
3245 |
2955 |
11.54 |
|
|
|
| 24 |
A" |
3224 |
2936 |
33.64 |
|
|
|
| 25 |
A" |
3184 |
2899 |
0.98 |
|
|
|
| 26 |
A" |
3161 |
2878 |
45.31 |
|
|
|
| 27 |
A" |
3081 |
2806 |
7.59 |
|
|
|
| 28 |
A" |
1634 |
1488 |
47.13 |
|
|
|
| 29 |
A" |
1618 |
1473 |
0.38 |
|
|
|
| 30 |
A" |
1592 |
1449 |
2.11 |
|
|
|
| 31 |
A" |
1581 |
1440 |
7.22 |
|
|
|
| 32 |
A" |
1519 |
1383 |
10.97 |
|
|
|
| 33 |
A" |
1463 |
1332 |
22.61 |
|
|
|
| 34 |
A" |
1380 |
1257 |
2.39 |
|
|
|
| 35 |
A" |
1250 |
1138 |
53.90 |
|
|
|
| 36 |
A" |
1196 |
1089 |
5.77 |
|
|
|
| 37 |
A" |
1140 |
1038 |
0.57 |
|
|
|
| 38 |
A" |
1004 |
914 |
0.71 |
|
|
|
| 39 |
A" |
857 |
780 |
0.49 |
|
|
|
| 40 |
A" |
456 |
415 |
0.27 |
|
|
|
| 41 |
A" |
276 |
251 |
0.53 |
|
|
|
| 42 |
A" |
113 |
103 |
0.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34662.4 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 31563.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.759 |
|
|
|
| 2 |
C |
0.609 |
|
|
|
| 3 |
C |
0.609 |
|
|
|
| 4 |
C |
0.372 |
|
|
|
| 5 |
C |
0.372 |
|
|
|
| 6 |
H |
0.005 |
|
|
|
| 7 |
H |
-0.181 |
|
|
|
| 8 |
H |
-0.234 |
|
|
|
| 9 |
H |
-0.181 |
|
|
|
| 10 |
H |
-0.234 |
|
|
|
| 11 |
H |
-0.067 |
|
|
|
| 12 |
H |
-0.096 |
|
|
|
| 13 |
H |
-0.026 |
|
|
|
| 14 |
H |
-0.067 |
|
|
|
| 15 |
H |
-0.096 |
|
|
|
| 16 |
H |
-0.026 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.835 |
0.493 |
0.000 |
0.969 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.981 |
1.674 |
0.000 |
| y |
1.674 |
-34.935 |
0.000 |
| z |
0.000 |
0.000 |
-33.826 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.399 |
1.674 |
0.000 |
| y |
1.674 |
-1.031 |
0.000 |
| z |
0.000 |
0.000 |
0.632 |
|
| Polar |
|---|
| 3z2-r2 | 1.264 |
|---|
| x2-y2 | 0.953 |
|---|
| xy | 1.674 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.687 |
-0.041 |
0.000 |
| y |
-0.041 |
8.278 |
0.000 |
| z |
0.000 |
0.000 |
10.188 |
<r2> (average value of r
2) Å
2
| <r2> |
187.384 |
| (<r2>)1/2 |
13.689 |