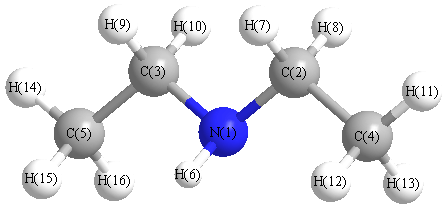
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3533 |
3386 |
0.10 |
|
|
|
| 2 |
A' |
3152 |
3021 |
41.45 |
|
|
|
| 3 |
A' |
3132 |
3002 |
65.64 |
|
|
|
| 4 |
A' |
3070 |
2942 |
51.40 |
|
|
|
| 5 |
A' |
3055 |
2927 |
14.49 |
|
|
|
| 6 |
A' |
2952 |
2829 |
182.04 |
|
|
|
| 7 |
A' |
1508 |
1445 |
2.52 |
|
|
|
| 8 |
A' |
1478 |
1416 |
2.76 |
|
|
|
| 9 |
A' |
1468 |
1407 |
14.38 |
|
|
|
| 10 |
A' |
1421 |
1362 |
2.29 |
|
|
|
| 11 |
A' |
1377 |
1319 |
1.11 |
|
|
|
| 12 |
A' |
1312 |
1257 |
4.05 |
|
|
|
| 13 |
A' |
1225 |
1174 |
1.93 |
|
|
|
| 14 |
A' |
1164 |
1115 |
13.64 |
|
|
|
| 15 |
A' |
1085 |
1040 |
7.95 |
|
|
|
| 16 |
A' |
901 |
863 |
4.33 |
|
|
|
| 17 |
A' |
822 |
788 |
1.52 |
|
|
|
| 18 |
A' |
759 |
727 |
71.45 |
|
|
|
| 19 |
A' |
427 |
410 |
0.26 |
|
|
|
| 20 |
A' |
250 |
239 |
1.11 |
|
|
|
| 21 |
A' |
185 |
178 |
0.90 |
|
|
|
| 22 |
A' |
111 |
106 |
1.19 |
|
|
|
| 23 |
A" |
3152 |
3020 |
11.93 |
|
|
|
| 24 |
A" |
3132 |
3001 |
14.92 |
|
|
|
| 25 |
A" |
3069 |
2941 |
1.02 |
|
|
|
| 26 |
A" |
3054 |
2927 |
33.74 |
|
|
|
| 27 |
A" |
2948 |
2825 |
16.53 |
|
|
|
| 28 |
A" |
1502 |
1439 |
30.56 |
|
|
|
| 29 |
A" |
1495 |
1433 |
0.54 |
|
|
|
| 30 |
A" |
1475 |
1413 |
4.58 |
|
|
|
| 31 |
A" |
1462 |
1401 |
9.00 |
|
|
|
| 32 |
A" |
1394 |
1336 |
15.57 |
|
|
|
| 33 |
A" |
1342 |
1286 |
20.55 |
|
|
|
| 34 |
A" |
1275 |
1222 |
3.73 |
|
|
|
| 35 |
A" |
1190 |
1140 |
47.26 |
|
|
|
| 36 |
A" |
1108 |
1062 |
5.86 |
|
|
|
| 37 |
A" |
1068 |
1023 |
3.69 |
|
|
|
| 38 |
A" |
955 |
915 |
0.13 |
|
|
|
| 39 |
A" |
802 |
769 |
0.43 |
|
|
|
| 40 |
A" |
426 |
409 |
0.41 |
|
|
|
| 41 |
A" |
250 |
240 |
0.62 |
|
|
|
| 42 |
A" |
116 |
111 |
0.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32799.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 31431.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.457 |
|
|
|
| 2 |
C |
0.780 |
|
|
|
| 3 |
C |
0.780 |
|
|
|
| 4 |
C |
0.428 |
|
|
|
| 5 |
C |
0.428 |
|
|
|
| 6 |
H |
0.008 |
|
|
|
| 7 |
H |
-0.290 |
|
|
|
| 8 |
H |
-0.338 |
|
|
|
| 9 |
H |
-0.290 |
|
|
|
| 10 |
H |
-0.338 |
|
|
|
| 11 |
H |
-0.120 |
|
|
|
| 12 |
H |
-0.151 |
|
|
|
| 13 |
H |
-0.084 |
|
|
|
| 14 |
H |
-0.120 |
|
|
|
| 15 |
H |
-0.151 |
|
|
|
| 16 |
H |
-0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.771 |
0.419 |
0.000 |
0.878 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.873 |
1.724 |
0.000 |
| y |
1.724 |
-34.854 |
0.000 |
| z |
0.000 |
0.000 |
-33.477 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.292 |
1.724 |
0.000 |
| y |
1.724 |
-1.179 |
0.000 |
| z |
0.000 |
0.000 |
0.887 |
|
| Polar |
|---|
| 3z2-r2 | 1.774 |
|---|
| x2-y2 | 0.981 |
|---|
| xy | 1.724 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.201 |
-0.147 |
0.000 |
| y |
-0.147 |
8.983 |
0.000 |
| z |
0.000 |
0.000 |
11.145 |
<r2> (average value of r
2) Å
2
| <r2> |
187.122 |
| (<r2>)1/2 |
13.679 |