return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: B2PLYP=FULLultrafine/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
 hartrees
Energy at 0K-213.575201
Energy at 298.15K 
HF Energy-213.311447
Nuclear repulsion energy187.893182
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3498 3367 0.20      
2 A' 3143 3024 45.25      
3 A' 3123 3005 66.71      
4 A' 3069 2954 47.48      
5 A' 3050 2935 14.64      
6 A' 2953 2842 179.77      
7 A' 1516 1459 1.58      
8 A' 1487 1431 2.55      
9 A' 1479 1423 12.81      
10 A' 1422 1368 2.58      
11 A' 1385 1333 0.63      
12 A' 1316 1266 4.45      
13 A' 1226 1180 1.39      
14 A' 1163 1119 11.44      
15 A' 1072 1032 7.66      
16 A' 899 865 4.37      
17 A' 824 793 1.30      
18 A' 763 734 68.73      
19 A' 425 409 0.20      
20 A' 253 243 1.18      
21 A' 186 179 0.84      
22 A' 110 106 1.28      
23 A" 3143 3024 9.80      
24 A" 3123 3005 18.52      
25 A" 3067 2951 0.97      
26 A" 3049 2934 34.67      
27 A" 2950 2838 14.74      
28 A" 1505 1449 5.71      
29 A" 1498 1441 14.68      
30 A" 1484 1428 4.71      
31 A" 1468 1413 14.21      
32 A" 1399 1346 11.61      
33 A" 1346 1296 23.14      
34 A" 1280 1232 3.81      
35 A" 1167 1123 49.26      
36 A" 1111 1069 7.23      
37 A" 1066 1026 2.23      
38 A" 948 913 0.45      
39 A" 804 774 0.41      
40 A" 427 411 0.37      
41 A" 253 243 0.58      
42 A" 110 106 0.90      

Unscaled Zero Point Vibrational Energy (zpe) 32779.3 cm-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 31543.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/aug-cc-pVDZ
ABC
0.59016 0.06965 0.06568

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.285 0.000
C2 0.017 0.518 1.222
C3 0.017 0.518 -1.222
C4 0.017 -0.371 2.461
C5 0.017 -0.371 -2.461
H6 -0.799 -0.896 0.000
H7 -0.841 1.220 1.263
H8 0.928 1.136 1.209
H9 -0.841 1.220 -1.263
H10 0.928 1.136 -1.209
H11 0.037 0.237 3.375
H12 -0.886 -0.997 2.496
H13 0.892 -1.033 2.459
H14 0.037 0.237 -3.375
H15 -0.886 -0.997 -2.496
H16 0.892 -1.033 -2.459

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46211.46212.46262.46261.01932.14412.07582.14412.07583.41542.74852.71533.41542.74852.7153
C21.46212.44411.52493.78882.03891.10951.10052.72142.66852.17142.17612.16854.60584.11534.0893
C31.46212.44413.78881.52492.03892.72142.66851.10951.10054.60584.11534.08932.17142.17612.1685
C42.46261.52493.78884.92212.64532.16862.16044.13994.07051.09821.10011.09755.86795.07745.0410
C52.46263.78881.52494.92212.64534.13994.07052.16862.16045.86795.07745.04101.09821.10011.0975
H61.01932.03892.03892.64532.64532.46472.92732.46472.92733.65692.49952.98763.65692.49952.9876
H72.14411.10952.72142.16864.13992.46471.77122.52663.04082.48912.53773.08414.82194.36494.6838
H82.07581.10052.66852.16044.07052.92731.77123.04082.41802.50873.08192.50394.75564.64424.2617
H92.14412.72141.10954.13992.16862.46472.52663.04081.77124.82194.36494.68382.48912.53773.0841
H102.07582.66851.10054.07052.16042.92733.04082.41801.77124.75564.64424.26172.50873.08192.5039
H113.41542.17144.60581.09825.86793.65692.48912.50874.82194.75561.77471.78506.75036.07016.0320
H122.74852.17614.11531.10015.07742.49952.53773.08194.36494.64421.77471.77936.07014.99185.2646
H132.71532.16854.08931.09755.04102.98763.08412.50394.68384.26171.78501.77936.03205.26464.9181
H143.41544.60582.17145.86791.09823.65694.82194.75562.48912.50876.75036.07016.03201.77471.7850
H152.74854.11532.17615.07741.10012.49954.36494.64422.53773.08196.07014.99185.26461.77471.7793
H162.71534.08932.16855.04101.09752.98764.68384.26173.08412.50396.03205.26464.91811.78501.7793

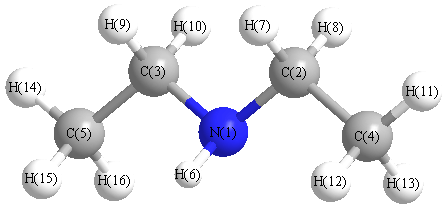
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 111.047 N1 C2 H7 112.249
N1 C2 H8 107.361 N1 C3 C5 111.047
N1 C3 H9 112.249 N1 C3 H10 107.361
C2 N1 C3 113.406 C2 N1 H6 109.207
C2 C4 H11 110.694 C2 C4 H12 110.962
C2 C4 H13 110.509 C3 N1 H6 109.207
C3 C5 H14 110.694 C3 C5 H15 110.962
C3 C5 H16 110.509 C4 C2 H7 109.804
C4 C2 H8 109.693 C5 C3 H8 151.073
C5 C3 H10 109.693 H7 C2 H8 106.535
H9 C3 H10 106.535 H11 C4 H12 107.671
H11 C4 H13 108.776 H12 C4 H13 108.130
H14 C5 H15 107.671 H14 C5 H16 108.776
H15 C5 H16 108.130
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability