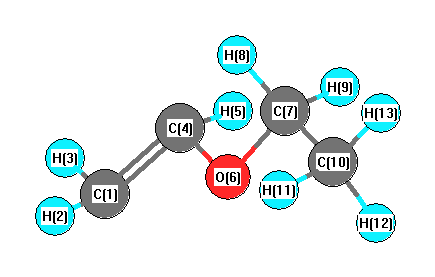
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3302 |
3164 |
4.94 |
|
|
|
| 2 |
A |
3198 |
3064 |
0.07 |
|
|
|
| 3 |
A |
3167 |
3035 |
17.07 |
|
|
|
| 4 |
A |
3161 |
3029 |
24.85 |
|
|
|
| 5 |
A |
3157 |
3025 |
20.64 |
|
|
|
| 6 |
A |
3073 |
2945 |
35.33 |
|
|
|
| 7 |
A |
3070 |
2942 |
16.17 |
|
|
|
| 8 |
A |
3027 |
2901 |
56.15 |
|
|
|
| 9 |
A |
1727 |
1655 |
192.53 |
|
|
|
| 10 |
A |
1502 |
1440 |
5.64 |
|
|
|
| 11 |
A |
1479 |
1418 |
1.94 |
|
|
|
| 12 |
A |
1462 |
1401 |
6.53 |
|
|
|
| 13 |
A |
1436 |
1376 |
2.96 |
|
|
|
| 14 |
A |
1408 |
1349 |
67.21 |
|
|
|
| 15 |
A |
1374 |
1317 |
20.26 |
|
|
|
| 16 |
A |
1339 |
1284 |
5.70 |
|
|
|
| 17 |
A |
1297 |
1243 |
0.64 |
|
|
|
| 18 |
A |
1237 |
1185 |
256.11 |
|
|
|
| 19 |
A |
1172 |
1123 |
4.09 |
|
|
|
| 20 |
A |
1160 |
1112 |
130.26 |
|
|
|
| 21 |
A |
1109 |
1062 |
10.17 |
|
|
|
| 22 |
A |
994 |
952 |
6.85 |
|
|
|
| 23 |
A |
976 |
936 |
22.66 |
|
|
|
| 24 |
A |
863 |
827 |
56.89 |
|
|
|
| 25 |
A |
862 |
826 |
4.26 |
|
|
|
| 26 |
A |
818 |
784 |
1.89 |
|
|
|
| 27 |
A |
707 |
677 |
7.29 |
|
|
|
| 28 |
A |
499 |
478 |
2.73 |
|
|
|
| 29 |
A |
438 |
419 |
1.39 |
|
|
|
| 30 |
A |
246 |
236 |
0.38 |
|
|
|
| 31 |
A |
210 |
201 |
0.21 |
|
|
|
| 32 |
A |
93 |
89 |
2.76 |
|
|
|
| 33 |
A |
43 |
41 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24801.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 23767.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.538 |
|
|
|
| 2 |
H |
-0.479 |
|
|
|
| 3 |
H |
-0.332 |
|
|
|
| 4 |
C |
1.043 |
|
|
|
| 5 |
H |
-0.573 |
|
|
|
| 6 |
O |
-0.645 |
|
|
|
| 7 |
C |
0.951 |
|
|
|
| 8 |
H |
-0.269 |
|
|
|
| 9 |
H |
-0.270 |
|
|
|
| 10 |
C |
0.285 |
|
|
|
| 11 |
H |
-0.089 |
|
|
|
| 12 |
H |
-0.090 |
|
|
|
| 13 |
H |
-0.072 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.267 |
1.251 |
0.001 |
1.780 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.069 |
0.091 |
0.003 |
| y |
0.091 |
-30.749 |
-0.012 |
| z |
0.003 |
-0.012 |
-33.616 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.114 |
0.091 |
0.003 |
| y |
0.091 |
1.093 |
-0.012 |
| z |
0.003 |
-0.012 |
-3.207 |
|
| Polar |
|---|
| 3z2-r2 | -6.414 |
|---|
| x2-y2 | 0.681 |
|---|
| xy | 0.091 |
|---|
| xz | 0.003 |
|---|
| yz | -0.012 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.760 |
0.723 |
-0.001 |
| y |
0.723 |
7.920 |
-0.001 |
| z |
-0.001 |
-0.001 |
6.922 |
<r2> (average value of r
2) Å
2
| <r2> |
162.322 |
| (<r2>)1/2 |
12.741 |