Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -231.165698 |
| Energy at 298.15K | -231.171529 |
| Nuclear repulsion energy | 158.871293 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3308 |
3170 |
3.95 |
|
|
|
| 2 |
A1 |
3204 |
3071 |
2.77 |
|
|
|
| 3 |
A1 |
3187 |
3054 |
16.35 |
|
|
|
| 4 |
A1 |
1756 |
1683 |
2.39 |
|
|
|
| 5 |
A1 |
1424 |
1364 |
0.88 |
|
|
|
| 6 |
A1 |
1348 |
1292 |
4.95 |
|
|
|
| 7 |
A1 |
1216 |
1165 |
14.42 |
|
|
|
| 8 |
A1 |
869 |
833 |
2.89 |
|
|
|
| 9 |
A1 |
523 |
502 |
0.27 |
|
|
|
| 10 |
A1 |
232 |
222 |
0.01 |
|
|
|
| 11 |
A2 |
968 |
927 |
0.00 |
|
|
|
| 12 |
A2 |
877 |
841 |
0.00 |
|
|
|
| 13 |
A2 |
720 |
690 |
0.00 |
|
|
|
| 14 |
A2 |
123 |
118 |
0.00 |
|
|
|
| 15 |
B1 |
975 |
934 |
49.72 |
|
|
|
| 16 |
B1 |
880 |
844 |
111.67 |
|
|
|
| 17 |
B1 |
694 |
665 |
12.49 |
|
|
|
| 18 |
B1 |
89 |
85 |
1.16 |
|
|
|
| 19 |
B2 |
3308 |
3170 |
1.11 |
|
|
|
| 20 |
B2 |
3204 |
3070 |
0.23 |
|
|
|
| 21 |
B2 |
3183 |
3050 |
6.09 |
|
|
|
| 22 |
B2 |
1705 |
1634 |
510.80 |
|
|
|
| 23 |
B2 |
1416 |
1357 |
70.66 |
|
|
|
| 24 |
B2 |
1327 |
1271 |
14.57 |
|
|
|
| 25 |
B2 |
1234 |
1183 |
451.75 |
|
|
|
| 26 |
B2 |
1029 |
986 |
4.85 |
|
|
|
| 27 |
B2 |
488 |
468 |
6.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19641.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18822.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.333 |
| C2 |
0.000 |
1.166 |
-0.371 |
| C3 |
0.000 |
-1.166 |
-0.371 |
| C4 |
0.000 |
2.352 |
0.234 |
| C5 |
0.000 |
-2.352 |
0.234 |
| H6 |
0.000 |
1.054 |
-1.457 |
| H7 |
0.000 |
-1.054 |
-1.457 |
| H8 |
0.000 |
3.255 |
-0.369 |
| H9 |
0.000 |
2.442 |
1.317 |
| H10 |
0.000 |
-3.255 |
-0.369 |
| H11 |
0.000 |
-2.442 |
1.317 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3622 | 1.3622 | 2.3541 | 2.3541 | 2.0773 | 2.0773 | 3.3297 | 2.6329 | 3.3297 | 2.6329 |
C2 | 1.3622 | | 2.3329 | 1.3307 | 3.5700 | 1.0920 | 2.4723 | 2.0883 | 2.1156 | 4.4212 | 3.9837 | C3 | 1.3622 | 2.3329 | | 3.5700 | 1.3307 | 2.4723 | 1.0920 | 4.4212 | 3.9837 | 2.0883 | 2.1156 | C4 | 2.3541 | 1.3307 | 3.5700 | | 4.7041 | 2.1312 | 3.8029 | 1.0857 | 1.0873 | 5.6392 | 4.9149 | C5 | 2.3541 | 3.5700 | 1.3307 | 4.7041 | | 3.8029 | 2.1312 | 5.6392 | 4.9149 | 1.0857 | 1.0873 | H6 | 2.0773 | 1.0920 | 2.4723 | 2.1312 | 3.8029 | | 2.1088 | 2.4544 | 3.1018 | 4.4443 | 4.4633 | H7 | 2.0773 | 2.4723 | 1.0920 | 3.8029 | 2.1312 | 2.1088 | | 4.4443 | 4.4633 | 2.4544 | 3.1018 | H8 | 3.3297 | 2.0883 | 4.4212 | 1.0857 | 5.6392 | 2.4544 | 4.4443 | | 1.8723 | 6.5096 | 5.9412 | H9 | 2.6329 | 2.1156 | 3.9837 | 1.0873 | 4.9149 | 3.1018 | 4.4633 | 1.8723 | | 5.9412 | 4.8839 | H10 | 3.3297 | 4.4212 | 2.0883 | 5.6392 | 1.0857 | 4.4443 | 2.4544 | 6.5096 | 5.9412 | | 1.8723 | H11 | 2.6329 | 3.9837 | 2.1156 | 4.9149 | 1.0873 | 4.4633 | 3.1018 | 5.9412 | 4.8839 | 1.8723 | |
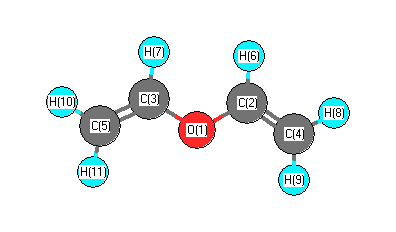 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
121.895 |
|
O1 |
C2 |
H6 |
115.208 |
| O1 |
C3 |
C5 |
121.895 |
|
O1 |
C3 |
H7 |
115.208 |
| C2 |
O1 |
C3 |
117.807 |
|
C2 |
C4 |
H8 |
119.244 |
| C2 |
C4 |
H9 |
121.752 |
|
C3 |
C5 |
H10 |
119.244 |
| C3 |
C5 |
H11 |
121.752 |
|
C4 |
C2 |
H6 |
122.897 |
| C5 |
C3 |
H7 |
122.897 |
|
H8 |
C4 |
H9 |
119.004 |
| H10 |
C5 |
H11 |
119.004 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.596 |
|
|
|
| 2 |
C |
1.008 |
|
|
|
| 3 |
C |
1.008 |
|
|
|
| 4 |
C |
0.618 |
|
|
|
| 5 |
C |
0.618 |
|
|
|
| 6 |
H |
-0.570 |
|
|
|
| 7 |
H |
-0.570 |
|
|
|
| 8 |
H |
-0.306 |
|
|
|
| 9 |
H |
-0.453 |
|
|
|
| 10 |
H |
-0.306 |
|
|
|
| 11 |
H |
-0.453 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.193 |
1.193 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.435 |
0.000 |
0.000 |
| y |
0.000 |
-28.092 |
0.000 |
| z |
0.000 |
0.000 |
-27.740 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.520 |
0.000 |
0.000 |
| y |
0.000 |
2.996 |
0.000 |
| z |
0.000 |
0.000 |
3.524 |
|
| Polar |
|---|
| 3z2-r2 | 7.048 |
|---|
| x2-y2 | -6.344 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.104 |
0.000 |
0.000 |
| y |
0.000 |
13.862 |
0.000 |
| z |
0.000 |
0.000 |
7.478 |
<r2> (average value of r
2) Å
2
| <r2> |
148.022 |
| (<r2>)1/2 |
12.166 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -231.165227 |
| Energy at 298.15K | -231.171308 |
| HF Energy | -231.165227 |
| Nuclear repulsion energy | 161.491084 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3304 |
3166 |
3.59 |
|
|
|
| 2 |
A |
3302 |
3165 |
4.97 |
|
|
|
| 3 |
A |
3237 |
3102 |
6.94 |
|
|
|
| 4 |
A |
3204 |
3070 |
3.74 |
|
|
|
| 5 |
A |
3200 |
3067 |
1.53 |
|
|
|
| 6 |
A |
3188 |
3055 |
5.35 |
|
|
|
| 7 |
A |
1729 |
1657 |
98.78 |
|
|
|
| 8 |
A |
1705 |
1634 |
222.44 |
|
|
|
| 9 |
A |
1430 |
1370 |
1.37 |
|
|
|
| 10 |
A |
1412 |
1353 |
23.59 |
|
|
|
| 11 |
A |
1361 |
1305 |
68.72 |
|
|
|
| 12 |
A |
1322 |
1267 |
4.85 |
|
|
|
| 13 |
A |
1253 |
1201 |
278.83 |
|
|
|
| 14 |
A |
1140 |
1093 |
14.53 |
|
|
|
| 15 |
A |
1013 |
971 |
31.93 |
|
|
|
| 16 |
A |
993 |
952 |
8.53 |
|
|
|
| 17 |
A |
982 |
941 |
42.11 |
|
|
|
| 18 |
A |
900 |
863 |
49.88 |
|
|
|
| 19 |
A |
884 |
847 |
14.65 |
|
|
|
| 20 |
A |
871 |
835 |
55.84 |
|
|
|
| 21 |
A |
735 |
705 |
8.48 |
|
|
|
| 22 |
A |
719 |
689 |
3.97 |
|
|
|
| 23 |
A |
591 |
567 |
2.43 |
|
|
|
| 24 |
A |
454 |
435 |
2.88 |
|
|
|
| 25 |
A |
283 |
271 |
1.04 |
|
|
|
| 26 |
A |
208 |
199 |
8.34 |
|
|
|
| 27 |
A |
93 |
90 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19757.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18933.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.046 |
-0.793 |
0.051 |
| C2 |
1.283 |
-0.495 |
0.006 |
| C3 |
-0.925 |
0.226 |
0.296 |
| C4 |
1.838 |
0.709 |
-0.149 |
| C5 |
-2.160 |
0.214 |
-0.195 |
| H6 |
1.873 |
-1.405 |
0.097 |
| H7 |
-0.547 |
1.003 |
0.963 |
| H8 |
2.921 |
0.784 |
-0.158 |
| H9 |
1.260 |
1.619 |
-0.287 |
| H10 |
-2.855 |
1.000 |
0.088 |
| H11 |
-2.502 |
-0.574 |
-0.862 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3624 | 1.3675 | 2.4169 | 2.3551 | 2.0146 | 2.0758 | 3.3666 | 2.7638 | 3.3329 | 2.6295 |
C2 | 1.3624 | | 2.3405 | 1.3343 | 3.5215 | 1.0884 | 2.5518 | 2.0851 | 2.1347 | 4.4005 | 3.8838 | C3 | 1.3675 | 2.3405 | | 2.8392 | 1.3297 | 3.2445 | 1.0916 | 3.9127 | 2.6563 | 2.0902 | 2.1138 | C4 | 2.4169 | 1.3343 | 2.8392 | | 4.0289 | 2.1284 | 2.6477 | 1.0863 | 1.0870 | 4.7073 | 4.5808 | C5 | 2.3551 | 3.5215 | 1.3297 | 4.0289 | | 4.3562 | 2.1367 | 5.1138 | 3.6994 | 1.0864 | 1.0875 | H6 | 2.0146 | 1.0884 | 3.2445 | 2.1284 | 4.3562 | | 3.5226 | 2.4409 | 3.1097 | 5.3047 | 4.5553 | H7 | 2.0758 | 2.5518 | 1.0916 | 2.6477 | 2.1367 | 3.5226 | | 3.6517 | 2.2826 | 2.4679 | 3.1046 | H8 | 3.3666 | 2.0851 | 3.9127 | 1.0863 | 5.1138 | 2.4409 | 3.6517 | | 1.8634 | 5.7852 | 5.6347 | H9 | 2.7638 | 2.1347 | 2.6563 | 1.0870 | 3.6994 | 3.1097 | 2.2826 | 1.8634 | | 4.1782 | 4.3924 | H10 | 3.3329 | 4.4005 | 2.0902 | 4.7073 | 1.0864 | 5.3047 | 2.4679 | 5.7852 | 4.1782 | | 1.8722 | H11 | 2.6295 | 3.8838 | 2.1138 | 4.5808 | 1.0875 | 4.5553 | 3.1046 | 5.6347 | 4.3924 | 1.8722 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.326 |
|
O1 |
C2 |
H6 |
110.071 |
| O1 |
C3 |
C5 |
121.646 |
|
O1 |
C3 |
H7 |
114.690 |
| C2 |
O1 |
C3 |
118.037 |
|
C2 |
C4 |
H8 |
118.587 |
| C2 |
C4 |
H9 |
123.351 |
|
C3 |
C5 |
H10 |
119.450 |
| C3 |
C5 |
H11 |
121.639 |
|
C4 |
C2 |
H6 |
122.595 |
| C5 |
C3 |
H7 |
123.575 |
|
H8 |
C4 |
H9 |
118.049 |
| H10 |
C5 |
H11 |
118.905 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.550 |
|
|
|
| 2 |
C |
0.874 |
|
|
|
| 3 |
C |
1.030 |
|
|
|
| 4 |
C |
0.636 |
|
|
|
| 5 |
C |
0.344 |
|
|
|
| 6 |
H |
-0.430 |
|
|
|
| 7 |
H |
-0.399 |
|
|
|
| 8 |
H |
-0.360 |
|
|
|
| 9 |
H |
-0.402 |
|
|
|
| 10 |
H |
-0.329 |
|
|
|
| 11 |
H |
-0.414 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.327 |
0.614 |
0.226 |
0.731 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.522 |
-1.593 |
0.560 |
| y |
-1.593 |
-29.690 |
1.007 |
| z |
0.560 |
1.007 |
-32.916 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.781 |
-1.593 |
0.560 |
| y |
-1.593 |
0.529 |
1.007 |
| z |
0.560 |
1.007 |
-4.310 |
|
| Polar |
|---|
| 3z2-r2 | -8.620 |
|---|
| x2-y2 | 2.168 |
|---|
| xy | -1.593 |
|---|
| xz | 0.560 |
|---|
| yz | 1.007 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.876 |
0.487 |
0.387 |
| y |
0.487 |
7.768 |
0.018 |
| z |
0.387 |
0.018 |
6.431 |
<r2> (average value of r
2) Å
2
| <r2> |
130.738 |
| (<r2>)1/2 |
11.434 |
