return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H5N (Pyrrole)

using model chemistry: B2PLYP=FULLultrafine/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
 hartrees
Energy at 0K-209.981768
Energy at 298.15K 
HF Energy-209.736479
Nuclear repulsion energy160.225759
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3684 3545 69.12      
2 A1 3296 3172 0.03      
3 A1 3274 3151 2.39      
4 A1 1499 1442 11.11      
5 A1 1426 1373 2.41      
6 A1 1171 1127 2.30      
7 A1 1092 1051 6.50      
8 A1 1033 994 30.91      
9 A1 893 859 0.21      
10 A2 852 820 0.00      
11 A2 690 664 0.00      
12 A2 623 599 0.00      
13 B1 821 790 0.12      
14 B1 729 701 171.18      
15 B1 639 615 0.01      
16 B1 505 486 67.03      
17 B2 3290 3166 1.76      
18 B2 3263 3140 2.54      
19 B2 1567 1507 1.19      
20 B2 1461 1406 6.60      
21 B2 1303 1254 1.30      
22 B2 1154 1111 2.57      
23 B2 1063 1023 24.99      
24 B2 871 838 1.48      

Unscaled Zero Point Vibrational Energy (zpe) 18098.4 cm-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 17416.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/aug-cc-pVDZ
ABC
0.30287 0.29828 0.15028

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 1.124
H2 0.000 0.000 2.132
C3 0.000 1.129 0.335
C4 0.000 -1.129 0.335
C5 0.000 0.714 -0.988
C6 0.000 -0.714 -0.988
H7 0.000 2.121 0.772
H8 0.000 -2.121 0.772
H9 0.000 1.367 -1.855
H10 0.000 -1.367 -1.855

Atom - Atom Distances (Å)
  N1 H2 C3 C4 C5 C6 H7 H8 H9 H10
N11.00811.37741.37742.22952.22952.14982.14983.27773.2777
H21.00812.12252.12253.20073.20072.51972.51974.21504.2150
C31.37742.12252.25741.38582.26851.08403.27872.20253.3201
C41.37742.12252.25742.26851.38583.27871.08403.32012.2025
C52.22953.20071.38582.26851.42902.25233.33671.08522.2549
C62.22953.20072.26851.38581.42903.33672.25232.25491.0852
H72.14982.51971.08403.27872.25233.33674.24152.73254.3661
H82.14982.51973.27871.08403.33672.25234.24154.36612.7325
H93.27774.21502.20253.32011.08522.25492.73254.36612.7341
H103.27774.21503.32012.20252.25491.08524.36612.73252.7341

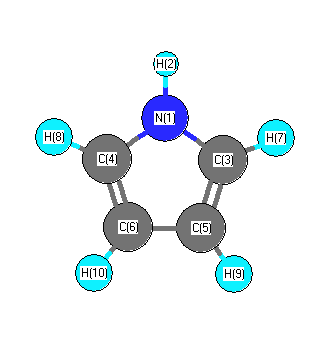
picture of Pyrrole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 107.578 N1 C3 H7 121.260
N1 C4 C6 107.578 N1 C4 H8 121.260
H2 N1 C3 124.970 H2 N1 C4 124.970
C3 N1 C4 110.060 C3 C5 C6 107.392
C3 C5 H9 125.645 C4 C6 C5 107.392
C4 C6 H10 125.645 C5 C3 H7 131.161
C5 C6 H10 126.963 C6 C4 H8 131.161
C6 C5 H9 126.963
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability