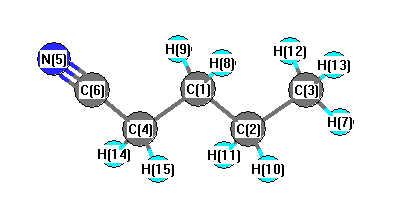
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3143 |
3012 |
30.35 |
|
|
|
| 2 |
A' |
3074 |
2946 |
31.53 |
|
|
|
| 3 |
A' |
3065 |
2937 |
3.57 |
|
|
|
| 4 |
A' |
3054 |
2927 |
23.09 |
|
|
|
| 5 |
A' |
3045 |
2918 |
22.11 |
|
|
|
| 6 |
A' |
2374 |
2275 |
17.85 |
|
|
|
| 7 |
A' |
1490 |
1428 |
9.91 |
|
|
|
| 8 |
A' |
1477 |
1416 |
0.49 |
|
|
|
| 9 |
A' |
1471 |
1409 |
1.18 |
|
|
|
| 10 |
A' |
1449 |
1389 |
3.83 |
|
|
|
| 11 |
A' |
1402 |
1343 |
1.26 |
|
|
|
| 12 |
A' |
1395 |
1337 |
2.85 |
|
|
|
| 13 |
A' |
1343 |
1287 |
1.64 |
|
|
|
| 14 |
A' |
1258 |
1205 |
1.40 |
|
|
|
| 15 |
A' |
1129 |
1082 |
3.39 |
|
|
|
| 16 |
A' |
1089 |
1044 |
0.19 |
|
|
|
| 17 |
A' |
1047 |
1003 |
1.04 |
|
|
|
| 18 |
A' |
953 |
914 |
0.49 |
|
|
|
| 19 |
A' |
913 |
875 |
2.34 |
|
|
|
| 20 |
A' |
544 |
522 |
0.73 |
|
|
|
| 21 |
A' |
386 |
370 |
0.80 |
|
|
|
| 22 |
A' |
280 |
268 |
1.16 |
|
|
|
| 23 |
A' |
125 |
120 |
6.21 |
|
|
|
| 24 |
A" |
3135 |
3005 |
62.23 |
|
|
|
| 25 |
A" |
3128 |
2997 |
2.36 |
|
|
|
| 26 |
A" |
3107 |
2978 |
2.75 |
|
|
|
| 27 |
A" |
3080 |
2952 |
6.79 |
|
|
|
| 28 |
A" |
1478 |
1416 |
8.09 |
|
|
|
| 29 |
A" |
1323 |
1267 |
0.12 |
|
|
|
| 30 |
A" |
1304 |
1250 |
0.39 |
|
|
|
| 31 |
A" |
1230 |
1178 |
0.04 |
|
|
|
| 32 |
A" |
1126 |
1079 |
0.05 |
|
|
|
| 33 |
A" |
925 |
886 |
0.58 |
|
|
|
| 34 |
A" |
785 |
753 |
0.05 |
|
|
|
| 35 |
A" |
728 |
697 |
4.38 |
|
|
|
| 36 |
A" |
385 |
369 |
0.47 |
|
|
|
| 37 |
A" |
240 |
230 |
0.12 |
|
|
|
| 38 |
A" |
112 |
107 |
0.43 |
|
|
|
| 39 |
A" |
83 |
80 |
4.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28837.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 27634.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.448 |
|
|
|
| 2 |
C |
0.677 |
|
|
|
| 3 |
C |
0.428 |
|
|
|
| 4 |
C |
0.802 |
|
|
|
| 5 |
N |
-0.309 |
|
|
|
| 6 |
C |
-0.162 |
|
|
|
| 7 |
H |
-0.121 |
|
|
|
| 8 |
H |
-0.333 |
|
|
|
| 9 |
H |
-0.333 |
|
|
|
| 10 |
H |
-0.285 |
|
|
|
| 11 |
H |
-0.285 |
|
|
|
| 12 |
H |
-0.111 |
|
|
|
| 13 |
H |
-0.111 |
|
|
|
| 14 |
H |
-0.153 |
|
|
|
| 15 |
H |
-0.153 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.956 |
2.004 |
0.000 |
4.435 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.518 |
-8.390 |
0.000 |
| y |
-8.390 |
-42.735 |
0.000 |
| z |
0.000 |
0.000 |
-36.896 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.702 |
-8.390 |
0.000 |
| y |
-8.390 |
-0.028 |
0.000 |
| z |
0.000 |
0.000 |
8.730 |
|
| Polar |
|---|
| 3z2-r2 | 17.461 |
|---|
| x2-y2 | -5.782 |
|---|
| xy | -8.390 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.090 |
1.916 |
0.000 |
| y |
1.916 |
10.624 |
0.000 |
| z |
0.000 |
0.000 |
8.059 |
<r2> (average value of r
2) Å
2
| <r2> |
257.904 |
| (<r2>)1/2 |
16.059 |