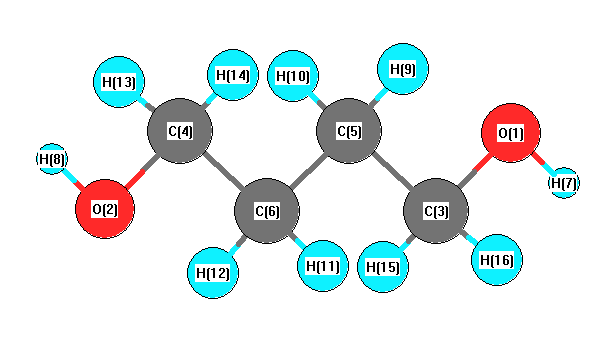
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3894 |
3731 |
0.00 |
|
|
|
| 2 |
Ag |
3055 |
2927 |
0.00 |
|
|
|
| 3 |
Ag |
3007 |
2882 |
0.00 |
|
|
|
| 4 |
Ag |
1504 |
1442 |
0.00 |
|
|
|
| 5 |
Ag |
1478 |
1416 |
0.00 |
|
|
|
| 6 |
Ag |
1451 |
1391 |
0.00 |
|
|
|
| 7 |
Ag |
1376 |
1318 |
0.00 |
|
|
|
| 8 |
Ag |
1270 |
1217 |
0.00 |
|
|
|
| 9 |
Ag |
1097 |
1051 |
0.00 |
|
|
|
| 10 |
Ag |
1075 |
1030 |
0.00 |
|
|
|
| 11 |
Ag |
1055 |
1011 |
0.00 |
|
|
|
| 12 |
Ag |
361 |
346 |
0.00 |
|
|
|
| 13 |
Ag |
341 |
327 |
0.00 |
|
|
|
| 14 |
Au |
3119 |
2989 |
76.64 |
|
|
|
| 15 |
Au |
3044 |
2917 |
68.27 |
|
|
|
| 16 |
Au |
1317 |
1262 |
1.52 |
|
|
|
| 17 |
Au |
1211 |
1160 |
1.73 |
|
|
|
| 18 |
Au |
938 |
898 |
1.03 |
|
|
|
| 19 |
Au |
748 |
717 |
0.79 |
|
|
|
| 20 |
Au |
287 |
275 |
218.85 |
|
|
|
| 21 |
Au |
88 |
84 |
16.15 |
|
|
|
| 22 |
Au |
80 |
76 |
4.49 |
|
|
|
| 23 |
Bg |
3095 |
2966 |
0.00 |
|
|
|
| 24 |
Bg |
3045 |
2918 |
0.00 |
|
|
|
| 25 |
Bg |
1305 |
1251 |
0.00 |
|
|
|
| 26 |
Bg |
1280 |
1227 |
0.00 |
|
|
|
| 27 |
Bg |
1168 |
1120 |
0.00 |
|
|
|
| 28 |
Bg |
810 |
776 |
0.00 |
|
|
|
| 29 |
Bg |
285 |
273 |
0.00 |
|
|
|
| 30 |
Bg |
149 |
143 |
0.00 |
|
|
|
| 31 |
Bu |
3893 |
3731 |
69.23 |
|
|
|
| 32 |
Bu |
3063 |
2935 |
67.55 |
|
|
|
| 33 |
Bu |
3007 |
2881 |
106.06 |
|
|
|
| 34 |
Bu |
1508 |
1445 |
7.61 |
|
|
|
| 35 |
Bu |
1486 |
1424 |
3.52 |
|
|
|
| 36 |
Bu |
1447 |
1387 |
11.26 |
|
|
|
| 37 |
Bu |
1301 |
1246 |
93.98 |
|
|
|
| 38 |
Bu |
1204 |
1153 |
17.23 |
|
|
|
| 39 |
Bu |
1095 |
1049 |
220.52 |
|
|
|
| 40 |
Bu |
1011 |
969 |
0.44 |
|
|
|
| 41 |
Bu |
525 |
503 |
37.99 |
|
|
|
| 42 |
Bu |
140 |
134 |
5.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31304.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29999.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.536 |
|
|
|
| 2 |
O |
-0.536 |
|
|
|
| 3 |
C |
1.002 |
|
|
|
| 4 |
C |
1.002 |
|
|
|
| 5 |
C |
0.451 |
|
|
|
| 6 |
C |
0.451 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
H |
-0.243 |
|
|
|
| 10 |
H |
-0.243 |
|
|
|
| 11 |
H |
-0.243 |
|
|
|
| 12 |
H |
-0.243 |
|
|
|
| 13 |
H |
-0.281 |
|
|
|
| 14 |
H |
-0.281 |
|
|
|
| 15 |
H |
-0.281 |
|
|
|
| 16 |
H |
-0.281 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.549 |
6.108 |
0.000 |
| y |
6.108 |
-45.807 |
0.000 |
| z |
0.000 |
0.000 |
-38.567 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
14.638 |
6.108 |
0.000 |
| y |
6.108 |
-12.749 |
0.000 |
| z |
0.000 |
0.000 |
-1.889 |
|
| Polar |
|---|
| 3z2-r2 | -3.778 |
|---|
| x2-y2 | 18.258 |
|---|
| xy | 6.108 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.587 |
1.193 |
0.000 |
| y |
1.193 |
10.294 |
0.000 |
| z |
0.000 |
0.000 |
7.903 |
<r2> (average value of r
2) Å
2
| <r2> |
284.630 |
| (<r2>)1/2 |
16.871 |