Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -250.221500 |
| Energy at 298.15K | -250.236247 |
| Nuclear repulsion energy | 259.605461 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3767 |
3430 |
0.59 |
|
|
|
| 2 |
A' |
3218 |
2930 |
131.18 |
|
|
|
| 3 |
A' |
3211 |
2924 |
10.87 |
|
|
|
| 4 |
A' |
3199 |
2913 |
89.21 |
|
|
|
| 5 |
A' |
3167 |
2884 |
12.72 |
|
|
|
| 6 |
A' |
3148 |
2866 |
36.81 |
|
|
|
| 7 |
A' |
3075 |
2800 |
155.30 |
|
|
|
| 8 |
A' |
1626 |
1480 |
1.85 |
|
|
|
| 9 |
A' |
1604 |
1461 |
5.71 |
|
|
|
| 10 |
A' |
1589 |
1447 |
5.06 |
|
|
|
| 11 |
A' |
1547 |
1408 |
4.25 |
|
|
|
| 12 |
A' |
1497 |
1363 |
0.17 |
|
|
|
| 13 |
A' |
1413 |
1286 |
1.88 |
|
|
|
| 14 |
A' |
1381 |
1257 |
3.94 |
|
|
|
| 15 |
A' |
1267 |
1154 |
5.98 |
|
|
|
| 16 |
A' |
1144 |
1042 |
13.15 |
|
|
|
| 17 |
A' |
1121 |
1021 |
1.25 |
|
|
|
| 18 |
A' |
966 |
879 |
3.31 |
|
|
|
| 19 |
A' |
940 |
856 |
18.32 |
|
|
|
| 20 |
A' |
874 |
796 |
0.40 |
|
|
|
| 21 |
A' |
844 |
768 |
67.11 |
|
|
|
| 22 |
A' |
584 |
532 |
20.96 |
|
|
|
| 23 |
A' |
463 |
422 |
6.43 |
|
|
|
| 24 |
A' |
425 |
387 |
1.52 |
|
|
|
| 25 |
A' |
259 |
236 |
1.89 |
|
|
|
| 26 |
A" |
3211 |
2924 |
52.99 |
|
|
|
| 27 |
A" |
3206 |
2920 |
94.37 |
|
|
|
| 28 |
A" |
3166 |
2883 |
40.28 |
|
|
|
| 29 |
A" |
3067 |
2793 |
25.80 |
|
|
|
| 30 |
A" |
1625 |
1480 |
4.12 |
|
|
|
| 31 |
A" |
1601 |
1458 |
1.99 |
|
|
|
| 32 |
A" |
1584 |
1442 |
4.16 |
|
|
|
| 33 |
A" |
1500 |
1366 |
0.50 |
|
|
|
| 34 |
A" |
1480 |
1347 |
29.57 |
|
|
|
| 35 |
A" |
1455 |
1325 |
6.02 |
|
|
|
| 36 |
A" |
1389 |
1265 |
2.03 |
|
|
|
| 37 |
A" |
1276 |
1162 |
8.74 |
|
|
|
| 38 |
A" |
1256 |
1144 |
5.84 |
|
|
|
| 39 |
A" |
1236 |
1125 |
10.92 |
|
|
|
| 40 |
A" |
1130 |
1029 |
2.00 |
|
|
|
| 41 |
A" |
1048 |
954 |
0.67 |
|
|
|
| 42 |
A" |
939 |
855 |
0.01 |
|
|
|
| 43 |
A" |
881 |
802 |
0.67 |
|
|
|
| 44 |
A" |
481 |
438 |
0.89 |
|
|
|
| 45 |
A" |
264 |
240 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37059.9 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 33746.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.627 |
1.333 |
0.000 |
| H2 |
0.542 |
2.421 |
0.000 |
| H3 |
1.697 |
1.101 |
0.000 |
| C4 |
-0.016 |
0.748 |
1.260 |
| C5 |
-0.016 |
0.748 |
-1.260 |
| C6 |
-0.016 |
-0.778 |
-1.212 |
| C7 |
-0.016 |
-0.778 |
1.212 |
| N8 |
-0.680 |
-1.235 |
0.000 |
| H9 |
-0.749 |
-2.233 |
0.000 |
| H10 |
0.515 |
1.087 |
2.152 |
| H11 |
0.515 |
1.087 |
-2.152 |
| H12 |
-1.048 |
1.096 |
1.341 |
| H13 |
-1.048 |
1.096 |
-1.341 |
| H14 |
1.022 |
-1.139 |
-1.272 |
| H15 |
1.022 |
-1.139 |
1.272 |
| H16 |
-0.549 |
-1.181 |
-2.073 |
| H17 |
-0.549 |
-1.181 |
2.073 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0913 | 1.0943 | 1.5311 | 1.5311 | 2.5173 | 2.5173 | 2.8810 | 3.8224 | 2.1685 | 2.1685 | 2.1588 | 2.1588 | 2.8076 | 2.8076 | 3.4640 | 3.4640 |
H2 | 1.0913 | | 1.7538 | 2.1675 | 2.1675 | 3.4656 | 3.4656 | 3.8542 | 4.8298 | 2.5315 | 2.5315 | 2.4661 | 2.4661 | 3.8105 | 3.8105 | 4.2965 | 4.2965 | H3 | 1.0943 | 1.7538 | | 2.1555 | 2.1555 | 2.8161 | 2.8161 | 3.3319 | 4.1350 | 2.4547 | 2.4547 | 3.0547 | 3.0547 | 2.6625 | 2.6625 | 3.8140 | 3.8140 | C4 | 1.5311 | 2.1675 | 2.1555 | | 2.5207 | 2.9051 | 1.5263 | 2.4412 | 3.3186 | 1.0917 | 3.4697 | 1.0920 | 2.8200 | 3.3241 | 2.1535 | 3.8877 | 2.1599 | C5 | 1.5311 | 2.1675 | 2.1555 | 2.5207 | | 1.5263 | 2.9051 | 2.4412 | 3.3186 | 3.4697 | 1.0917 | 2.8200 | 1.0920 | 2.1535 | 3.3241 | 2.1599 | 3.8877 | C6 | 2.5173 | 3.4656 | 2.8161 | 2.9051 | 1.5263 | | 2.4238 | 1.4554 | 2.0310 | 3.8825 | 2.1549 | 3.3304 | 2.1427 | 1.1007 | 2.7161 | 1.0899 | 3.3520 | C7 | 2.5173 | 3.4656 | 2.8161 | 1.5263 | 2.9051 | 2.4238 | | 1.4554 | 2.0310 | 2.1549 | 3.8825 | 2.1427 | 3.3304 | 2.7161 | 1.1007 | 3.3520 | 1.0899 | N8 | 2.8810 | 3.8542 | 3.3319 | 2.4412 | 2.4412 | 1.4554 | 1.4554 | | 1.0009 | 3.3837 | 3.3837 | 2.7138 | 2.7138 | 2.1267 | 2.1267 | 2.0775 | 2.0775 | H9 | 3.8224 | 4.8298 | 4.1350 | 3.3186 | 3.3186 | 2.0310 | 2.0310 | 1.0009 | | 4.1539 | 4.1539 | 3.6012 | 3.6012 | 2.4398 | 2.4398 | 2.3330 | 2.3330 | H10 | 2.1685 | 2.5315 | 2.4547 | 1.0917 | 3.4697 | 3.8825 | 2.1549 | 3.3837 | 4.1539 | | 4.3033 | 1.7609 | 3.8265 | 4.1150 | 2.4468 | 4.9116 | 2.5070 | H11 | 2.1685 | 2.5315 | 2.4547 | 3.4697 | 1.0917 | 2.1549 | 3.8825 | 3.3837 | 4.1539 | 4.3033 | | 3.8265 | 1.7609 | 2.4468 | 4.1150 | 2.5070 | 4.9116 | H12 | 2.1588 | 2.4661 | 3.0547 | 1.0920 | 2.8200 | 3.3304 | 2.1427 | 2.7138 | 3.6012 | 1.7609 | 3.8265 | | 2.6819 | 4.0131 | 3.0469 | 4.1335 | 2.4430 | H13 | 2.1588 | 2.4661 | 3.0547 | 2.8200 | 1.0920 | 2.1427 | 3.3304 | 2.7138 | 3.6012 | 3.8265 | 1.7609 | 2.6819 | | 3.0469 | 4.0131 | 2.4430 | 4.1335 | H14 | 2.8076 | 3.8105 | 2.6625 | 3.3241 | 2.1535 | 1.1007 | 2.7161 | 2.1267 | 2.4398 | 4.1150 | 2.4468 | 4.0131 | 3.0469 | | 2.5437 | 1.7640 | 3.6954 | H15 | 2.8076 | 3.8105 | 2.6625 | 2.1535 | 3.3241 | 2.7161 | 1.1007 | 2.1267 | 2.4398 | 2.4468 | 4.1150 | 3.0469 | 4.0131 | 2.5437 | | 3.6954 | 1.7640 | H16 | 3.4640 | 4.2965 | 3.8140 | 3.8877 | 2.1599 | 1.0899 | 3.3520 | 2.0775 | 2.3330 | 4.9116 | 2.5070 | 4.1335 | 2.4430 | 1.7640 | 3.6954 | | 4.1454 | H17 | 3.4640 | 4.2965 | 3.8140 | 2.1599 | 3.8877 | 3.3520 | 1.0899 | 2.0775 | 2.3330 | 2.5070 | 4.9116 | 2.4430 | 4.1335 | 3.6954 | 1.7640 | 4.1454 | |
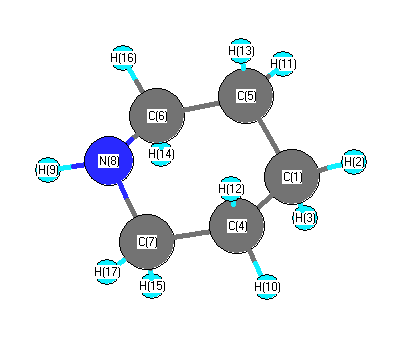 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.837 |
|
C1 |
C4 |
H10 |
110.416 |
| C1 |
C4 |
H12 |
109.638 |
|
C1 |
C5 |
C6 |
110.837 |
| C1 |
C5 |
H11 |
110.416 |
|
C1 |
C5 |
H13 |
109.638 |
| H2 |
C1 |
H3 |
106.726 |
|
H2 |
C1 |
C4 |
110.365 |
| H2 |
C1 |
C5 |
110.365 |
|
H3 |
C1 |
C4 |
109.241 |
| H3 |
C1 |
C5 |
109.241 |
|
C4 |
C1 |
C5 |
110.804 |
| C4 |
C7 |
N8 |
109.892 |
|
C4 |
C7 |
H15 |
109.043 |
| C4 |
C7 |
H17 |
110.178 |
|
C5 |
C6 |
N8 |
109.892 |
| C5 |
C6 |
H14 |
109.043 |
|
C5 |
C6 |
H16 |
110.178 |
| C6 |
C5 |
H11 |
109.674 |
|
C6 |
C5 |
H13 |
108.707 |
| C6 |
N8 |
C7 |
112.755 |
|
C6 |
N8 |
H9 |
110.176 |
| C7 |
C4 |
H10 |
109.674 |
|
C7 |
C4 |
H12 |
108.707 |
| C7 |
N8 |
H9 |
110.176 |
|
N8 |
C6 |
H14 |
111.864 |
| N8 |
C6 |
H16 |
108.561 |
|
N8 |
C7 |
H15 |
111.864 |
| N8 |
C7 |
H17 |
108.561 |
|
H10 |
C4 |
H12 |
107.491 |
| H11 |
C5 |
H13 |
107.491 |
|
H14 |
C6 |
H16 |
107.266 |
| H15 |
C7 |
H17 |
107.266 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.189 |
|
|
|
| 2 |
H |
-0.109 |
|
|
|
| 3 |
H |
-0.160 |
|
|
|
| 4 |
C |
0.278 |
|
|
|
| 5 |
C |
0.278 |
|
|
|
| 6 |
C |
0.748 |
|
|
|
| 7 |
C |
0.748 |
|
|
|
| 8 |
N |
-0.628 |
|
|
|
| 9 |
H |
-0.116 |
|
|
|
| 10 |
H |
-0.101 |
|
|
|
| 11 |
H |
-0.101 |
|
|
|
| 12 |
H |
-0.128 |
|
|
|
| 13 |
H |
-0.128 |
|
|
|
| 14 |
H |
-0.149 |
|
|
|
| 15 |
H |
-0.149 |
|
|
|
| 16 |
H |
-0.236 |
|
|
|
| 17 |
H |
-0.236 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.836 |
-0.422 |
0.000 |
0.937 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.769 |
-0.375 |
0.000 |
| y |
-0.375 |
-37.111 |
0.000 |
| z |
0.000 |
0.000 |
-38.556 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.935 |
-0.375 |
0.000 |
| y |
-0.375 |
3.051 |
0.000 |
| z |
0.000 |
0.000 |
0.884 |
|
| Polar |
|---|
| 3z2-r2 | 1.767 |
|---|
| x2-y2 | -4.658 |
|---|
| xy | -0.375 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.767 |
0.304 |
0.000 |
| y |
0.304 |
9.662 |
0.000 |
| z |
0.000 |
0.000 |
10.073 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -250.219985 |
| Energy at 298.15K | -250.234696 |
| Nuclear repulsion energy | 259.306022 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3735 |
3401 |
0.51 |
|
|
|
| 2 |
A' |
3218 |
2931 |
87.47 |
|
|
|
| 3 |
A' |
3202 |
2916 |
95.08 |
|
|
|
| 4 |
A' |
3197 |
2911 |
76.13 |
|
|
|
| 5 |
A' |
3163 |
2880 |
47.49 |
|
|
|
| 6 |
A' |
3147 |
2866 |
28.37 |
|
|
|
| 7 |
A' |
3145 |
2864 |
34.03 |
|
|
|
| 8 |
A' |
1616 |
1471 |
1.23 |
|
|
|
| 9 |
A' |
1599 |
1456 |
6.31 |
|
|
|
| 10 |
A' |
1590 |
1448 |
4.59 |
|
|
|
| 11 |
A' |
1514 |
1379 |
9.27 |
|
|
|
| 12 |
A' |
1497 |
1363 |
0.56 |
|
|
|
| 13 |
A' |
1439 |
1310 |
0.60 |
|
|
|
| 14 |
A' |
1376 |
1253 |
8.30 |
|
|
|
| 15 |
A' |
1288 |
1173 |
2.32 |
|
|
|
| 16 |
A' |
1125 |
1024 |
2.35 |
|
|
|
| 17 |
A' |
1092 |
994 |
11.72 |
|
|
|
| 18 |
A' |
975 |
888 |
10.22 |
|
|
|
| 19 |
A' |
932 |
848 |
35.26 |
|
|
|
| 20 |
A' |
871 |
793 |
1.65 |
|
|
|
| 21 |
A' |
832 |
758 |
87.95 |
|
|
|
| 22 |
A' |
595 |
542 |
1.81 |
|
|
|
| 23 |
A' |
467 |
426 |
1.02 |
|
|
|
| 24 |
A' |
411 |
374 |
5.36 |
|
|
|
| 25 |
A' |
256 |
233 |
4.36 |
|
|
|
| 26 |
A" |
3215 |
2928 |
64.02 |
|
|
|
| 27 |
A" |
3193 |
2907 |
107.55 |
|
|
|
| 28 |
A" |
3156 |
2874 |
11.45 |
|
|
|
| 29 |
A" |
3142 |
2861 |
28.76 |
|
|
|
| 30 |
A" |
1621 |
1476 |
14.42 |
|
|
|
| 31 |
A" |
1597 |
1454 |
2.54 |
|
|
|
| 32 |
A" |
1584 |
1443 |
3.34 |
|
|
|
| 33 |
A" |
1497 |
1363 |
0.22 |
|
|
|
| 34 |
A" |
1484 |
1351 |
2.38 |
|
|
|
| 35 |
A" |
1450 |
1321 |
0.93 |
|
|
|
| 36 |
A" |
1392 |
1267 |
0.83 |
|
|
|
| 37 |
A" |
1310 |
1193 |
15.48 |
|
|
|
| 38 |
A" |
1229 |
1119 |
21.88 |
|
|
|
| 39 |
A" |
1201 |
1094 |
0.46 |
|
|
|
| 40 |
A" |
1129 |
1028 |
2.84 |
|
|
|
| 41 |
A" |
1020 |
929 |
3.62 |
|
|
|
| 42 |
A" |
935 |
852 |
0.14 |
|
|
|
| 43 |
A" |
868 |
791 |
0.45 |
|
|
|
| 44 |
A" |
488 |
444 |
0.68 |
|
|
|
| 45 |
A" |
254 |
231 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37023.2 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 33713.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.637 |
1.325 |
0.000 |
| H2 |
0.561 |
2.413 |
0.000 |
| H3 |
1.705 |
1.085 |
0.000 |
| C4 |
-0.009 |
0.741 |
1.260 |
| C5 |
-0.009 |
0.741 |
-1.260 |
| C6 |
-0.009 |
-0.789 |
-1.213 |
| C7 |
-0.009 |
-0.789 |
1.213 |
| N8 |
-0.601 |
-1.339 |
0.000 |
| H9 |
-1.589 |
-1.165 |
0.000 |
| H10 |
0.516 |
1.086 |
2.154 |
| H11 |
0.516 |
1.086 |
-2.154 |
| H12 |
-1.040 |
1.099 |
1.339 |
| H13 |
-1.040 |
1.099 |
-1.339 |
| H14 |
1.020 |
-1.153 |
-1.275 |
| H15 |
1.020 |
-1.153 |
1.275 |
| H16 |
-0.540 |
-1.199 |
-2.072 |
| H17 |
-0.540 |
-1.199 |
2.072 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0914 | 1.0943 | 1.5315 | 1.5315 | 2.5212 | 2.5212 | 2.9370 | 3.3397 | 2.1706 | 2.1706 | 2.1585 | 2.1585 | 2.8128 | 2.8128 | 3.4712 | 3.4712 |
H2 | 1.0914 | | 1.7536 | 2.1696 | 2.1696 | 3.4715 | 3.4715 | 3.9276 | 4.1741 | 2.5305 | 2.5305 | 2.4667 | 2.4667 | 3.8154 | 3.8154 | 4.3076 | 4.3076 | H3 | 1.0943 | 1.7536 | | 2.1547 | 2.1547 | 2.8141 | 2.8141 | 3.3449 | 3.9887 | 2.4605 | 2.4605 | 3.0547 | 3.0547 | 2.6651 | 2.6651 | 3.8142 | 3.8142 | C4 | 1.5315 | 2.1696 | 2.1547 | | 2.5201 | 2.9082 | 1.5315 | 2.5030 | 2.7781 | 1.0924 | 3.4713 | 1.0946 | 2.8189 | 3.3277 | 2.1563 | 3.8924 | 2.1697 | C5 | 1.5315 | 2.1696 | 2.1547 | 2.5201 | | 1.5315 | 2.9082 | 2.5030 | 2.7781 | 3.4713 | 1.0924 | 2.8189 | 1.0946 | 2.1563 | 3.3277 | 2.1697 | 3.8924 | C6 | 2.5212 | 3.4715 | 2.8141 | 2.9082 | 1.5315 | | 2.4253 | 1.4569 | 2.0268 | 3.8893 | 2.1629 | 3.3375 | 2.1551 | 1.0933 | 2.7162 | 1.0901 | 3.3524 | C7 | 2.5212 | 3.4715 | 2.8141 | 1.5315 | 2.9082 | 2.4253 | | 1.4569 | 2.0268 | 2.1629 | 3.8893 | 2.1551 | 3.3375 | 2.7162 | 1.0933 | 3.3524 | 1.0901 | N8 | 2.9370 | 3.9276 | 3.3449 | 2.5030 | 2.5030 | 1.4569 | 1.4569 | | 1.0033 | 3.4301 | 3.4301 | 2.8154 | 2.8154 | 2.0702 | 2.0702 | 2.0776 | 2.0776 | H9 | 3.3397 | 4.1741 | 3.9887 | 2.7781 | 2.7781 | 2.0268 | 2.0268 | 1.0033 | | 3.7598 | 3.7598 | 2.6863 | 2.6863 | 2.9037 | 2.9037 | 2.3227 | 2.3227 | H10 | 2.1706 | 2.5305 | 2.4605 | 1.0924 | 3.4713 | 3.8893 | 2.1629 | 3.4301 | 3.7598 | | 4.3081 | 1.7571 | 3.8240 | 4.1261 | 2.4583 | 4.9190 | 2.5187 | H11 | 2.1706 | 2.5305 | 2.4605 | 3.4713 | 1.0924 | 2.1629 | 3.8893 | 3.4301 | 3.7598 | 4.3081 | | 3.8240 | 1.7571 | 2.4583 | 4.1261 | 2.5187 | 4.9190 | H12 | 2.1585 | 2.4667 | 3.0547 | 1.0946 | 2.8189 | 3.3375 | 2.1551 | 2.8154 | 2.6863 | 1.7571 | 3.8240 | | 2.6777 | 4.0185 | 3.0533 | 4.1431 | 2.4636 | H13 | 2.1585 | 2.4667 | 3.0547 | 2.8189 | 1.0946 | 2.1551 | 3.3375 | 2.8154 | 2.6863 | 3.8240 | 1.7571 | 2.6777 | | 3.0533 | 4.0185 | 2.4636 | 4.1431 | H14 | 2.8128 | 3.8154 | 2.6651 | 3.3277 | 2.1563 | 1.0933 | 2.7162 | 2.0702 | 2.9037 | 4.1261 | 2.4583 | 4.0185 | 3.0533 | | 2.5491 | 1.7524 | 3.6925 | H15 | 2.8128 | 3.8154 | 2.6651 | 2.1563 | 3.3277 | 2.7162 | 1.0933 | 2.0702 | 2.9037 | 2.4583 | 4.1261 | 3.0533 | 4.0185 | 2.5491 | | 3.6925 | 1.7524 | H16 | 3.4712 | 4.3076 | 3.8142 | 3.8924 | 2.1697 | 1.0901 | 3.3524 | 2.0776 | 2.3227 | 4.9190 | 2.5187 | 4.1431 | 2.4636 | 1.7524 | 3.6925 | | 4.1440 | H17 | 3.4712 | 4.3076 | 3.8142 | 2.1697 | 3.8924 | 3.3524 | 1.0901 | 2.0776 | 2.3227 | 2.5187 | 4.9190 | 2.4636 | 4.1431 | 3.6925 | 1.7524 | 4.1440 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.801 |
|
C1 |
C4 |
H10 |
110.521 |
| C1 |
C4 |
H12 |
109.432 |
|
C1 |
C5 |
C6 |
110.801 |
| C1 |
C5 |
H11 |
110.521 |
|
C1 |
C5 |
H13 |
109.432 |
| H2 |
C1 |
H3 |
106.702 |
|
H2 |
C1 |
C4 |
110.496 |
| H2 |
C1 |
C5 |
110.496 |
|
H3 |
C1 |
C4 |
109.159 |
| H3 |
C1 |
C5 |
109.159 |
|
C4 |
C1 |
C5 |
110.727 |
| C4 |
C7 |
N8 |
113.745 |
|
C4 |
C7 |
H15 |
109.342 |
| C4 |
C7 |
H17 |
110.585 |
|
C5 |
C6 |
N8 |
113.745 |
| C5 |
C6 |
H14 |
109.342 |
|
C5 |
C6 |
H16 |
110.585 |
| C6 |
C5 |
H11 |
109.913 |
|
C6 |
C5 |
H13 |
109.170 |
| C6 |
N8 |
C7 |
112.678 |
|
C6 |
N8 |
H9 |
109.557 |
| C7 |
C4 |
H10 |
109.913 |
|
C7 |
C4 |
H12 |
109.170 |
| C7 |
N8 |
H9 |
109.557 |
|
N8 |
C6 |
H14 |
107.688 |
| N8 |
C6 |
H16 |
108.449 |
|
N8 |
C7 |
H15 |
107.688 |
| N8 |
C7 |
H17 |
108.449 |
|
H10 |
C4 |
H12 |
106.913 |
| H11 |
C5 |
H13 |
106.913 |
|
H14 |
C6 |
H16 |
106.759 |
| H15 |
C7 |
H17 |
106.759 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.267 |
|
|
|
| 2 |
H |
-0.102 |
|
|
|
| 3 |
H |
-0.150 |
|
|
|
| 4 |
C |
0.259 |
|
|
|
| 5 |
C |
0.259 |
|
|
|
| 6 |
C |
0.657 |
|
|
|
| 7 |
C |
0.657 |
|
|
|
| 8 |
N |
-0.597 |
|
|
|
| 9 |
H |
-0.033 |
|
|
|
| 10 |
H |
-0.113 |
|
|
|
| 11 |
H |
-0.113 |
|
|
|
| 12 |
H |
-0.135 |
|
|
|
| 13 |
H |
-0.135 |
|
|
|
| 14 |
H |
-0.167 |
|
|
|
| 15 |
H |
-0.167 |
|
|
|
| 16 |
H |
-0.192 |
|
|
|
| 17 |
H |
-0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.225 |
1.232 |
0.000 |
1.252 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.365 |
-0.424 |
0.000 |
| y |
-0.424 |
-44.308 |
0.000 |
| z |
0.000 |
0.000 |
-38.453 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.016 |
-0.424 |
0.000 |
| y |
-0.424 |
-6.399 |
0.000 |
| z |
0.000 |
0.000 |
2.383 |
|
| Polar |
|---|
| 3z2-r2 | 4.766 |
|---|
| x2-y2 | 6.943 |
|---|
| xy | -0.424 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.736 |
0.330 |
0.000 |
| y |
0.330 |
9.541 |
0.000 |
| z |
0.000 |
0.000 |
10.063 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
