Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -251.881097 |
| Energy at 298.15K | -251.895477 |
| Nuclear repulsion energy | 259.513019 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3572 |
3423 |
0.22 |
|
|
|
| 2 |
A' |
3115 |
2985 |
72.81 |
|
|
|
| 3 |
A' |
3101 |
2972 |
6.95 |
|
|
|
| 4 |
A' |
3098 |
2969 |
60.89 |
|
|
|
| 5 |
A' |
3058 |
2930 |
18.98 |
|
|
|
| 6 |
A' |
3040 |
2913 |
31.59 |
|
|
|
| 7 |
A' |
2934 |
2811 |
154.20 |
|
|
|
| 8 |
A' |
1494 |
1432 |
2.21 |
|
|
|
| 9 |
A' |
1477 |
1416 |
9.27 |
|
|
|
| 10 |
A' |
1467 |
1405 |
8.22 |
|
|
|
| 11 |
A' |
1416 |
1357 |
1.97 |
|
|
|
| 12 |
A' |
1375 |
1317 |
0.33 |
|
|
|
| 13 |
A' |
1312 |
1257 |
1.08 |
|
|
|
| 14 |
A' |
1282 |
1229 |
2.95 |
|
|
|
| 15 |
A' |
1165 |
1116 |
4.41 |
|
|
|
| 16 |
A' |
1072 |
1027 |
1.60 |
|
|
|
| 17 |
A' |
1059 |
1015 |
11.00 |
|
|
|
| 18 |
A' |
923 |
884 |
2.66 |
|
|
|
| 19 |
A' |
870 |
834 |
13.96 |
|
|
|
| 20 |
A' |
843 |
808 |
0.14 |
|
|
|
| 21 |
A' |
770 |
738 |
72.83 |
|
|
|
| 22 |
A' |
541 |
518 |
20.84 |
|
|
|
| 23 |
A' |
434 |
416 |
6.20 |
|
|
|
| 24 |
A' |
394 |
378 |
1.19 |
|
|
|
| 25 |
A' |
246 |
236 |
1.29 |
|
|
|
| 26 |
A" |
3110 |
2980 |
19.71 |
|
|
|
| 27 |
A" |
3099 |
2970 |
76.30 |
|
|
|
| 28 |
A" |
3058 |
2930 |
30.12 |
|
|
|
| 29 |
A" |
2929 |
2807 |
35.45 |
|
|
|
| 30 |
A" |
1491 |
1429 |
2.84 |
|
|
|
| 31 |
A" |
1477 |
1415 |
2.08 |
|
|
|
| 32 |
A" |
1458 |
1397 |
5.39 |
|
|
|
| 33 |
A" |
1377 |
1320 |
0.17 |
|
|
|
| 34 |
A" |
1357 |
1300 |
20.39 |
|
|
|
| 35 |
A" |
1336 |
1281 |
8.10 |
|
|
|
| 36 |
A" |
1291 |
1237 |
2.77 |
|
|
|
| 37 |
A" |
1199 |
1149 |
12.97 |
|
|
|
| 38 |
A" |
1165 |
1116 |
6.74 |
|
|
|
| 39 |
A" |
1158 |
1109 |
1.93 |
|
|
|
| 40 |
A" |
1077 |
1032 |
2.72 |
|
|
|
| 41 |
A" |
974 |
933 |
0.61 |
|
|
|
| 42 |
A" |
902 |
864 |
0.45 |
|
|
|
| 43 |
A" |
816 |
782 |
0.38 |
|
|
|
| 44 |
A" |
452 |
433 |
0.76 |
|
|
|
| 45 |
A" |
246 |
236 |
0.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35011.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33551.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.631 |
1.326 |
0.000 |
| H2 |
0.559 |
2.422 |
0.000 |
| H3 |
1.705 |
1.080 |
0.000 |
| C4 |
-0.015 |
0.747 |
1.258 |
| C5 |
-0.015 |
0.747 |
-1.258 |
| C6 |
-0.015 |
-0.776 |
-1.209 |
| C7 |
-0.015 |
-0.776 |
1.209 |
| N8 |
-0.690 |
-1.228 |
0.000 |
| H9 |
-0.767 |
-2.239 |
0.000 |
| H10 |
0.516 |
1.089 |
2.156 |
| H11 |
0.516 |
1.089 |
-2.156 |
| H12 |
-1.054 |
1.096 |
1.335 |
| H13 |
-1.054 |
1.096 |
-1.335 |
| H14 |
1.035 |
-1.135 |
-1.260 |
| H15 |
1.035 |
-1.135 |
1.260 |
| H16 |
-0.543 |
-1.187 |
-2.079 |
| H17 |
-0.543 |
-1.187 |
2.079 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0984 | 1.1017 | 1.5276 | 1.5276 | 2.5093 | 2.5093 | 2.8758 | 3.8290 | 2.1722 | 2.1722 | 2.1622 | 2.1622 | 2.7944 | 2.7944 | 3.4661 | 3.4661 |
H2 | 1.0984 | | 1.7642 | 2.1719 | 2.1719 | 3.4668 | 3.4668 | 3.8583 | 4.8459 | 2.5352 | 2.5352 | 2.4787 | 2.4787 | 3.8037 | 3.8037 | 4.3081 | 4.3081 | H3 | 1.1017 | 1.7642 | | 2.1564 | 2.1564 | 2.8045 | 2.8045 | 3.3268 | 4.1385 | 2.4623 | 2.4623 | 3.0654 | 3.0654 | 2.6355 | 2.6355 | 3.8099 | 3.8099 | C4 | 1.5276 | 2.1719 | 2.1564 | | 2.5150 | 2.8987 | 1.5238 | 2.4370 | 3.3259 | 1.0986 | 3.4717 | 1.0995 | 2.8150 | 3.3140 | 2.1552 | 3.8923 | 2.1662 | C5 | 1.5276 | 2.1719 | 2.1564 | 2.5150 | | 1.5238 | 2.8987 | 2.4370 | 3.3259 | 3.4717 | 1.0986 | 2.8150 | 1.0995 | 2.1552 | 3.3140 | 2.1662 | 3.8923 | C6 | 2.5093 | 3.4668 | 2.8045 | 2.8987 | 1.5238 | | 2.4176 | 1.4567 | 2.0412 | 3.8840 | 2.1586 | 3.3253 | 2.1454 | 1.1107 | 2.7066 | 1.0977 | 3.3553 | C7 | 2.5093 | 3.4668 | 2.8045 | 1.5238 | 2.8987 | 2.4176 | | 1.4567 | 2.0412 | 2.1586 | 3.8840 | 2.1454 | 3.3253 | 2.7066 | 1.1107 | 3.3553 | 1.0977 | N8 | 2.8758 | 3.8583 | 3.3268 | 2.4370 | 2.4370 | 1.4567 | 1.4567 | | 1.0134 | 3.3877 | 3.3877 | 2.7053 | 2.7053 | 2.1384 | 2.1384 | 2.0847 | 2.0847 | H9 | 3.8290 | 4.8459 | 4.1385 | 3.3259 | 3.3259 | 2.0412 | 2.0412 | 1.0134 | | 4.1680 | 4.1680 | 3.6039 | 3.6039 | 2.4600 | 2.4600 | 2.3409 | 2.3409 | H10 | 2.1722 | 2.5352 | 2.4623 | 1.0986 | 3.4717 | 3.8840 | 2.1586 | 3.3877 | 4.1680 | | 4.3125 | 1.7722 | 3.8282 | 4.1096 | 2.4540 | 4.9234 | 2.5115 | H11 | 2.1722 | 2.5352 | 2.4623 | 3.4717 | 1.0986 | 2.1586 | 3.8840 | 3.3877 | 4.1680 | 4.3125 | | 3.8282 | 1.7722 | 2.4540 | 4.1096 | 2.5115 | 4.9234 | H12 | 2.1622 | 2.4787 | 3.0654 | 1.0995 | 2.8150 | 3.3253 | 2.1454 | 2.7053 | 3.6039 | 1.7722 | 3.8282 | | 2.6701 | 4.0099 | 3.0580 | 4.1387 | 2.4549 | H13 | 2.1622 | 2.4787 | 3.0654 | 2.8150 | 1.0995 | 2.1454 | 3.3253 | 2.7053 | 3.6039 | 3.8282 | 1.7722 | 2.6701 | | 3.0580 | 4.0099 | 2.4549 | 4.1387 | H14 | 2.7944 | 3.8037 | 2.6355 | 3.3140 | 2.1552 | 1.1107 | 2.7066 | 2.1384 | 2.4600 | 4.1096 | 2.4540 | 4.0099 | 3.0580 | | 2.5199 | 1.7788 | 3.6935 | H15 | 2.7944 | 3.8037 | 2.6355 | 2.1552 | 3.3140 | 2.7066 | 1.1107 | 2.1384 | 2.4600 | 2.4540 | 4.1096 | 3.0580 | 4.0099 | 2.5199 | | 3.6935 | 1.7788 | H16 | 3.4661 | 4.3081 | 3.8099 | 3.8923 | 2.1662 | 1.0977 | 3.3553 | 2.0847 | 2.3409 | 4.9234 | 2.5115 | 4.1387 | 2.4549 | 1.7788 | 3.6935 | | 4.1582 | H17 | 3.4661 | 4.3081 | 3.8099 | 2.1662 | 3.8923 | 3.3553 | 1.0977 | 2.0847 | 2.3409 | 2.5115 | 4.9234 | 2.4549 | 4.1387 | 3.6935 | 1.7788 | 4.1582 | |
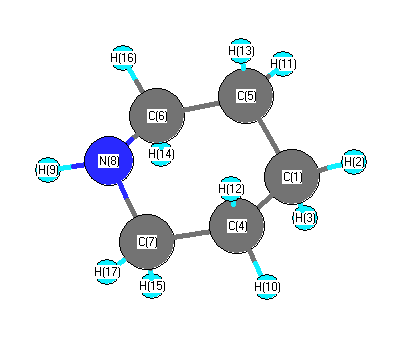 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.642 |
|
C1 |
C4 |
H10 |
110.548 |
| C1 |
C4 |
H12 |
109.711 |
|
C1 |
C5 |
C6 |
110.642 |
| C1 |
C5 |
H11 |
110.548 |
|
C1 |
C5 |
H13 |
109.711 |
| H2 |
C1 |
H3 |
106.615 |
|
H2 |
C1 |
C4 |
110.529 |
| H2 |
C1 |
C5 |
110.529 |
|
H3 |
C1 |
C4 |
109.123 |
| H3 |
C1 |
C5 |
109.123 |
|
C4 |
C1 |
C5 |
110.811 |
| C4 |
C7 |
N8 |
109.683 |
|
C4 |
C7 |
H15 |
108.774 |
| C4 |
C7 |
H17 |
110.388 |
|
C5 |
C6 |
N8 |
109.683 |
| C5 |
C6 |
H14 |
108.774 |
|
C5 |
C6 |
H16 |
110.388 |
| C6 |
C5 |
H11 |
109.740 |
|
C6 |
C5 |
H13 |
108.657 |
| C6 |
N8 |
C7 |
112.158 |
|
C6 |
N8 |
H9 |
110.157 |
| C7 |
C4 |
H10 |
109.740 |
|
C7 |
C4 |
H12 |
108.657 |
| C7 |
N8 |
H9 |
110.157 |
|
N8 |
C6 |
H14 |
112.088 |
| N8 |
C6 |
H16 |
108.578 |
|
N8 |
C7 |
H15 |
112.088 |
| N8 |
C7 |
H17 |
108.578 |
|
H10 |
C4 |
H12 |
107.467 |
| H11 |
C5 |
H13 |
107.467 |
|
H14 |
C6 |
H16 |
107.304 |
| H15 |
C7 |
H17 |
107.304 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.443 |
|
|
|
| 2 |
H |
-0.216 |
|
|
|
| 3 |
H |
-0.284 |
|
|
|
| 4 |
C |
0.548 |
|
|
|
| 5 |
C |
0.548 |
|
|
|
| 6 |
C |
0.775 |
|
|
|
| 7 |
C |
0.775 |
|
|
|
| 8 |
N |
-0.339 |
|
|
|
| 9 |
H |
-0.099 |
|
|
|
| 10 |
H |
-0.212 |
|
|
|
| 11 |
H |
-0.212 |
|
|
|
| 12 |
H |
-0.261 |
|
|
|
| 13 |
H |
-0.261 |
|
|
|
| 14 |
H |
-0.263 |
|
|
|
| 15 |
H |
-0.263 |
|
|
|
| 16 |
H |
-0.340 |
|
|
|
| 17 |
H |
-0.340 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.727 |
-0.424 |
0.000 |
0.842 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.584 |
-0.151 |
0.000 |
| y |
-0.151 |
-36.766 |
0.000 |
| z |
0.000 |
0.000 |
-38.232 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.085 |
-0.151 |
0.000 |
| y |
-0.151 |
3.142 |
0.000 |
| z |
0.000 |
0.000 |
0.943 |
|
| Polar |
|---|
| 3z2-r2 | 1.887 |
|---|
| x2-y2 | -4.818 |
|---|
| xy | -0.151 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.558 |
0.315 |
0.000 |
| y |
0.315 |
10.368 |
0.000 |
| z |
0.000 |
0.000 |
10.822 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -251.879744 |
| Energy at 298.15K | -251.894054 |
| Nuclear repulsion energy | 259.075314 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3534 |
3386 |
0.73 |
|
|
|
| 2 |
A' |
3107 |
2977 |
69.27 |
|
|
|
| 3 |
A' |
3099 |
2970 |
42.53 |
|
|
|
| 4 |
A' |
3092 |
2963 |
48.55 |
|
|
|
| 5 |
A' |
3041 |
2914 |
64.99 |
|
|
|
| 6 |
A' |
3037 |
2911 |
29.51 |
|
|
|
| 7 |
A' |
3036 |
2909 |
8.88 |
|
|
|
| 8 |
A' |
1487 |
1425 |
2.48 |
|
|
|
| 9 |
A' |
1469 |
1408 |
11.14 |
|
|
|
| 10 |
A' |
1467 |
1405 |
4.77 |
|
|
|
| 11 |
A' |
1388 |
1330 |
3.87 |
|
|
|
| 12 |
A' |
1374 |
1316 |
2.59 |
|
|
|
| 13 |
A' |
1335 |
1279 |
0.21 |
|
|
|
| 14 |
A' |
1276 |
1223 |
6.39 |
|
|
|
| 15 |
A' |
1186 |
1137 |
1.82 |
|
|
|
| 16 |
A' |
1058 |
1014 |
2.18 |
|
|
|
| 17 |
A' |
1024 |
982 |
7.61 |
|
|
|
| 18 |
A' |
925 |
887 |
0.94 |
|
|
|
| 19 |
A' |
865 |
829 |
36.55 |
|
|
|
| 20 |
A' |
834 |
799 |
0.83 |
|
|
|
| 21 |
A' |
754 |
722 |
94.87 |
|
|
|
| 22 |
A' |
545 |
522 |
2.66 |
|
|
|
| 23 |
A' |
436 |
418 |
1.00 |
|
|
|
| 24 |
A' |
380 |
364 |
5.87 |
|
|
|
| 25 |
A' |
237 |
227 |
5.08 |
|
|
|
| 26 |
A" |
3105 |
2975 |
34.26 |
|
|
|
| 27 |
A" |
3091 |
2962 |
76.52 |
|
|
|
| 28 |
A" |
3038 |
2911 |
0.44 |
|
|
|
| 29 |
A" |
3032 |
2905 |
50.83 |
|
|
|
| 30 |
A" |
1491 |
1429 |
9.22 |
|
|
|
| 31 |
A" |
1466 |
1405 |
5.48 |
|
|
|
| 32 |
A" |
1457 |
1396 |
3.51 |
|
|
|
| 33 |
A" |
1376 |
1318 |
0.27 |
|
|
|
| 34 |
A" |
1361 |
1304 |
1.48 |
|
|
|
| 35 |
A" |
1333 |
1278 |
1.38 |
|
|
|
| 36 |
A" |
1294 |
1240 |
0.74 |
|
|
|
| 37 |
A" |
1225 |
1174 |
17.74 |
|
|
|
| 38 |
A" |
1154 |
1106 |
12.46 |
|
|
|
| 39 |
A" |
1118 |
1071 |
0.98 |
|
|
|
| 40 |
A" |
1074 |
1029 |
3.17 |
|
|
|
| 41 |
A" |
945 |
906 |
3.71 |
|
|
|
| 42 |
A" |
893 |
856 |
0.25 |
|
|
|
| 43 |
A" |
803 |
769 |
0.13 |
|
|
|
| 44 |
A" |
456 |
437 |
0.82 |
|
|
|
| 45 |
A" |
231 |
222 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34962.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 33504.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.639 |
1.320 |
0.000 |
| H2 |
0.575 |
2.417 |
0.000 |
| H3 |
1.711 |
1.069 |
0.000 |
| C4 |
-0.009 |
0.743 |
1.257 |
| C5 |
-0.009 |
0.743 |
-1.257 |
| C6 |
-0.009 |
-0.786 |
-1.210 |
| C7 |
-0.009 |
-0.786 |
1.210 |
| N8 |
-0.603 |
-1.343 |
0.000 |
| H9 |
-1.602 |
-1.157 |
0.000 |
| H10 |
0.515 |
1.092 |
2.159 |
| H11 |
0.515 |
1.092 |
-2.159 |
| H12 |
-1.048 |
1.101 |
1.335 |
| H13 |
-1.048 |
1.101 |
-1.335 |
| H14 |
1.029 |
-1.152 |
-1.266 |
| H15 |
1.029 |
-1.152 |
1.266 |
| H16 |
-0.536 |
-1.203 |
-2.079 |
| H17 |
-0.536 |
-1.203 |
2.079 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0985 | 1.1018 | 1.5279 | 1.5279 | 2.5143 | 2.5143 | 2.9387 | 3.3400 | 2.1742 | 2.1742 | 2.1624 | 2.1624 | 2.8043 | 2.8043 | 3.4741 | 3.4741 |
H2 | 1.0985 | | 1.7633 | 2.1736 | 2.1736 | 3.4735 | 3.4735 | 3.9402 | 4.1841 | 2.5337 | 2.5337 | 2.4792 | 2.4792 | 3.8134 | 3.8134 | 4.3198 | 4.3198 | H3 | 1.1018 | 1.7633 | | 2.1559 | 2.1559 | 2.8049 | 2.8049 | 3.3431 | 3.9916 | 2.4679 | 2.4679 | 3.0658 | 3.0658 | 2.6455 | 2.6455 | 3.8125 | 3.8125 | C4 | 1.5279 | 2.1736 | 2.1559 | | 2.5149 | 2.9030 | 1.5296 | 2.5070 | 2.7795 | 1.0995 | 3.4737 | 1.1018 | 2.8162 | 3.3214 | 2.1601 | 3.8981 | 2.1770 | C5 | 1.5279 | 2.1736 | 2.1559 | 2.5149 | | 1.5296 | 2.9030 | 2.5070 | 2.7795 | 3.4737 | 1.0995 | 2.8162 | 1.1018 | 2.1601 | 3.3214 | 2.1770 | 3.8981 | C6 | 2.5143 | 3.4735 | 2.8049 | 2.9030 | 1.5296 | | 2.4206 | 1.4587 | 2.0346 | 3.8924 | 2.1680 | 3.3350 | 2.1581 | 1.1020 | 2.7095 | 1.0980 | 3.3570 | C7 | 2.5143 | 3.4735 | 2.8049 | 1.5296 | 2.9030 | 2.4206 | | 1.4587 | 2.0346 | 2.1680 | 3.8924 | 2.1581 | 3.3350 | 2.7095 | 1.1020 | 3.3570 | 1.0980 | N8 | 2.9387 | 3.9402 | 3.3431 | 2.5070 | 2.5070 | 1.4587 | 1.4587 | | 1.0164 | 3.4408 | 3.4408 | 2.8208 | 2.8208 | 2.0741 | 2.0741 | 2.0846 | 2.0846 | H9 | 3.3400 | 4.1841 | 3.9916 | 2.7795 | 2.7795 | 2.0346 | 2.0346 | 1.0164 | | 3.7680 | 3.7680 | 2.6810 | 2.6810 | 2.9197 | 2.9197 | 2.3369 | 2.3369 | H10 | 2.1742 | 2.5337 | 2.4679 | 1.0995 | 3.4737 | 3.8924 | 2.1680 | 3.4408 | 3.7680 | | 4.3172 | 1.7670 | 3.8279 | 4.1256 | 2.4684 | 4.9323 | 2.5255 | H11 | 2.1742 | 2.5337 | 2.4679 | 3.4737 | 1.0995 | 2.1680 | 3.8924 | 3.4408 | 3.7680 | 4.3172 | | 3.8279 | 1.7670 | 2.4684 | 4.1256 | 2.5255 | 4.9323 | H12 | 2.1624 | 2.4792 | 3.0658 | 1.1018 | 2.8162 | 3.3350 | 2.1581 | 2.8208 | 2.6810 | 1.7670 | 3.8279 | | 2.6708 | 4.0192 | 3.0650 | 4.1510 | 2.4752 | H13 | 2.1624 | 2.4792 | 3.0658 | 2.8162 | 1.1018 | 2.1581 | 3.3350 | 2.8208 | 2.6810 | 3.8279 | 1.7670 | 2.6708 | | 3.0650 | 4.0192 | 2.4752 | 4.1510 | H14 | 2.8043 | 3.8134 | 2.6455 | 3.3214 | 2.1601 | 1.1020 | 2.7095 | 2.0741 | 2.9197 | 4.1256 | 2.4684 | 4.0192 | 3.0650 | | 2.5311 | 1.7642 | 3.6927 | H15 | 2.8043 | 3.8134 | 2.6455 | 2.1601 | 3.3214 | 2.7095 | 1.1020 | 2.0741 | 2.9197 | 2.4684 | 4.1256 | 3.0650 | 4.0192 | 2.5311 | | 3.6927 | 1.7642 | H16 | 3.4741 | 4.3198 | 3.8125 | 3.8981 | 2.1770 | 1.0980 | 3.3570 | 2.0846 | 2.3369 | 4.9323 | 2.5255 | 4.1510 | 2.4752 | 1.7642 | 3.6927 | | 4.1576 | H17 | 3.4741 | 4.3198 | 3.8125 | 2.1770 | 3.8981 | 3.3570 | 1.0980 | 2.0846 | 2.3369 | 2.5255 | 4.9323 | 2.4752 | 4.1510 | 3.6927 | 1.7642 | 4.1576 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.641 |
|
C1 |
C4 |
H10 |
110.629 |
| C1 |
C4 |
H12 |
109.567 |
|
C1 |
C5 |
C6 |
110.641 |
| C1 |
C5 |
H11 |
110.629 |
|
C1 |
C5 |
H13 |
109.567 |
| H2 |
C1 |
H3 |
106.527 |
|
H2 |
C1 |
C4 |
110.649 |
| H2 |
C1 |
C5 |
110.649 |
|
H3 |
C1 |
C4 |
109.062 |
| H3 |
C1 |
C5 |
109.062 |
|
C4 |
C1 |
C5 |
110.774 |
| C4 |
C7 |
N8 |
114.034 |
|
C4 |
C7 |
H15 |
109.261 |
| C4 |
C7 |
H17 |
110.820 |
|
C5 |
C6 |
N8 |
114.034 |
| C5 |
C6 |
H14 |
109.261 |
|
C5 |
C6 |
H16 |
110.820 |
| C6 |
C5 |
H11 |
110.025 |
|
C6 |
C5 |
H13 |
109.116 |
| C6 |
N8 |
C7 |
112.137 |
|
C6 |
N8 |
H9 |
109.271 |
| C7 |
C4 |
H10 |
110.025 |
|
C7 |
C4 |
H12 |
109.116 |
| C7 |
N8 |
H9 |
109.271 |
|
N8 |
C6 |
H14 |
107.369 |
| N8 |
C6 |
H16 |
108.415 |
|
N8 |
C7 |
H15 |
107.369 |
| N8 |
C7 |
H17 |
108.415 |
|
H10 |
C4 |
H12 |
106.774 |
| H11 |
C5 |
H13 |
106.774 |
|
H14 |
C6 |
H16 |
106.629 |
| H15 |
C7 |
H17 |
106.629 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.494 |
|
|
|
| 2 |
H |
-0.206 |
|
|
|
| 3 |
H |
-0.270 |
|
|
|
| 4 |
C |
0.538 |
|
|
|
| 5 |
C |
0.538 |
|
|
|
| 6 |
C |
0.659 |
|
|
|
| 7 |
C |
0.659 |
|
|
|
| 8 |
N |
-0.336 |
|
|
|
| 9 |
H |
-0.016 |
|
|
|
| 10 |
H |
-0.225 |
|
|
|
| 11 |
H |
-0.225 |
|
|
|
| 12 |
H |
-0.263 |
|
|
|
| 13 |
H |
-0.263 |
|
|
|
| 14 |
H |
-0.263 |
|
|
|
| 15 |
H |
-0.263 |
|
|
|
| 16 |
H |
-0.278 |
|
|
|
| 17 |
H |
-0.278 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.227 |
1.181 |
0.000 |
1.203 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.197 |
-0.360 |
0.000 |
| y |
-0.360 |
-43.816 |
0.000 |
| z |
0.000 |
0.000 |
-38.153 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.787 |
-0.360 |
0.000 |
| y |
-0.360 |
-6.141 |
0.000 |
| z |
0.000 |
0.000 |
2.354 |
|
| Polar |
|---|
| 3z2-r2 | 4.708 |
|---|
| x2-y2 | 6.618 |
|---|
| xy | -0.360 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.358 |
0.379 |
0.000 |
| y |
0.379 |
10.315 |
0.000 |
| z |
0.000 |
0.000 |
10.821 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |
