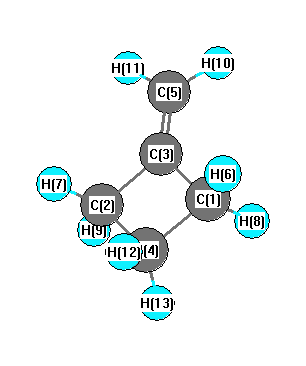
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3161 |
3029 |
5.07 |
|
|
|
| 2 |
A' |
3159 |
3028 |
41.66 |
|
|
|
| 3 |
A' |
3127 |
2996 |
11.42 |
|
|
|
| 4 |
A' |
3095 |
2966 |
44.52 |
|
|
|
| 5 |
A' |
3069 |
2941 |
10.32 |
|
|
|
| 6 |
A' |
1771 |
1697 |
33.90 |
|
|
|
| 7 |
A' |
1489 |
1427 |
0.11 |
|
|
|
| 8 |
A' |
1460 |
1399 |
2.55 |
|
|
|
| 9 |
A' |
1428 |
1369 |
1.50 |
|
|
|
| 10 |
A' |
1247 |
1195 |
0.01 |
|
|
|
| 11 |
A' |
1216 |
1165 |
0.43 |
|
|
|
| 12 |
A' |
1084 |
1039 |
0.78 |
|
|
|
| 13 |
A' |
992 |
951 |
0.53 |
|
|
|
| 14 |
A' |
910 |
872 |
46.33 |
|
|
|
| 15 |
A' |
895 |
858 |
0.97 |
|
|
|
| 16 |
A' |
750 |
719 |
1.81 |
|
|
|
| 17 |
A' |
667 |
639 |
2.19 |
|
|
|
| 18 |
A' |
384 |
368 |
7.48 |
|
|
|
| 19 |
A' |
109 |
104 |
0.10 |
|
|
|
| 20 |
A" |
3254 |
3119 |
10.97 |
|
|
|
| 21 |
A" |
3131 |
3000 |
18.12 |
|
|
|
| 22 |
A" |
3065 |
2937 |
43.68 |
|
|
|
| 23 |
A" |
1445 |
1385 |
6.71 |
|
|
|
| 24 |
A" |
1283 |
1230 |
0.09 |
|
|
|
| 25 |
A" |
1235 |
1184 |
0.34 |
|
|
|
| 26 |
A" |
1205 |
1155 |
0.61 |
|
|
|
| 27 |
A" |
1190 |
1140 |
1.51 |
|
|
|
| 28 |
A" |
1032 |
989 |
0.15 |
|
|
|
| 29 |
A" |
946 |
907 |
1.08 |
|
|
|
| 30 |
A" |
914 |
876 |
1.31 |
|
|
|
| 31 |
A" |
803 |
770 |
0.06 |
|
|
|
| 32 |
A" |
635 |
609 |
0.09 |
|
|
|
| 33 |
A" |
354 |
339 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25252.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 24199.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.368 |
|
|
|
| 2 |
C |
0.368 |
|
|
|
| 3 |
C |
0.306 |
|
|
|
| 4 |
C |
0.878 |
|
|
|
| 5 |
C |
0.744 |
|
|
|
| 6 |
H |
-0.349 |
|
|
|
| 7 |
H |
-0.349 |
|
|
|
| 8 |
H |
-0.200 |
|
|
|
| 9 |
H |
-0.200 |
|
|
|
| 10 |
H |
-0.441 |
|
|
|
| 11 |
H |
-0.441 |
|
|
|
| 12 |
H |
-0.377 |
|
|
|
| 13 |
H |
-0.306 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.222 |
-0.573 |
0.000 |
0.614 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.142 |
-0.627 |
0.000 |
| y |
-0.627 |
-32.433 |
0.000 |
| z |
0.000 |
0.000 |
-30.843 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.504 |
-0.627 |
0.000 |
| y |
-0.627 |
-0.940 |
0.000 |
| z |
0.000 |
0.000 |
1.444 |
|
| Polar |
|---|
| 3z2-r2 | 2.889 |
|---|
| x2-y2 | 0.291 |
|---|
| xy | -0.627 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.828 |
-1.482 |
0.000 |
| y |
-1.482 |
10.773 |
0.000 |
| z |
0.000 |
0.000 |
8.725 |
<r2> (average value of r
2) Å
2
| <r2> |
115.908 |
| (<r2>)1/2 |
10.766 |