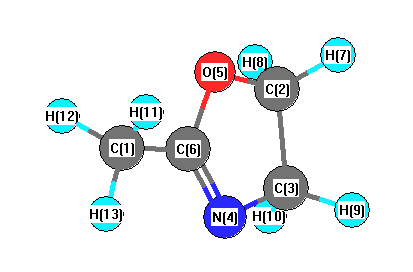
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3307 |
3011 |
10.13 |
|
|
|
| 2 |
A |
3223 |
2935 |
73.11 |
|
|
|
| 3 |
A |
3197 |
2911 |
55.07 |
|
|
|
| 4 |
A |
3190 |
2905 |
14.76 |
|
|
|
| 5 |
A |
1909 |
1739 |
232.68 |
|
|
|
| 6 |
A |
1646 |
1499 |
2.85 |
|
|
|
| 7 |
A |
1619 |
1474 |
0.31 |
|
|
|
| 8 |
A |
1574 |
1433 |
8.85 |
|
|
|
| 9 |
A |
1539 |
1401 |
58.11 |
|
|
|
| 10 |
A |
1495 |
1361 |
7.69 |
|
|
|
| 11 |
A |
1460 |
1329 |
22.27 |
|
|
|
| 12 |
A |
1356 |
1235 |
120.83 |
|
|
|
| 13 |
A |
1183 |
1077 |
4.01 |
|
|
|
| 14 |
A |
1134 |
1032 |
111.10 |
|
|
|
| 15 |
A |
1073 |
977 |
3.79 |
|
|
|
| 16 |
A |
1027 |
935 |
29.98 |
|
|
|
| 17 |
A |
991 |
902 |
21.63 |
|
|
|
| 18 |
A |
823 |
750 |
6.48 |
|
|
|
| 19 |
A |
701 |
638 |
2.99 |
|
|
|
| 20 |
A |
374 |
341 |
7.18 |
|
|
|
| 21 |
A |
3273 |
2980 |
54.88 |
|
|
|
| 22 |
A |
3254 |
2964 |
14.31 |
|
|
|
| 23 |
A |
3228 |
2940 |
12.88 |
|
|
|
| 24 |
A |
1576 |
1435 |
7.36 |
|
|
|
| 25 |
A |
1345 |
1225 |
0.05 |
|
|
|
| 26 |
A |
1316 |
1198 |
4.87 |
|
|
|
| 27 |
A |
1215 |
1106 |
0.18 |
|
|
|
| 28 |
A |
1151 |
1048 |
2.92 |
|
|
|
| 29 |
A |
894 |
814 |
1.13 |
|
|
|
| 30 |
A |
661 |
602 |
8.39 |
|
|
|
| 31 |
A |
260 |
236 |
11.40 |
|
|
|
| 32 |
A |
160 |
146 |
0.77 |
|
|
|
| 33 |
A |
59 |
54 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25605.0 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 23315.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.554 |
|
|
|
| 2 |
C |
1.004 |
|
|
|
| 3 |
C |
0.555 |
|
|
|
| 4 |
N |
-0.686 |
|
|
|
| 5 |
O |
-0.990 |
|
|
|
| 6 |
C |
0.352 |
|
|
|
| 7 |
H |
-0.204 |
|
|
|
| 8 |
H |
-0.204 |
|
|
|
| 9 |
H |
-0.190 |
|
|
|
| 10 |
H |
-0.190 |
|
|
|
| 11 |
H |
0.022 |
|
|
|
| 12 |
H |
0.022 |
|
|
|
| 13 |
H |
-0.045 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.098 |
-0.876 |
0.000 |
1.405 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.910 |
1.920 |
0.000 |
| y |
1.920 |
-29.945 |
0.000 |
| z |
0.000 |
0.000 |
-35.527 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.174 |
1.920 |
0.000 |
| y |
1.920 |
8.773 |
0.000 |
| z |
0.000 |
0.000 |
0.401 |
|
| Polar |
|---|
| 3z2-r2 | 0.801 |
|---|
| x2-y2 | -11.965 |
|---|
| xy | 1.920 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
147.347 |
| (<r2>)1/2 |
12.139 |