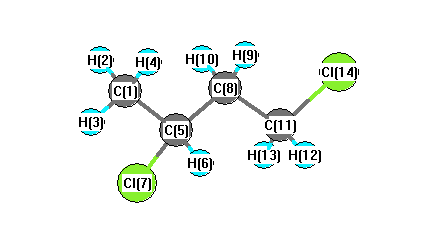
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3192 |
3058 |
3.09 |
|
|
|
| 2 |
A |
3170 |
3038 |
13.43 |
|
|
|
| 3 |
A |
3147 |
3016 |
19.73 |
|
|
|
| 4 |
A |
3130 |
3000 |
7.53 |
|
|
|
| 5 |
A |
3113 |
2983 |
10.97 |
|
|
|
| 6 |
A |
3107 |
2978 |
2.47 |
|
|
|
| 7 |
A |
3071 |
2943 |
14.72 |
|
|
|
| 8 |
A |
3061 |
2934 |
8.59 |
|
|
|
| 9 |
A |
1476 |
1415 |
3.19 |
|
|
|
| 10 |
A |
1471 |
1409 |
3.76 |
|
|
|
| 11 |
A |
1466 |
1405 |
10.14 |
|
|
|
| 12 |
A |
1457 |
1396 |
1.29 |
|
|
|
| 13 |
A |
1399 |
1341 |
10.21 |
|
|
|
| 14 |
A |
1388 |
1330 |
8.51 |
|
|
|
| 15 |
A |
1328 |
1273 |
7.56 |
|
|
|
| 16 |
A |
1310 |
1255 |
10.36 |
|
|
|
| 17 |
A |
1270 |
1217 |
14.95 |
|
|
|
| 18 |
A |
1248 |
1196 |
18.64 |
|
|
|
| 19 |
A |
1177 |
1128 |
9.90 |
|
|
|
| 20 |
A |
1128 |
1081 |
0.36 |
|
|
|
| 21 |
A |
1121 |
1075 |
7.62 |
|
|
|
| 22 |
A |
1076 |
1031 |
2.16 |
|
|
|
| 23 |
A |
1032 |
989 |
17.68 |
|
|
|
| 24 |
A |
956 |
916 |
1.17 |
|
|
|
| 25 |
A |
921 |
883 |
5.67 |
|
|
|
| 26 |
A |
778 |
746 |
19.28 |
|
|
|
| 27 |
A |
764 |
733 |
27.49 |
|
|
|
| 28 |
A |
631 |
605 |
28.30 |
|
|
|
| 29 |
A |
439 |
420 |
2.44 |
|
|
|
| 30 |
A |
412 |
395 |
5.32 |
|
|
|
| 31 |
A |
340 |
326 |
3.46 |
|
|
|
| 32 |
A |
250 |
240 |
0.38 |
|
|
|
| 33 |
A |
244 |
234 |
0.10 |
|
|
|
| 34 |
A |
151 |
145 |
2.09 |
|
|
|
| 35 |
A |
114 |
109 |
1.34 |
|
|
|
| 36 |
A |
73 |
70 |
3.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25204.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 24153.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.669 |
|
|
|
| 2 |
H |
-0.049 |
|
|
|
| 3 |
H |
-0.111 |
|
|
|
| 4 |
H |
-0.085 |
|
|
|
| 5 |
C |
-0.296 |
|
|
|
| 6 |
H |
-0.245 |
|
|
|
| 7 |
Cl |
-0.158 |
|
|
|
| 8 |
C |
0.839 |
|
|
|
| 9 |
H |
-0.208 |
|
|
|
| 10 |
H |
-0.215 |
|
|
|
| 11 |
C |
0.105 |
|
|
|
| 12 |
H |
-0.085 |
|
|
|
| 13 |
H |
-0.098 |
|
|
|
| 14 |
Cl |
-0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.415 |
1.747 |
1.030 |
2.473 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-59.535 |
-3.821 |
0.065 |
| y |
-3.821 |
-51.194 |
0.136 |
| z |
0.065 |
0.136 |
-50.406 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.735 |
-3.821 |
0.065 |
| y |
-3.821 |
3.777 |
0.136 |
| z |
0.065 |
0.136 |
4.958 |
|
| Polar |
|---|
| 3z2-r2 | 9.917 |
|---|
| x2-y2 | -8.341 |
|---|
| xy | -3.821 |
|---|
| xz | 0.065 |
|---|
| yz | 0.136 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.029 |
0.188 |
-0.158 |
| y |
0.188 |
11.632 |
0.030 |
| z |
-0.158 |
0.030 |
9.716 |
<r2> (average value of r
2) Å
2
| <r2> |
367.149 |
| (<r2>)1/2 |
19.161 |