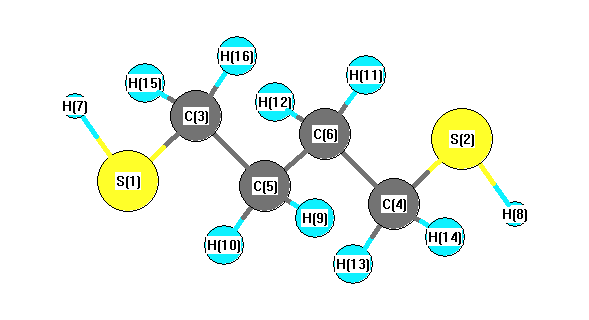
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3080 |
2951 |
0.00 |
|
|
|
| 2 |
Ag |
3050 |
2923 |
0.00 |
|
|
|
| 3 |
Ag |
2715 |
2602 |
0.00 |
|
|
|
| 4 |
Ag |
1474 |
1412 |
0.00 |
|
|
|
| 5 |
Ag |
1468 |
1407 |
0.00 |
|
|
|
| 6 |
Ag |
1394 |
1336 |
0.00 |
|
|
|
| 7 |
Ag |
1263 |
1210 |
0.00 |
|
|
|
| 8 |
Ag |
1120 |
1074 |
0.00 |
|
|
|
| 9 |
Ag |
1075 |
1031 |
0.00 |
|
|
|
| 10 |
Ag |
854 |
818 |
0.00 |
|
|
|
| 11 |
Ag |
777 |
744 |
0.00 |
|
|
|
| 12 |
Ag |
336 |
322 |
0.00 |
|
|
|
| 13 |
Ag |
206 |
197 |
0.00 |
|
|
|
| 14 |
Au |
3146 |
3015 |
29.95 |
|
|
|
| 15 |
Au |
3107 |
2977 |
14.68 |
|
|
|
| 16 |
Au |
1309 |
1255 |
0.81 |
|
|
|
| 17 |
Au |
1096 |
1050 |
3.25 |
|
|
|
| 18 |
Au |
888 |
851 |
1.67 |
|
|
|
| 19 |
Au |
736 |
706 |
3.77 |
|
|
|
| 20 |
Au |
193 |
185 |
28.34 |
|
|
|
| 21 |
Au |
98 |
94 |
6.13 |
|
|
|
| 22 |
Au |
51 |
48 |
5.50 |
|
|
|
| 23 |
Bg |
3143 |
3012 |
0.00 |
|
|
|
| 24 |
Bg |
3087 |
2958 |
0.00 |
|
|
|
| 25 |
Bg |
1321 |
1266 |
0.00 |
|
|
|
| 26 |
Bg |
1246 |
1194 |
0.00 |
|
|
|
| 27 |
Bg |
1036 |
993 |
0.00 |
|
|
|
| 28 |
Bg |
774 |
742 |
0.00 |
|
|
|
| 29 |
Bg |
191 |
183 |
0.00 |
|
|
|
| 30 |
Bg |
129 |
124 |
0.00 |
|
|
|
| 31 |
Bu |
3082 |
2954 |
58.81 |
|
|
|
| 32 |
Bu |
3057 |
2929 |
15.49 |
|
|
|
| 33 |
Bu |
2715 |
2602 |
6.76 |
|
|
|
| 34 |
Bu |
1488 |
1426 |
7.10 |
|
|
|
| 35 |
Bu |
1466 |
1405 |
3.78 |
|
|
|
| 36 |
Bu |
1337 |
1282 |
34.90 |
|
|
|
| 37 |
Bu |
1212 |
1162 |
19.76 |
|
|
|
| 38 |
Bu |
1071 |
1026 |
0.24 |
|
|
|
| 39 |
Bu |
885 |
848 |
1.89 |
|
|
|
| 40 |
Bu |
739 |
708 |
5.42 |
|
|
|
| 41 |
Bu |
390 |
374 |
4.47 |
|
|
|
| 42 |
Bu |
100 |
96 |
3.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28951.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 27743.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.199 |
|
|
|
| 2 |
S |
0.199 |
|
|
|
| 3 |
C |
0.237 |
|
|
|
| 4 |
C |
0.237 |
|
|
|
| 5 |
C |
0.556 |
|
|
|
| 6 |
C |
0.556 |
|
|
|
| 7 |
H |
0.095 |
|
|
|
| 8 |
H |
0.095 |
|
|
|
| 9 |
H |
-0.301 |
|
|
|
| 10 |
H |
-0.301 |
|
|
|
| 11 |
H |
-0.301 |
|
|
|
| 12 |
H |
-0.301 |
|
|
|
| 13 |
H |
-0.243 |
|
|
|
| 14 |
H |
-0.243 |
|
|
|
| 15 |
H |
-0.243 |
|
|
|
| 16 |
H |
-0.243 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.771 |
1.494 |
0.000 |
| y |
1.494 |
-66.763 |
0.000 |
| z |
0.000 |
0.000 |
-56.188 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
15.704 |
1.494 |
0.000 |
| y |
1.494 |
-15.784 |
0.000 |
| z |
0.000 |
0.000 |
0.079 |
|
| Polar |
|---|
| 3z2-r2 | 0.158 |
|---|
| x2-y2 | 20.992 |
|---|
| xy | 1.494 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.613 |
2.654 |
0.000 |
| y |
2.654 |
17.579 |
0.000 |
| z |
0.000 |
0.000 |
11.542 |
<r2> (average value of r
2) Å
2
| <r2> |
533.195 |
| (<r2>)1/2 |
23.091 |