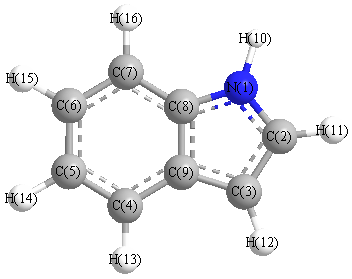
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3916 |
3566 |
94.19 |
|
|
|
| 2 |
A' |
3418 |
3112 |
4.24 |
|
|
|
| 3 |
A' |
3394 |
3091 |
1.53 |
|
|
|
| 4 |
A' |
3357 |
3057 |
20.23 |
|
|
|
| 5 |
A' |
3345 |
3046 |
32.78 |
|
|
|
| 6 |
A' |
3332 |
3034 |
3.59 |
|
|
|
| 7 |
A' |
3324 |
3027 |
1.03 |
|
|
|
| 8 |
A' |
1800 |
1639 |
4.80 |
|
|
|
| 9 |
A' |
1762 |
1605 |
0.73 |
|
|
|
| 10 |
A' |
1683 |
1533 |
20.07 |
|
|
|
| 11 |
A' |
1639 |
1493 |
5.25 |
|
|
|
| 12 |
A' |
1598 |
1456 |
33.09 |
|
|
|
| 13 |
A' |
1556 |
1417 |
31.67 |
|
|
|
| 14 |
A' |
1485 |
1352 |
5.00 |
|
|
|
| 15 |
A' |
1382 |
1259 |
0.00 |
|
|
|
| 16 |
A' |
1380 |
1256 |
47.47 |
|
|
|
| 17 |
A' |
1352 |
1231 |
10.80 |
|
|
|
| 18 |
A' |
1301 |
1185 |
4.22 |
|
|
|
| 19 |
A' |
1227 |
1117 |
1.97 |
|
|
|
| 20 |
A' |
1210 |
1102 |
4.71 |
|
|
|
| 21 |
A' |
1173 |
1068 |
26.68 |
|
|
|
| 22 |
A' |
1162 |
1059 |
20.13 |
|
|
|
| 23 |
A' |
1097 |
999 |
5.21 |
|
|
|
| 24 |
A' |
971 |
884 |
8.00 |
|
|
|
| 25 |
A' |
939 |
855 |
0.56 |
|
|
|
| 26 |
A' |
819 |
746 |
5.55 |
|
|
|
| 27 |
A' |
658 |
599 |
1.55 |
|
|
|
| 28 |
A' |
583 |
531 |
0.19 |
|
|
|
| 29 |
A' |
427 |
388 |
5.06 |
|
|
|
| 30 |
A" |
1073 |
977 |
0.01 |
|
|
|
| 31 |
A" |
1048 |
954 |
2.27 |
|
|
|
| 32 |
A" |
986 |
897 |
0.00 |
|
|
|
| 33 |
A" |
944 |
860 |
0.64 |
|
|
|
| 34 |
A" |
854 |
778 |
39.89 |
|
|
|
| 35 |
A" |
823 |
750 |
99.04 |
|
|
|
| 36 |
A" |
801 |
729 |
25.21 |
|
|
|
| 37 |
A" |
660 |
601 |
6.21 |
|
|
|
| 38 |
A" |
625 |
569 |
0.84 |
|
|
|
| 39 |
A" |
470 |
428 |
7.29 |
|
|
|
| 40 |
A" |
377 |
343 |
75.01 |
|
|
|
| 41 |
A" |
263 |
239 |
0.07 |
|
|
|
| 42 |
A" |
230 |
210 |
18.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30220.2 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 27518.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.685 |
|
|
|
| 2 |
C |
0.692 |
|
|
|
| 3 |
C |
0.586 |
|
|
|
| 4 |
C |
1.134 |
|
|
|
| 5 |
C |
0.547 |
|
|
|
| 6 |
C |
0.325 |
|
|
|
| 7 |
C |
0.924 |
|
|
|
| 8 |
C |
-0.714 |
|
|
|
| 9 |
C |
0.085 |
|
|
|
| 10 |
H |
-0.153 |
|
|
|
| 11 |
H |
-0.331 |
|
|
|
| 12 |
H |
-0.427 |
|
|
|
| 13 |
H |
-0.480 |
|
|
|
| 14 |
H |
-0.489 |
|
|
|
| 15 |
H |
-0.475 |
|
|
|
| 16 |
H |
-0.541 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.839 |
1.863 |
0.000 |
2.043 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.385 |
-2.004 |
0.000 |
| y |
-2.004 |
-42.858 |
0.000 |
| z |
0.000 |
0.000 |
-59.680 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.884 |
-2.004 |
0.000 |
| y |
-2.004 |
10.675 |
0.000 |
| z |
0.000 |
0.000 |
-14.559 |
|
| Polar |
|---|
| 3z2-r2 | -29.117 |
|---|
| x2-y2 | -4.527 |
|---|
| xy | -2.004 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
18.762 |
-1.494 |
0.000 |
| y |
-1.494 |
15.713 |
0.000 |
| z |
0.000 |
0.000 |
8.820 |
<r2> (average value of r
2) Å
2
| <r2> |
280.970 |
| (<r2>)1/2 |
16.762 |