Jump to
S2C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -152.058605 |
| Energy at 298.15K | |
| HF Energy | -152.058605 |
| Nuclear repulsion energy | 63.053794 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
2156 |
2066 |
0.00 |
64.00 |
0.51 |
0.67 |
| 2 |
Σg |
947 |
908 |
0.00 |
108.00 |
0.30 |
0.47 |
| 3 |
Σu |
1602 |
1535 |
324.98 |
0.00 |
0.00 |
0.00 |
| 4 |
Πg |
229 |
220 |
0.00 |
16.61 |
0.75 |
0.86 |
| 4 |
Πg |
229 |
220 |
0.00 |
16.61 |
0.75 |
0.86 |
| 5 |
Πu |
154 |
148 |
24.48 |
0.00 |
0.00 |
0.00 |
| 5 |
Πu |
154 |
148 |
24.48 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 2735.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 2621.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.647 |
| C2 |
0.000 |
0.000 |
-0.647 |
| C3 |
0.000 |
0.000 |
1.965 |
| C4 |
0.000 |
0.000 |
-1.965 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
| C1 | | 1.2949 | 1.3178 | 2.6127 |
C2 | 1.2949 | | 2.6127 | 1.3178 | C3 | 1.3178 | 2.6127 | | 3.9305 | C4 | 2.6127 | 1.3178 | 3.9305 | |
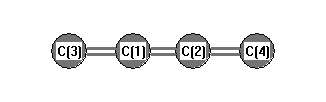 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
180.000 |
|
C2 |
C1 |
C3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.773 |
|
|
|
| 2 |
C |
0.773 |
|
|
|
| 3 |
C |
-0.773 |
|
|
|
| 4 |
C |
-0.773 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.161 |
0.000 |
0.000 |
| y |
0.000 |
-22.161 |
0.000 |
| z |
0.000 |
0.000 |
-33.145 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.492 |
0.000 |
0.000 |
| y |
0.000 |
5.492 |
0.000 |
| z |
0.000 |
0.000 |
-10.985 |
|
| Polar |
|---|
| 3z2-r2 | -21.969 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.715 |
0.000 |
0.000 |
| y |
0.000 |
4.715 |
0.000 |
| z |
0.000 |
0.000 |
13.968 |
<r2> (average value of r
2) Å
2
| <r2> |
67.505 |
| (<r2>)1/2 |
8.216 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -152.025433 |
| Energy at 298.15K | |
| HF Energy | -152.025433 |
| Nuclear repulsion energy | 62.964184 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
2158 |
2068 |
0.00 |
91.82 |
0.34 |
0.51 |
| 2 |
Σg |
944 |
905 |
0.00 |
90.18 |
0.24 |
0.38 |
| 3 |
Σu |
1613 |
1546 |
424.82 |
0.00 |
0.00 |
0.00 |
| 4 |
Πg |
392 |
376 |
0.00 |
0.21 |
0.75 |
0.86 |
| 4 |
Πg |
108i |
103i |
0.00 |
14.70 |
0.75 |
0.86 |
| 5 |
Πu |
193 |
185 |
6.73 |
0.00 |
0.00 |
0.00 |
| 5 |
Πu |
120 |
115 |
35.33 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 2656.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 2545.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.650 |
| C2 |
0.000 |
0.000 |
-0.650 |
| C3 |
0.000 |
0.000 |
1.968 |
| C4 |
0.000 |
0.000 |
-1.968 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
| C1 | | 1.2996 | 1.3179 | 2.6176 |
C2 | 1.2996 | | 2.6176 | 1.3179 | C3 | 1.3179 | 2.6176 | | 3.9355 | C4 | 2.6176 | 1.3179 | 3.9355 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.746 |
|
|
|
| 2 |
C |
0.746 |
|
|
|
| 3 |
C |
-0.746 |
|
|
|
| 4 |
C |
-0.746 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.123 |
0.000 |
0.000 |
| y |
0.000 |
-20.413 |
0.000 |
| z |
0.000 |
0.000 |
-32.886 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.526 |
0.000 |
0.000 |
| y |
0.000 |
8.092 |
0.000 |
| z |
0.000 |
0.000 |
-10.618 |
|
| Polar |
|---|
| 3z2-r2 | -21.235 |
|---|
| x2-y2 | -3.711 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.544 |
0.000 |
0.000 |
| y |
0.000 |
4.525 |
0.000 |
| z |
0.000 |
0.000 |
13.464 |
<r2> (average value of r
2) Å
2
| <r2> |
67.650 |
| (<r2>)1/2 |
8.225 |
