return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CHO (Propanal)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-192.671434
Energy at 298.15K-192.677723
HF Energy-191.975003
Nuclear repulsion energy117.232742
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3156 3083        
2 A' 3074 3003        
3 A' 3036 2966        
4 A' 2917 2849        
5 A' 1760 1719        
6 A' 1544 1509        
7 A' 1508 1474        
8 A' 1464 1430        
9 A' 1430 1397        
10 A' 1400 1368        
11 A' 1138 1112        
12 A' 1023 999        
13 A' 879 859        
14 A' 674 658        
15 A' 265 259        
16 A" 3161 3088        
17 A" 3072 3001        
18 A" 1542 1506        
19 A" 1305 1275        
20 A" 1169 1142        
21 A" 912 891        
22 A" 693 677        
23 A" 238 232        
24 A" 136 133        

Unscaled Zero Point Vibrational Energy (zpe) 18746.6 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 18313.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.54366 0.19341 0.15074

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.461 0.457 0.000
C2 0.000 0.927 0.000
C3 -1.007 -0.213 0.000
O4 -0.709 -1.406 0.000
H5 2.146 1.323 0.000
H6 1.673 -0.159 0.890
H7 1.673 -0.159 -0.890
H8 -0.229 1.559 0.883
H9 -0.229 1.559 -0.883
H10 -2.084 0.099 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.53462.55732.86011.10381.10351.10352.20232.20233.5636
C21.53461.52042.43792.18202.18402.18401.10961.10962.2426
C32.55731.52041.23013.50682.82422.82422.12612.12611.1217
O42.86012.43791.23013.94922.83172.83173.13013.13012.0392
H51.10382.18203.50683.94921.79281.79282.54482.54484.4036
H61.10352.18402.82422.83171.79281.78092.56303.11663.8698
H71.10352.18402.82422.83171.79281.78093.11662.56303.8698
H82.20231.10962.12613.13012.54482.56303.11661.76552.5198
H92.20231.10962.12613.13012.54483.11662.56301.76552.5198
H103.56362.24261.12172.03924.40363.86983.86982.51982.5198

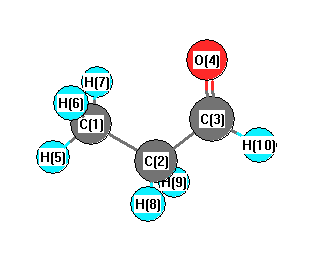
picture of Propanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 113.669 C1 C2 H8 111.781
C1 C2 H9 111.781 C2 C1 H5 110.523
C2 C1 H6 110.694 C2 C1 H7 110.694
C2 C3 O4 124.497 C2 C3 H10 115.335
C3 C2 H8 106.842 C3 C2 H9 106.842
O4 C3 H10 120.168 H5 C1 H6 108.624
H5 C1 H7 108.624 H6 C1 H7 107.593
H8 C2 H9 105.413
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability