Jump to
S1C2
S1C3
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.993297 |
| Energy at 298.15K | -207.999394 |
| HF Energy | -207.993297 |
| Nuclear repulsion energy | 120.193781 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3842 |
3499 |
41.07 |
|
|
|
| 2 |
A' |
3271 |
2978 |
28.35 |
|
|
|
| 3 |
A' |
3179 |
2895 |
73.89 |
|
|
|
| 4 |
A' |
3162 |
2879 |
49.56 |
|
|
|
| 5 |
A' |
1925 |
1753 |
698.63 |
|
|
|
| 6 |
A' |
1647 |
1500 |
28.17 |
|
|
|
| 7 |
A' |
1606 |
1463 |
4.17 |
|
|
|
| 8 |
A' |
1579 |
1438 |
5.65 |
|
|
|
| 9 |
A' |
1503 |
1369 |
8.85 |
|
|
|
| 10 |
A' |
1395 |
1271 |
190.55 |
|
|
|
| 11 |
A' |
1255 |
1143 |
43.16 |
|
|
|
| 12 |
A' |
1081 |
985 |
45.26 |
|
|
|
| 13 |
A' |
655 |
596 |
17.62 |
|
|
|
| 14 |
A' |
373 |
339 |
9.94 |
|
|
|
| 15 |
A" |
3236 |
2947 |
38.24 |
|
|
|
| 16 |
A" |
1582 |
1441 |
5.86 |
|
|
|
| 17 |
A" |
1238 |
1127 |
1.42 |
|
|
|
| 18 |
A" |
1164 |
1060 |
0.00 |
|
|
|
| 19 |
A" |
636 |
579 |
140.41 |
|
|
|
| 20 |
A" |
197 |
180 |
0.11 |
|
|
|
| 21 |
A" |
55 |
50 |
1.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17290.9 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 15745.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.289 |
-0.752 |
0.000 |
| O2 |
1.395 |
-1.210 |
0.000 |
| N3 |
0.000 |
0.563 |
0.000 |
| C4 |
-1.334 |
1.118 |
0.000 |
| H5 |
-0.603 |
-1.393 |
0.000 |
| H6 |
0.785 |
1.179 |
0.000 |
| H7 |
-2.059 |
0.307 |
0.000 |
| H8 |
-1.508 |
1.728 |
0.886 |
| H9 |
-1.508 |
1.728 |
-0.886 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.1973 | 1.3469 | 2.4760 | 1.0978 | 1.9940 | 2.5755 | 3.1883 | 3.1883 |
O2 | 1.1973 | | 2.2565 | 3.5871 | 2.0062 | 2.4660 | 3.7724 | 4.2244 | 4.2244 | N3 | 1.3469 | 2.2565 | | 1.4445 | 2.0467 | 0.9976 | 2.0745 | 2.1012 | 2.1012 | C4 | 2.4760 | 3.5871 | 1.4445 | | 2.6148 | 2.1196 | 1.0876 | 1.0898 | 1.0898 | H5 | 1.0978 | 2.0062 | 2.0467 | 2.6148 | | 2.9222 | 2.2380 | 3.3680 | 3.3680 | H6 | 1.9940 | 2.4660 | 0.9976 | 2.1196 | 2.9222 | | 2.9743 | 2.5185 | 2.5185 | H7 | 2.5755 | 3.7724 | 2.0745 | 1.0876 | 2.2380 | 2.9743 | | 1.7630 | 1.7630 | H8 | 3.1883 | 4.2244 | 2.1012 | 1.0898 | 3.3680 | 2.5185 | 1.7630 | | 1.7721 | H9 | 3.1883 | 4.2244 | 2.1012 | 1.0898 | 3.3680 | 2.5185 | 1.7630 | 1.7721 | |
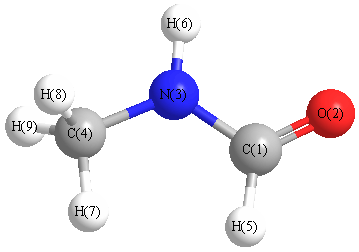 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.964 |
|
C1 |
N3 |
H6 |
115.728 |
| O2 |
C1 |
N3 |
124.874 |
|
O2 |
C1 |
H5 |
121.823 |
| N3 |
C1 |
H5 |
113.302 |
|
N3 |
C4 |
H7 |
109.218 |
| N3 |
C4 |
H8 |
111.245 |
|
N3 |
C4 |
H9 |
111.245 |
| C4 |
N3 |
H6 |
119.308 |
|
H7 |
C4 |
H8 |
108.123 |
| H7 |
C4 |
H9 |
108.123 |
|
H8 |
C4 |
H9 |
108.784 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.965 |
|
|
|
| 2 |
O |
-0.666 |
|
|
|
| 3 |
N |
-0.373 |
|
|
|
| 4 |
C |
0.766 |
|
|
|
| 5 |
H |
-0.185 |
|
|
|
| 6 |
H |
-0.157 |
|
|
|
| 7 |
H |
-0.159 |
|
|
|
| 8 |
H |
-0.096 |
|
|
|
| 9 |
H |
-0.096 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.521 |
2.774 |
0.000 |
4.483 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.503 |
4.737 |
0.000 |
| y |
4.737 |
-24.611 |
0.000 |
| z |
0.000 |
0.000 |
-24.958 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.718 |
4.737 |
0.000 |
| y |
4.737 |
1.620 |
0.000 |
| z |
0.000 |
0.000 |
1.098 |
|
| Polar |
|---|
| 3z2-r2 | 2.197 |
|---|
| x2-y2 | -2.892 |
|---|
| xy | 4.737 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.029 |
-1.003 |
0.000 |
| y |
-1.003 |
5.857 |
0.000 |
| z |
0.000 |
0.000 |
4.044 |
<r2> (average value of r
2) Å
2
| <r2> |
88.355 |
| (<r2>)1/2 |
9.400 |
Jump to
S1C1
S1C3
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.993297 |
| Energy at 298.15K | -207.999394 |
| HF Energy | -207.993297 |
| Nuclear repulsion energy | 120.193781 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -207.995258 |
| Energy at 298.15K | -208.001286 |
| HF Energy | -207.995258 |
| Nuclear repulsion energy | 122.445406 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3891 |
3543 |
46.22 |
|
|
|
| 2 |
A |
3287 |
2994 |
20.15 |
|
|
|
| 3 |
A |
3260 |
2969 |
32.22 |
|
|
|
| 4 |
A |
3189 |
2904 |
34.32 |
|
|
|
| 5 |
A |
3171 |
2888 |
99.89 |
|
|
|
| 6 |
A |
1916 |
1744 |
463.87 |
|
|
|
| 7 |
A |
1675 |
1525 |
206.70 |
|
|
|
| 8 |
A |
1608 |
1464 |
2.80 |
|
|
|
| 9 |
A |
1581 |
1439 |
11.87 |
|
|
|
| 10 |
A |
1562 |
1423 |
12.16 |
|
|
|
| 11 |
A |
1526 |
1390 |
11.59 |
|
|
|
| 12 |
A |
1328 |
1210 |
94.62 |
|
|
|
| 13 |
A |
1264 |
1151 |
5.25 |
|
|
|
| 14 |
A |
1240 |
1129 |
2.51 |
|
|
|
| 15 |
A |
1156 |
1053 |
1.31 |
|
|
|
| 16 |
A |
1022 |
931 |
19.95 |
|
|
|
| 17 |
A |
826 |
753 |
10.20 |
|
|
|
| 18 |
A |
494 |
450 |
18.88 |
|
|
|
| 19 |
A |
294 |
267 |
14.40 |
|
|
|
| 20 |
A |
230 |
209 |
101.17 |
|
|
|
| 21 |
A |
59 |
54 |
9.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17289.2 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 15743.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.865 |
0.421 |
0.004 |
| O2 |
1.373 |
-0.663 |
-0.000 |
| N3 |
-0.462 |
0.649 |
-0.022 |
| C4 |
-1.430 |
-0.430 |
0.005 |
| H5 |
1.459 |
1.343 |
0.021 |
| H6 |
-0.774 |
1.589 |
0.060 |
| H7 |
-2.416 |
-0.019 |
-0.193 |
| H8 |
-1.190 |
-1.164 |
-0.760 |
| H9 |
-1.441 |
-0.933 |
0.972 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.1979 | 1.3462 | 2.4474 | 1.0970 | 2.0126 | 3.3157 | 2.7050 | 2.8436 |
O2 | 1.1979 | | 2.2561 | 2.8129 | 2.0081 | 3.1121 | 3.8487 | 2.7201 | 2.9897 | N3 | 1.3462 | 2.2561 | | 1.4498 | 2.0432 | 0.9939 | 2.0718 | 2.0878 | 2.1092 | C4 | 2.4474 | 2.8129 | 1.4498 | | 3.3899 | 2.1237 | 1.0871 | 1.0866 | 1.0897 | H5 | 1.0970 | 2.0081 | 2.0432 | 3.3899 | | 2.2469 | 4.1129 | 3.7298 | 3.8073 | H6 | 2.0126 | 3.1121 | 0.9939 | 2.1237 | 2.2469 | | 2.3116 | 2.9020 | 2.7632 | H7 | 3.3157 | 3.8487 | 2.0718 | 1.0871 | 4.1129 | 2.3116 | | 1.7707 | 1.7737 | H8 | 2.7050 | 2.7201 | 2.0878 | 1.0866 | 3.7298 | 2.9020 | 1.7707 | | 1.7653 | H9 | 2.8436 | 2.9897 | 2.1092 | 1.0897 | 3.8073 | 2.7632 | 1.7737 | 1.7653 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
122.123 |
|
C1 |
N3 |
H6 |
117.866 |
| O2 |
C1 |
N3 |
124.851 |
|
O2 |
C1 |
H5 |
122.036 |
| N3 |
C1 |
H5 |
113.111 |
|
N3 |
C4 |
H7 |
108.662 |
| N3 |
C4 |
H8 |
109.976 |
|
N3 |
C4 |
H9 |
111.520 |
| C4 |
N3 |
H6 |
119.535 |
|
H7 |
C4 |
H8 |
109.098 |
| H7 |
C4 |
H9 |
109.135 |
|
H8 |
C4 |
H9 |
108.416 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.882 |
|
|
|
| 2 |
O |
-0.656 |
|
|
|
| 3 |
N |
-0.381 |
|
|
|
| 4 |
C |
0.767 |
|
|
|
| 5 |
H |
-0.146 |
|
|
|
| 6 |
H |
-0.124 |
|
|
|
| 7 |
H |
-0.176 |
|
|
|
| 8 |
H |
-0.099 |
|
|
|
| 9 |
H |
-0.066 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.923 |
2.984 |
0.146 |
4.180 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.681 |
3.295 |
-0.050 |
| y |
3.295 |
-22.708 |
0.331 |
| z |
-0.050 |
0.331 |
-25.034 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.810 |
3.295 |
-0.050 |
| y |
3.295 |
2.649 |
0.331 |
| z |
-0.050 |
0.331 |
-0.840 |
|
| Polar |
|---|
| 3z2-r2 | -1.679 |
|---|
| x2-y2 | -2.973 |
|---|
| xy | 3.295 |
|---|
| xz | -0.050 |
|---|
| yz | 0.331 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.168 |
-0.127 |
-0.007 |
| y |
-0.127 |
5.513 |
-0.014 |
| z |
-0.007 |
-0.014 |
4.098 |
<r2> (average value of r
2) Å
2
| <r2> |
77.439 |
| (<r2>)1/2 |
8.800 |
