Jump to
S1C2
S1C3
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.034334 |
| Energy at 298.15K | |
| HF Energy | -208.811722 |
| Nuclear repulsion energy | 118.732690 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3614 |
3478 |
30.42 |
|
|
|
| 2 |
A' |
3154 |
3035 |
14.34 |
|
|
|
| 3 |
A' |
3054 |
2939 |
54.82 |
|
|
|
| 4 |
A' |
2985 |
2872 |
73.15 |
|
|
|
| 5 |
A' |
1755 |
1689 |
512.65 |
|
|
|
| 6 |
A' |
1526 |
1468 |
16.67 |
|
|
|
| 7 |
A' |
1476 |
1420 |
3.68 |
|
|
|
| 8 |
A' |
1455 |
1401 |
5.96 |
|
|
|
| 9 |
A' |
1384 |
1332 |
12.94 |
|
|
|
| 10 |
A' |
1304 |
1255 |
112.06 |
|
|
|
| 11 |
A' |
1167 |
1123 |
27.64 |
|
|
|
| 12 |
A' |
1011 |
973 |
38.86 |
|
|
|
| 13 |
A' |
607 |
584 |
12.94 |
|
|
|
| 14 |
A' |
342 |
329 |
7.80 |
|
|
|
| 15 |
A" |
3120 |
3003 |
23.53 |
|
|
|
| 16 |
A" |
1474 |
1418 |
5.91 |
|
|
|
| 17 |
A" |
1149 |
1105 |
0.54 |
|
|
|
| 18 |
A" |
1027 |
988 |
3.05 |
|
|
|
| 19 |
A" |
610 |
587 |
110.89 |
|
|
|
| 20 |
A" |
200 |
192 |
0.32 |
|
|
|
| 21 |
A" |
91 |
87 |
0.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16252.0 cm
-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 15639.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.281 |
-0.759 |
0.000 |
| O2 |
1.409 |
-1.237 |
0.000 |
| N3 |
0.000 |
0.573 |
0.000 |
| C4 |
-1.339 |
1.136 |
0.000 |
| H5 |
-0.634 |
-1.388 |
0.000 |
| H6 |
0.802 |
1.191 |
0.000 |
| H7 |
-2.065 |
0.315 |
0.000 |
| H8 |
-1.514 |
1.750 |
0.894 |
| H9 |
-1.514 |
1.750 |
-0.894 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2248 | 1.3619 | 2.4935 | 1.1100 | 2.0186 | 2.5800 | 3.2126 | 3.2126 |
O2 | 1.2248 | | 2.2939 | 3.6307 | 2.0483 | 2.5025 | 3.8045 | 4.2742 | 4.2742 | N3 | 1.3619 | 2.2939 | | 1.4523 | 2.0609 | 1.0125 | 2.0807 | 2.1161 | 2.1161 | C4 | 2.4935 | 3.6307 | 1.4523 | | 2.6207 | 2.1417 | 1.0963 | 1.0983 | 1.0983 | H5 | 1.1100 | 2.0483 | 2.0609 | 2.6207 | | 2.9516 | 2.2240 | 3.3796 | 3.3796 | H6 | 2.0186 | 2.5025 | 1.0125 | 2.1417 | 2.9516 | | 2.9978 | 2.5454 | 2.5454 | H7 | 2.5800 | 3.8045 | 2.0807 | 1.0963 | 2.2240 | 2.9978 | | 1.7780 | 1.7780 | H8 | 3.2126 | 4.2742 | 2.1161 | 1.0983 | 3.3796 | 2.5454 | 1.7780 | | 1.7870 | H9 | 3.2126 | 4.2742 | 2.1161 | 1.0983 | 3.3796 | 2.5454 | 1.7780 | 1.7870 | |
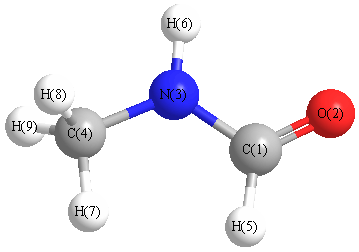 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.728 |
|
C1 |
N3 |
H6 |
115.677 |
| O2 |
C1 |
N3 |
124.868 |
|
O2 |
C1 |
H5 |
122.564 |
| N3 |
C1 |
H5 |
112.568 |
|
N3 |
C4 |
H7 |
108.654 |
| N3 |
C4 |
H8 |
111.368 |
|
N3 |
C4 |
H9 |
111.368 |
| C4 |
N3 |
H6 |
119.595 |
|
H7 |
C4 |
H8 |
108.230 |
| H7 |
C4 |
H9 |
108.230 |
|
H8 |
C4 |
H9 |
108.889 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.034334 |
| Energy at 298.15K | |
| HF Energy | -208.811722 |
| Nuclear repulsion energy | 118.732690 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -209.036298 |
| Energy at 298.15K | |
| HF Energy | -208.813526 |
| Nuclear repulsion energy | 120.986335 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3652 |
3514 |
28.96 |
|
|
|
| 2 |
A |
3180 |
3060 |
4.39 |
|
|
|
| 3 |
A |
3139 |
3021 |
21.05 |
|
|
|
| 4 |
A |
3060 |
2945 |
39.32 |
|
|
|
| 5 |
A |
3000 |
2887 |
97.39 |
|
|
|
| 6 |
A |
1744 |
1678 |
347.76 |
|
|
|
| 7 |
A |
1553 |
1495 |
111.02 |
|
|
|
| 8 |
A |
1495 |
1439 |
7.40 |
|
|
|
| 9 |
A |
1477 |
1421 |
18.33 |
|
|
|
| 10 |
A |
1436 |
1382 |
15.22 |
|
|
|
| 11 |
A |
1406 |
1353 |
6.93 |
|
|
|
| 12 |
A |
1231 |
1184 |
61.41 |
|
|
|
| 13 |
A |
1164 |
1120 |
0.92 |
|
|
|
| 14 |
A |
1147 |
1104 |
9.29 |
|
|
|
| 15 |
A |
1015 |
977 |
0.45 |
|
|
|
| 16 |
A |
965 |
929 |
17.61 |
|
|
|
| 17 |
A |
763 |
734 |
3.67 |
|
|
|
| 18 |
A |
513 |
493 |
32.92 |
|
|
|
| 19 |
A |
283 |
272 |
12.10 |
|
|
|
| 20 |
A |
259 |
249 |
74.50 |
|
|
|
| 21 |
A |
16 |
16 |
0.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16247.9 cm
-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 15635.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.867 |
0.440 |
0.003 |
| O2 |
1.382 |
-0.675 |
0.000 |
| N3 |
-0.474 |
0.659 |
-0.020 |
| C4 |
-1.429 |
-0.443 |
0.005 |
| H5 |
1.462 |
1.374 |
0.018 |
| H6 |
-0.801 |
1.610 |
0.059 |
| H7 |
-2.429 |
-0.050 |
-0.207 |
| H8 |
-1.159 |
-1.180 |
-0.760 |
| H9 |
-1.434 |
-0.947 |
0.981 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2278 | 1.3594 | 2.4600 | 1.1077 | 2.0387 | 3.3388 | 2.7033 | 2.8587 |
O2 | 1.2278 | | 2.2859 | 2.8206 | 2.0504 | 3.1610 | 3.8675 | 2.6995 | 2.9939 | N3 | 1.3594 | 2.2859 | | 1.4582 | 2.0646 | 1.0091 | 2.0877 | 2.0971 | 2.1212 | C4 | 2.4600 | 2.8206 | 1.4582 | | 3.4148 | 2.1480 | 1.0950 | 1.0959 | 1.0986 | H5 | 1.1077 | 2.0504 | 2.0646 | 3.4148 | | 2.2759 | 4.1495 | 3.7410 | 3.8339 | H6 | 2.0387 | 3.1610 | 1.0091 | 2.1480 | 2.2759 | | 2.3403 | 2.9300 | 2.7909 | H7 | 3.3388 | 3.8675 | 2.0877 | 1.0950 | 4.1495 | 2.3403 | | 1.7882 | 1.7904 | H8 | 2.7033 | 2.6995 | 2.0971 | 1.0959 | 3.7410 | 2.9300 | 1.7882 | | 1.7778 | H9 | 2.8587 | 2.9939 | 2.1212 | 1.0986 | 3.8339 | 2.7909 | 1.7904 | 1.7778 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.597 |
|
C1 |
N3 |
H6 |
118.056 |
| O2 |
C1 |
N3 |
124.071 |
|
O2 |
C1 |
H5 |
122.706 |
| N3 |
C1 |
H5 |
113.223 |
|
N3 |
C4 |
H7 |
108.866 |
| N3 |
C4 |
H8 |
109.567 |
|
N3 |
C4 |
H9 |
111.347 |
| C4 |
N3 |
H6 |
119.945 |
|
H7 |
C4 |
H8 |
109.411 |
| H7 |
C4 |
H9 |
109.410 |
|
H8 |
C4 |
H9 |
108.217 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
